Original paper: Released original PDF, physical pages 71–77.
Write , and . The product rule applied to a smooth test function givesThe mixed second partial derivatives cancel because they commute. Since the orbital angular momentum components are , their commutator isThis is one of the orbital angular momentum commutation relations; the Planck constant enters through the definition of each linear operator.
Cyclic permutation of the coordinates in the preceding commutator calculation gives the remaining orbital angular momentum commutation relations:Reversing the order of a commutator reverses its sign.
Use the commutator derivation identity and the orbital angular momentum commutation relations. They giveThe first two expressions cancel on addition, soThe same argument works for each component of the angular momentum operator.
The squared angular momentum operator and one selected component are commuting operators, so a complete set of common eigenfunctions can be chosen: for the component these are the usual angular momentum eigenstates. Distinct components do not have a common complete eigenbasis, because their commutator is another nonzero component. A state is an exception in which all components vanish; noncommutation does not forbid every individual common eigenfunction.
The squared angular momentum and one chosen component can have simultaneously sharp values. All three components cannot have simultaneously sharp values in a nonzero- state. The possible measured component values are , while the squared magnitude is . This concerns the quantum state, rather than imperfect experimental resolution. The Robertson uncertainty relation, for example,quantifies the restriction; its right-hand side may vanish in some states, without making all three components simultaneously sharp.
It represents the squared orbital angular momentum. In atomic units, , so its eigenvalues are . In ordinary units the corresponding observable is , with orbital angular momentum quantum number .
The spherical harmonic is also an eigenfunction of the component of orbital angular momentum:These are atomic units; restoring units gives . The integer is the magnetic quantum number.
Separation of variables makes the angular factor a spherical harmonic. Applying the Quantum Hamiltonian givesConsequently is an energy eigenfunction provided the radial factor satisfiesThis is the radial Schrödinger equation for a central potential, here with the attractive Coulomb potential. An arbitrary does not give an eigenfunction; regularity, decay and wavefunction normalization select admissible bound quantum states.
For a atomic orbital, . The radial factor obeysSubstitution into the radial Schrödinger equation for a central potential givesThus the energy eigenvalue isThe normalization constant cancels; is the Hartree energy.
The quantum variational principle says that every admissible nonzero trial wavefunction obeyswhere is the ground-state energy. Thus the Rayleigh quotient is an upper bound. Equality holds when the trial lies entirely in the ground-state subspace.
Expand the trial in a complete orthonormal eigenbasis of the Quantum Hamiltonian, , with . Orthonormality givesEvery summand in the numerator is nonnegative because . This proves the quantum variational principle for the stated discrete expansion; the trial energy is a weighted mean of exact energy eigenvalues.
Inside the particle in a box, the Quantum Hamiltonian isThe infinite potential outside imposes the Dirichlet boundary conditions and a zero wavefunction outside the interval. These boundary conditions are part of the Hamiltonian operator, rather than an optional addition.
Twice differentiation the sine givesBoth Dirichlet boundary conditions hold when is an integer. The zero function at is not an eigenfunction, and negative merely changes an overall sign. ThereforeThe corresponding normalized eigenfunction is .
The trial already vanishes at . The second Dirichlet boundary condition requires , hence for a box of positive length. This makes the trial an admissible particle in a box wavefunction.
Up to a global phase, is real, positive throughout the interior, has no interior nodes of a wavefunction, and is symmetric about . It is smooth inside the box and has one central maximum. These are the shape and reflection symmetry properties of the lowest particle in a box eigenfunction, ; they make it a plausible variational method trial without making it exact.
Sincean explicitly normalized trial isIt is zero outside the box. The original expression differs by a minus sign, which changes only the global phase and leaves probabilities and expectation values unchanged.
Integration by parts and the zero endpoint values turn the Rayleigh quotient into a positive kinetic energy integral:This is the parabolic variational trial for a particle in a box. The potential energy vanishes inside the box.
Comparing the variational method estimate with the exact ground-state energy givesThe estimate is about too high, consistently with the quantum variational principle. Its closeness does not mean that the trial wavefunction is an exact energy eigenfunction.
The trial is even under reflection about the midpoint. The normalized box eigenfunctions satisfy , so orthogonality between even and odd functions gives for even . If the higher odd contributions are neglected, the weighted mean from the quantum variational principle becomesRearranging yields the two-level variational contamination estimateThe required assumption concerns the neglected contributions to the energy as well as their small probabilities. Numerically the two-level estimate is . It is not an exact identity: direct expansion of this parabolic trial gives for odd , hence the exact ratio is . The omitted higher odd states account for the difference.
Place the carbon and sulfur nuclei on a line separated by , with masses and . The center of mass lies between them and closer to the heavier sulfur nucleus. In the linear rigid rotor approximation, remains fixed while the molecule rotates.
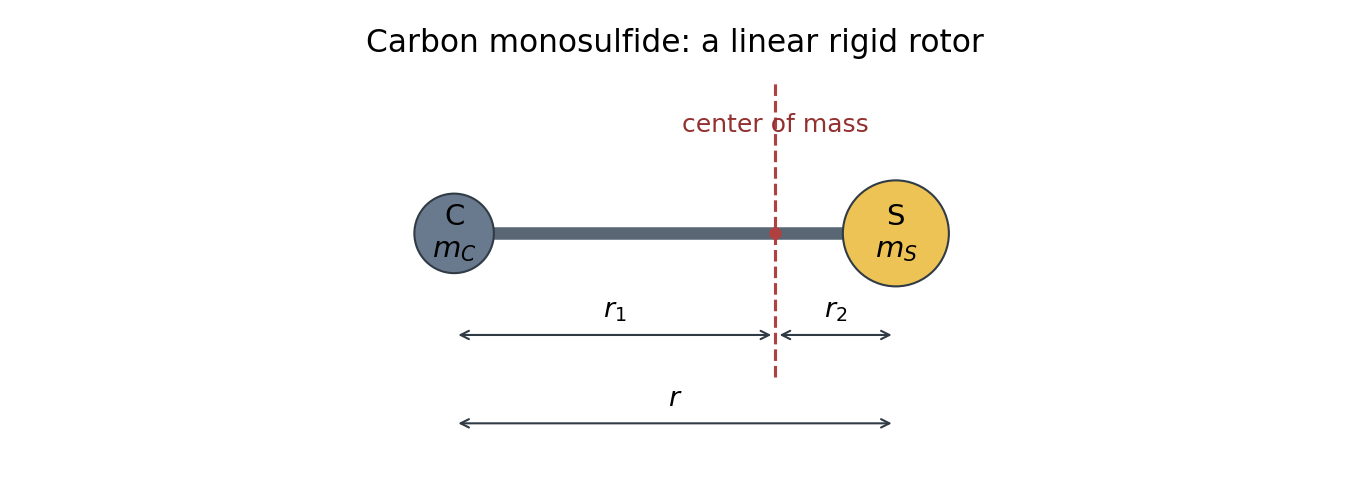
Original carbon monosulfide rotor diagram showing masses, bond length and distances from the center of mass
. Details. Measure from the carbon nucleus toward the sulfur nucleus. The definition of the center of mass givesTherefore the requested distance isThe other distance is , consistent with .
For an axis through the center of mass perpendicular to the bond, the point-mass moment of inertia isThe prefactor is the reduced mass ; this is the standard reduction of a two-body linear rigid rotor to .
Using the stipulated integer isotope masses givesThe rotational constant in wavenumber units isHere must be expressed in when is in SI units. Equivalently calculate in first, then divide by in .
The rotational state populations start at . Their statistical weight factor initially favors higher , while the Boltzmann factor eventually suppresses them, producing one maximum followed by a tail tending to zero. The physically allowed values occur at integer ; a continuous curve only guides the eye.
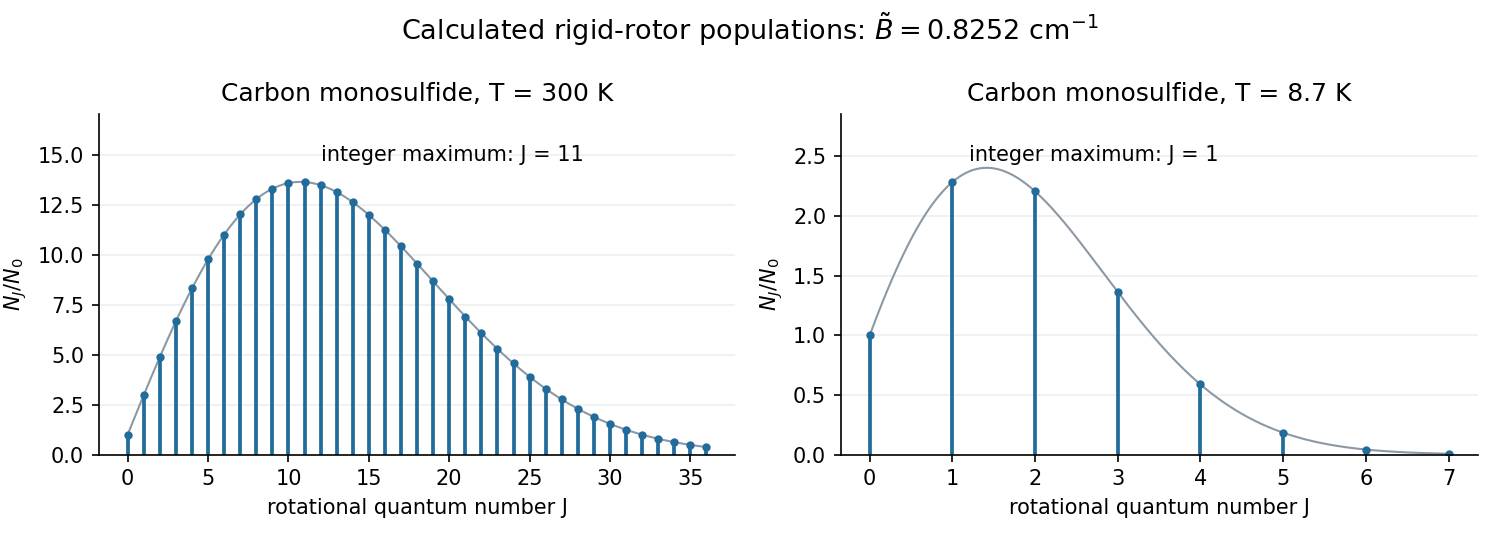
Set , the rotational temperature. For a continuous nonnegative variable ,Equating this derivative to zero gives the most populated rotational levelThe second derivative is , so this is a maximum when . At lower temperatures the constrained continuous maximum is . For the physical integer maximum compare the nearby allowed levels explicitly.
The calculated rotational constant gives . At room temperature, taken as , the continuous maximum is ; comparing the adjacent integer populations gives . Taking instead gives and the same integer answer.
At , . In particular,and , so . The integer comparison avoids treating a noninteger angular momentum quantum number as a physical state.
Low rotational energy spacings can be excited in cold interstellar gas when electronic and most vibrational excitations are frozen out. A polar molecule such as carbon monosulfide has allowed electric dipole transition lines whose frequencies depend on its moment of inertia, providing a distinctive pattern for rotational spectral line assignment. Several matching spectral lines and their isotope shifts give a much stronger identification than one coincidence. The method is especially effective for polar species; a molecule without a permanent electric dipole moment lacks an ordinary pure electric-dipole rotational spectrum.
The trigonal pyramidal molecular geometry has C3v point group symmetry: identity, two rotations about the threefold axis, and three vertical mirror reflections. Choose along the threefold axis through nitrogen and the center of the hydrogen triangle.
In the C3v point group, the nitrogen valence atomic orbitals transform asThe orbital is invariant under every symmetry operation; is also invariant because each operation preserves the coordinate. The other two components mix as a two-dimensional irreducible representation.
Use the classes of the C3v point group. For the three hydrogen atomic orbitals, the character of a representation counts orbitals fixed by each symmetry operation. Identity fixes three, a nontrivial threefold rotation fixes none, and a vertical mirror fixes one. ThereforeThe character inner product gives multiplicitiesThus , with the contribution accounting for two independent symmetry orbitals.
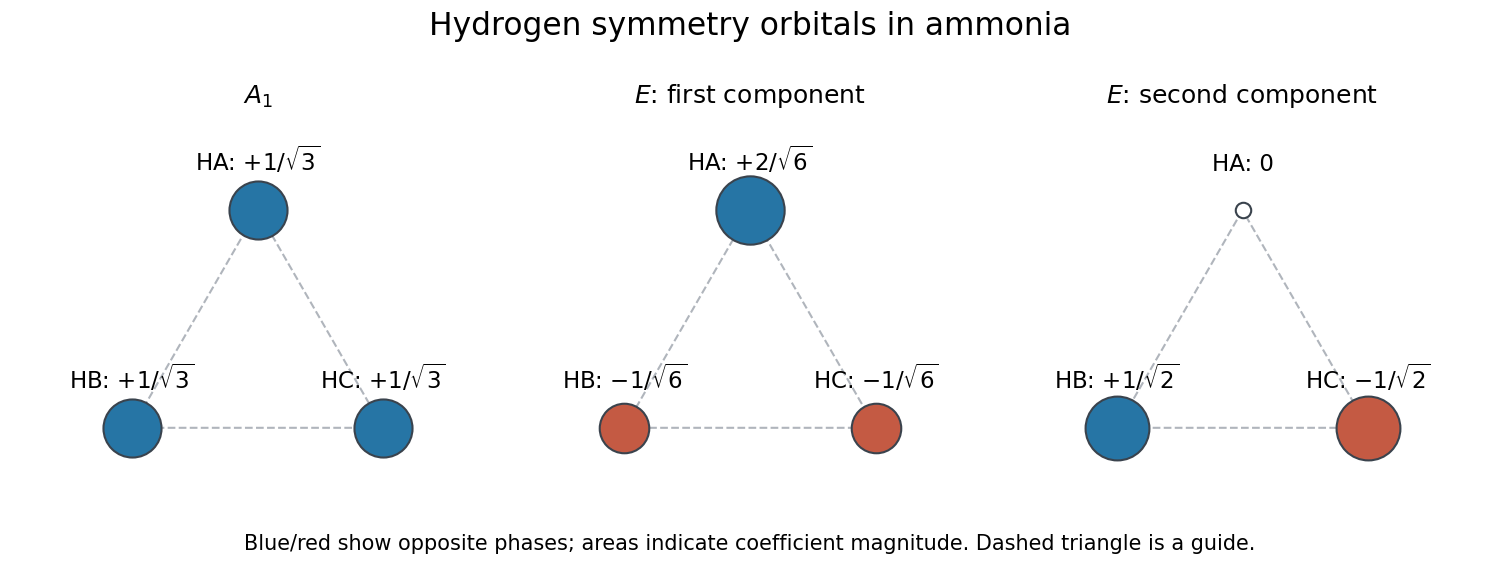
Neglecting intersite orbital overlap, one normalized real set of symmetry-adapted linear combinations isThe coefficient vectors are orthonormal. The first has equal phase at all three sites; the second opposes site to ; the third has zero amplitude at and opposite phases at . Any orthonormal rotation of the two functions is equally valid. With nonzero intersite orbital overlap, normalization must instead use the overlap matrix.
The hydrogen symmetry orbital mixes with both nitrogen and , giving three molecular orbitals. The two hydrogen functions mix with , giving a bonding pair and an antibonding pair. Eight valence electrons fill the lower levels asThis uses conventional numbering that reserves for the omitted nitrogen core orbital. The low and levels provide the principal N–H bonding molecular orbitals; the highest occupied level is the mainly nitrogen lone pair, with some bonding admixture permitted by the pyramidal geometry. Empty and levels are antibonding. Symmetry fixes which orbitals can mix, rather than their quantitative energy separations.
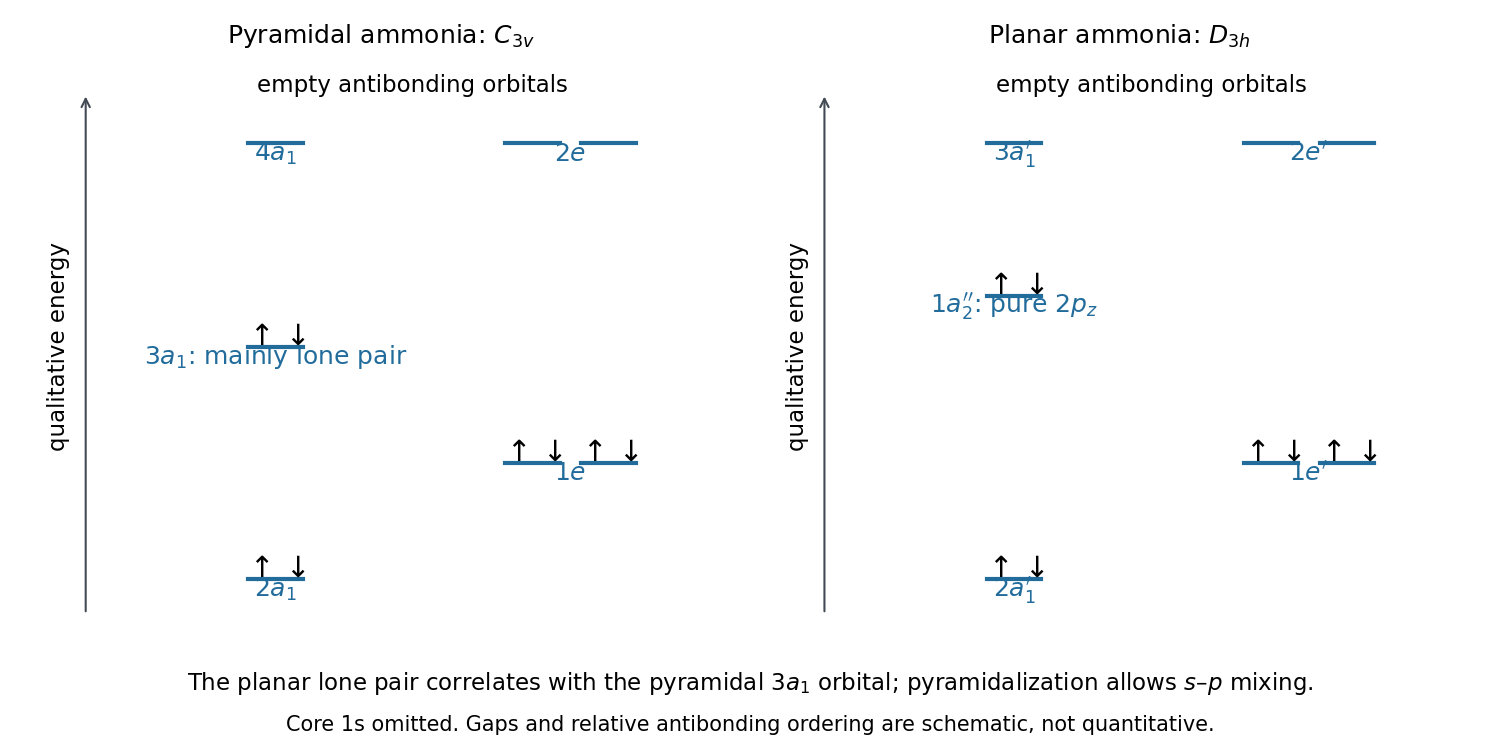
Original qualitative ammonia molecular orbital diagram comparing pyramidal and planar geometries; the core orbital is omitted
. Details. The planar equilateral structure has D3h point group symmetry. In addition to the threefold axis, it has a horizontal mirror plane containing all four nuclei and three twofold axes lying in that plane.
For planar ammonia, the nitrogen valence atomic orbitals have symmetries , and . The hydrogen symmetry orbitals span . Consequently the in-plane orbitals form a bonding orbital, a bonding pair and their antibonding partners. The out-of-plane has no matching hydrogen combination and remains a nonbonding orbital. With the core omitted, the conventional valence occupation isThe preceding ammonia molecular orbital diagram shows the correlation. On pyramidalization, and both correlate with of the C3v point group, permitting s–p orbital mixing. The pure planar lone pair can acquire bonding character, lowering its energy and helping stabilize the pyramid. Symmetry alone does not provide a numerical inversion barrier.[ref]
The nitrogen-centered highest occupied molecular orbital of ammonia supplies a lone pair; planar borane has an empty out-of-plane -derived lowest unoccupied molecular orbital. Approach along this acceptor axis gives constructive orbital overlap and donation of the nitrogen pair into the boron-centered acceptor. The resulting N-to-B coordinate covalent bond forms the ammonia borane adduct.
The occupied molecular orbital formed by this interaction is bonding, while the corresponding antibonding combination remains empty. Borane pyramidalizes as coordination increases from three to four. Ammonia can use its existing outward-pointing lone pair; its inversion to the planar transition geometry is not a prerequisite. The Lewis base and Lewis acid descriptions summarize the donor and acceptor roles, while the frontier molecular orbitals explain their directional interaction.
A symmetry orbital is a linear combination of atomic orbitals transforming as a component of an irreducible representation of the molecular point group. A molecular orbital is a one-electron wavefunction for the molecule, obtained as an eigenfunction of the chosen effective one-electron Hamiltonian operator. Symmetry organizes the eigenproblem, but a symmetry label alone does not determine its coefficients.
A symmetry orbital is a molecular orbital when it is also an eigenfunction of the effective one-electron Hamiltonian operator. If an irreducible representation occurs once in a complete invariant model space, a symmetry-invariant Hamiltonian acts as a scalar on that copy by Schur's lemma. Its basis functions are then degenerate eigenfunctions. When the same symmetry occurs in several copies, those copies can mix; one must diagonalize the corresponding block. Merely having a one-dimensional symmetry label is insufficient if other orbitals have that same label.
The axis is a axis through the centers of opposite cube faces: a half turn permutes atoms within each species. A body diagonal through sites 1 and 8 is a axis: a third turn fixes those two opposite vertices and cycles the other three vertices of each tetrahedral sublattice. These are symmetry operations of the tetrahedral symmetry group ; a quarter turn about would interchange the two chemical species and is not a symmetry of the specified cluster.
For the tetrahedral symmetry classes , each set of four atomic orbitals hasThe counts are the fixed sites of each symmetry operation: a threefold rotation fixes one vertex of each tetrahedron, a twofold or improper fourfold operation fixes none, and a diagonal mirror fixes two. Character orthogonality reduces each set toThe dimensions provide a check. Thus the complete eight-orbital model contains two copies of each of and , which can mix across the two species.
Write for the lithium orbital at site and for the sodium orbital. Reading the printed axes, the lithium coordinates have signs , , and ; the sodium sites are , , and . The normalized tetrahedral s-orbital symmetry combinations areEach row takes its signs from the corresponding coordinate, so the three functions transform together like . These coefficient vectors are orthonormal when intersite orbital overlap is neglected. A common sign reversal of any row is physically immaterial.
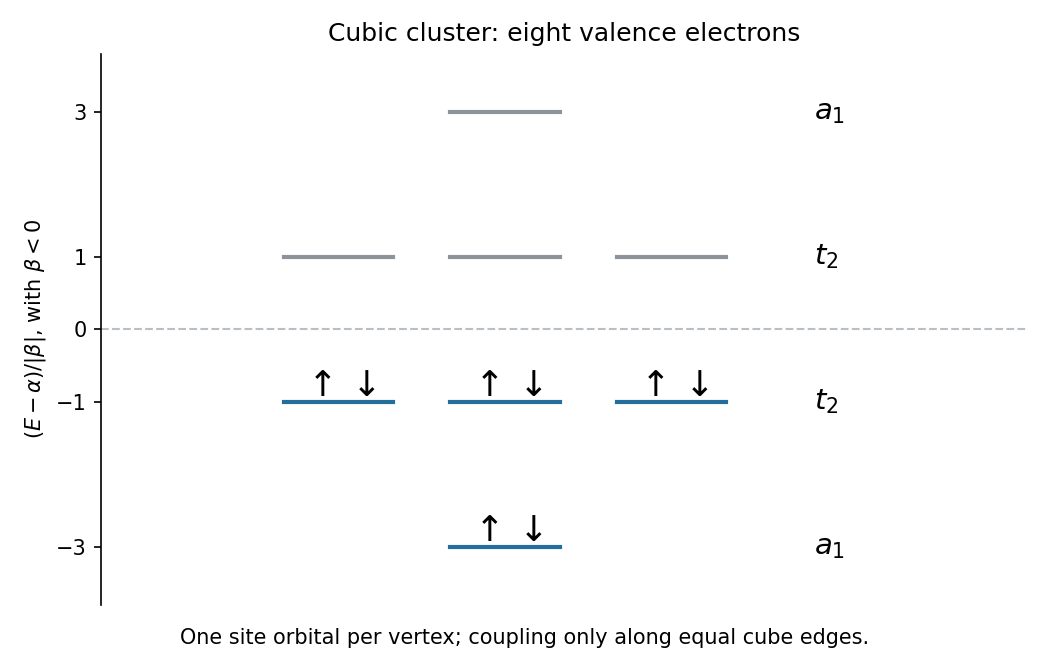
Use the stipulated common Hückel Coulomb parameter , an equal Hückel resonance integral along each cube edge, and zero coupling elsewhere; take an orthonormal site basis. Each lithium vertex has three sodium neighbors. The totally symmetric combinations therefore have the blockFor a coordinate-like combination, the sum of the three neighboring sodium coordinate signs equals the lithium coordinate sign. Thus each of the three equivalent blocks isTheir eigenvalues give the cube nearest-neighbour Hückel model:For the bonding convention these are in increasing energy order, with statistical weights . The eight valence electrons occupy , filling the four bonding spatial molecular orbitals. Their total one-electron energy is . This is a deliberately simplified model: unequal atomic energies or unequal overlaps require a generalized calculation.


Articles by others on the same topic
There are currently no matching articles.