Source bundle pages 129–133. The cover establishes 2022 despite the attachment filename. Solutions use the paper's rounded numerical data.
In LS coupling, first add the individual orbital angular momenta and spins separately, then add their totals:The Coulomb interaction between electrons must separate different atomic terms more strongly than the spin-orbit interaction mixes them. Then are useful quantum numbers, with . This approximation is usually better for lighter chemical atoms and well-separated atomic terms; strong configuration interaction or strong spin-orbit coupling can spoil it. The term symbol is .
The general electric-dipole selection rules, for isolated chemical atoms with definite parity and total electronic angular momentum, are a change of parity, excluding , and , with the accessible magnetic sublevels determined by the light's polarization. In pure LS coupling, the additional selection rules are and , excluding . In a one-electron description the electric dipole operator also requires for the active electron. These latter statements can fail as exact labels when atomic terms mix; the general parity and total-angular momentum restrictions still apply. Unresolved hyperfine structure is neglected here.
For strontium, the closed core contributes . The Pauli exclusion principle makes the two equivalent electrons a singlet state. Different radial orbitals permit both a singlet state and a triplet state. Thus the complete lists of atomic fine-structure levels areThe electron configuration has odd parity; the other two electron configurations have even parity.
Measure each energy level by the wavenumber in . An upward electronic transition of vacuum wavelength adds . Treat the quoted wavelengths as vacuum values to their limited accuracy; air corrections would be smaller than the precision justified here.
The strong ground-state absorption must be at . The weak ground-state electronic transition at must reach : the electric-dipole selection rule on excludes the other two triplet state energy levels, while weak spin-orbit coupling admixes singlet state character and relaxes the approximate spin selection rule.
Assume the usual ordering , as expected from a positive effective spin-orbit coupling constant. The three nearby strong spectral lines then join these energy levels to a common energy level. Larger lower-state energy means a smaller gap and longer wavelength, so correspond to respectively. The remaining spectral line joins and . Assume the latter is the upper energy level. These assignments and orderings agree with independently tabulated strontium energy levels.[ref]
Energy Levels of Neutral Strontium by NIST. Entries for 5s², 5s5p and 5s6s; the tabulated energies use cm⁻¹ rather than m⁻¹
Starting with the two ground-state gaps, the construction isThe resulting energy levels, rounded to avoid suggesting spectroscopic precision, are
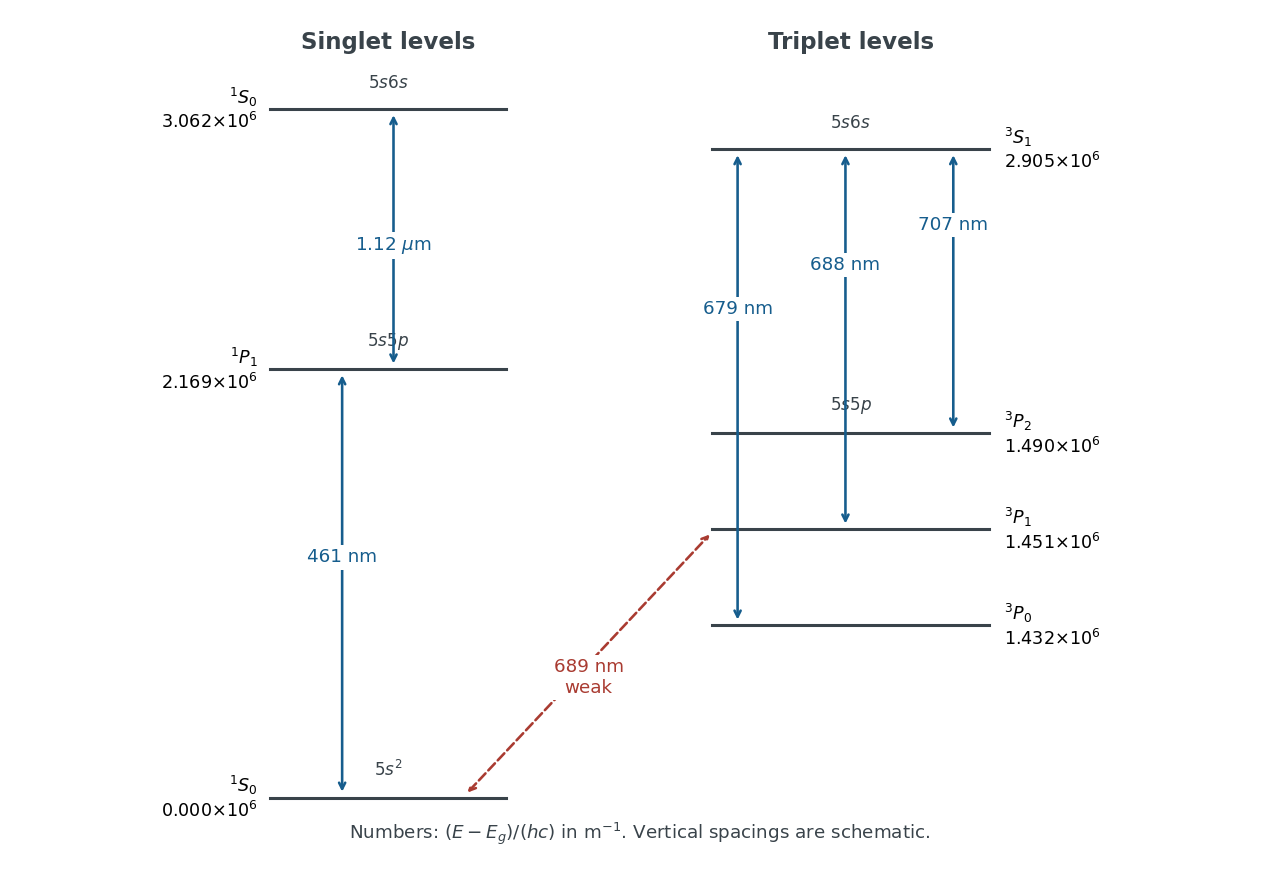
Both and exhibit the normal Zeeman effect. Each connects two singlet state energy levels, with Landé g-factor for the energy level and no first-order shift for . Their frequency shifts are . The weak spectral line also has three components, but gives shifts ; a three-component pattern alone does not establish the normal Zeeman effect. The spectral line likewise has three components with .
For , addition of angular momentum gives and . The even-parity atomic terms and atomic fine-structure levels areThe electric-dipole selection rules exclude all connections to the even-parity and electron configurations. To the odd-parity electron configuration, all seven allowed connections in pure LS coupling areHere a connection is listed independently of whether the energy level is above or below its partner.
In a weak magnetic field, use the Landé g-factorFor the three energy levels, this gives ; for it gives . The six-component spectral line is thereforeA component has frequency displacement . The electric dipole transition with has zero matrix element, since the relevant rank-one Clebsch-Gordan coefficients vanish there. The remaining allowed pairs giveThus all six frequency positions are distinct. Perpendicular to the magnetic field, both the and components are visible, with mutually perpendicular linear polarizations. Along the magnetic field the components disappear. This is the weak-field Zeeman effect, before the Paschen-Back effect changes the coupling scheme.
The closest components above differ by . At , their separation is about . A visible spectral line near would consequently require spectral resolving power of order . An ordinary low-resolution spectrometer would merge the components.
A suitable arrangement places an excited low-pressure strontium discharge or a transversely observed atomic beam between wide electromagnet pole pieces. Collect the emission perpendicular to the magnetic field, select the desired spectral line with a prefilter or monochromator, and send it through a high-finesse scanning Fabry–Pérot interferometer to a detector. A rotatable linear polarizer separates the two components from the four components. Calibrate both the interferometer's free spectral range and the local magnetic field; take zero-field and several field-strength scans, and fit component positions against .
The magnetic field should be uniform across the viewed volume, with its associated Zeeman splitting larger than the combined natural linewidth, Doppler broadening, pressure broadening and instrumental width, but much smaller than the separations of the relevant atomic fine-structure levels. A collimated transverse atomic beam, low collision rate and restricted collection angle reduce unwanted spectral line broadening. Avoid appreciable power broadening, resolve or model isotope and hyperfine structure, and check for self-absorption. A zero-field scan and the predicted linear scaling help distinguish actual Zeeman splitting components from neighbouring spectral lines.
Let the dimensionful angular momentum operators obey and take the interaction Hamiltonian as . Sincethe energy eigenvalues at fixed arewhere . Subtraction gives the Landé interval ruleMany atomic conventions use dimensionless angular momentum operators and call the energy-valued constant itself ; then the right-hand side is . Each energy level has magnetic sublevels with the same energy in zero field.
For fine structure, the effective interaction is with dimensionless angular momentum operators. It originates in the spin-orbit interaction, including the magnetic interaction of electron spin with orbital motion and its relativistic correction. Within an isolated LS coupling atomic term, the labels are the electron configuration, , and the interval between adjacent energy levels is . This is smaller than the electrostatic separation of atomic terms.
For magnetic-dipole hyperfine structure, the nuclear magnetic dipole moment interacts with the magnetic field generated by the electrons. Contributions include the Fermi contact interaction of penetrating electrons, orbital magnetic fields, and the dipolar field of electron spin. Projected onto one electronic energy level, their rotationally invariant magnetic-dipole part has the form . The labels are , together with the approximate electronic labels such as . This splitting is generally smaller than fine structure, reflecting the much smaller nuclear magnetic dipole moment. An electric nuclear quadrupole moment can add a different interaction and violate the simple Landé interval rule.
At the larger electrostatic energy scale, an effective two-spin exchange interaction can also be written within fixed orbitals. Its origin is the Coulomb interaction combined with the Pauli exclusion principle, rather than a large direct magnetic interaction. It separates singlet state and triplet state atomic terms; before spin-orbit coupling, useful labels include the electron configuration, . General electrostatic atomic term separations need not obey a single Landé interval rule as varies.
The shell carries a large spin (its ground atomic term under Hund's rules has ), and the electron penetrates the inner shells. Its nonzero probability density at the atomic nucleus supplies a potentially large Fermi contact interaction with the nuclear magnetic dipole moment. The shell here is singly occupied, so the obligatory paired- cancellation is absent. Both stable europium isotopes have nonzero nuclear spin, as the spectrum below also establishes.
The term symbol specifies , , . Its large , combined with appreciable nuclear spin, permits a wide range of total angular momentum and hence a substantial total hyperfine structure span. For , and the span is . The penetrating contribution and the available range of explain the large hyperfine structure; the term symbol alone does not determine its constant, which also depends on electronic recoupling, radial wavefunctions and configuration interaction.
Write , so the Landé interval rule for hyperfine structure intervals in ordinary frequency is . For the first four selected positions, the successive positive differences areTheir ratios are approximately , sinceThe decreasing frequencies therefore correspond to increasing , with , if increasing printed position means increasing physical frequency. Continuing the intervals predicts additional positions near and then . They identify peaks and unambiguously. The full assignment for isWith , these endpoints requireEquivalently, a complete set of six hyperfine levels gives when . This inference assumes that every upper hyperfine level contributes an observed position and the negligible lower hyperfine structure lets coincident transitions be grouped by their upper energy. A line count without these completeness assumptions would not alone fix nuclear spin.
The supplied positions obey the Landé interval rule only approximately. For example, , whereas . Even allowing each printed position a rounding error of does not fully reconcile these two relations. A least-squares dipole-only fit has a largest position residual of about . Small additional hyperfine interactions, measurement uncertainty beyond the displayed precision, or an error in the tabulated data could account for this; it is inappropriate to claim an exact identity. The grouping and nuclear spin are nevertheless robust. Reversing the convention for the printed frequency axis reverses the inferred signs of both constants, without changing the following ratio.
The remaining six positions give the other isotope's sequenceThe adjacent differences divided by are , respectively. Thus and the same endpoint argument gives .
For the same electronic energy level, the magnetic-dipole hyperfine structure constant is proportional to , to leading order in the small isotope-dependent change of the electronic wavefunction. Since the two nuclear spins are equal,Using the full spans averages over the approximate Landé interval rule: , soThe constants have the same sign, so the nuclear magnetic dipole moments have the same relative sign. Their absolute sign cannot be obtained from their ratio without the electronic field's sign. Finite nuclear-size corrections to the hyperfine interaction, often called the hyperfine anomaly, and electric-quadrupole corrections are neglected at this accuracy.
Choose the relative phases of the two energy eigenstates so that is real and nonnegative. Define the resonant Rabi angular frequency , the Rabi detuning , and . Substitution into the Schrödinger equation, followed by projection onto the two eigenstates, gives the exact interaction picture equationsKeep both probability amplitudes dynamical: replacing by one would incorrectly impose weak excitation and miss complete population transfer.
For , apply the rotating-wave approximation to discard the rapidly oscillating terms at . Differentiating the second remaining equation and using the first givesThe initial conditions are and . With , their solution isConsequently,This result permits order-one excitation, including full population transfer on resonance, even though the drive is small compared with the optical angular frequency.
For an average from switch-on to time , integrate the squared sine:On resonance this limiting average is , although the instantaneous probability repeatedly reaches one. The zero-coupling, zero-Rabi detuning case is understood separately as .
Near resonance alone does not make this an exact result for arbitrarily strong driving. The stated sinusoidal Hamiltonian also needs the rotating-wave approximation. A concrete counterexample is and : the exact equations give at switch-on, while the claimed rotating-wave sinusoid gives . Thus “without assuming weak radiation” can mean without assuming small excitation, but cannot remove the drive-to-carrier scale separation. The infinite-time average above belongs to the rotating-wave model; omitted corrections can accumulate over sufficiently long times.
Rabi oscillations require a coherent drive with a stable phase over several population cycles and an isolated, appreciably coupled pair of quantum states. For large-amplitude oscillations, the Rabi detuning must be comparable to or smaller than the Rabi angular frequency. The population lifetime, optical coherence time, transit time and drive coherence time should all exceed by enough to resolve the cycles. An ensemble also needs sufficiently uniform Rabi frequencies and Rabi detunings that phase averaging does not wash the oscillations out. The usual sinusoidal formula additionally assumes the rotating-wave approximation.
The Einstein coefficients describe incoherent transition rates, rather than the evolving optical probability amplitudes. Their simplest use is appropriate for broadband, randomly phased radiation, with field correlation time short relative to population evolution, or for rapidly dephased chemical atoms whose optical coherences can be eliminated. The radiation energy density per unit frequency must be smooth over the relevant spectral line. Stimulated upward and downward transitions and spontaneous emission then enter rate equations, without predicting coherent population return. A short observation time can give a small excitation probability even for a coherent field; that condition alone does not justify an Einstein coefficients rate description. Coherent laser irradiation can also admit steady rate equations after optical coherence has relaxed, with the appropriate saturation response.
Let the upper-state population lifetime be , and take the lower energy level to be stable. Spontaneous emission gives survival probability , hence a field or excited-state probability amplitude proportional to for . Its Fourier transform at angular frequency is proportional toThe squared modulus therefore has the Lorentzian line shapeHere is normalized over the full mathematical frequency axis. The full width at half maximum in ordinary frequency is . If both energy levels decay, their population decay rates add in the natural linewidth; pure dephasing adds to the amplitude-decay rate instead.
Impact-regime pressure broadening provides another example. Independent phase-randomizing collisions with rate give optical correlation proportional to , whose Fourier transform is a Lorentzian line shape of full width at half maximum . This assumes short, uncorrelated impacts: a general static distribution of perturbing fields need not give a Lorentzian line shape.
Use first-order time-dependent perturbation theory, setting in the equation for and retaining the generally complex matrix element . With and , the probability amplitude isFor , split the integral at the pulse maximum. For either or ,Thus the full first-order answer, without discarding the counter-rotating contribution, isAt long times it givesThis full expression is not exactly one squared Lorentzian line shape. To obtain the intended near-resonance spectral line profile, additionally assume , so the pulse has a narrow positive-frequency lobe and is negligible. ThenThe weak-excitation condition near resonance is . The squared Lorentzian line shape occurs because the symmetric exponential pulse has a Lorentzian line shape amplitude Fourier transform, which is then squared to obtain probability.
At half the maximum, , soThe full width at half maximum refers to varying the drive frequency through the positive-frequency resonance. For a concrete failure of the unqualified squared-Lorentzian line shape claim, set while keeping . The full first-order resonant probability is times the one-lobe prediction. Weak radiation alone therefore does not justify dropping the second Fourier transform lobe.
Define and let be the stimulated-emission cross section, averaged over upper magnetic sublevels. The Einstein coefficients relation gives the absorption cross section . Assume equally populated magnetic sublevels within each energy level, a steady pump, and a monochromatic beam. The photon flux is ; define and the effective population inversion . Then the rate equations areThe term includes both stimulated emission and absorption. In steady state, adding the two rate equations gives . The first equation then yieldsThe directed beam gains energy at rate per volume, giving and the optical gain coefficient . At line centre, write . ThereforeHere is the small-signal optical gain coefficient and is the saturation intensity, at which the local optical gain is half its small-signal value. Positive optical gain requires : the lower energy level must empty sufficiently quickly.
The cancellation of from is specific to this model's steady fixed pump and the fact that every upper spontaneous emission feeds the lower energy level. Adding an extra lower-state accumulation factor to would contradict these rate equations. Alternative conventions using the absorption cross section must replace by throughout. Homogeneous spectral broadening permits the whole selected ensemble to share this local saturation response.
In a uniform steady amplifier, with no extra propagation losses, the local optical gain law givesIntegrate between the two faces of length to obtainFor , this reduces to exponential amplification. In deep saturation, the intensity increment approaches : finite pumping limits the extracted power.
Let and . Applying the integrated amplifier law to each section givesEliminating givesAs a consistency check, both amplifiers have . The second section receives a larger intensity and adds , compared with in the first, while its multiplicative optical gain is much smaller. This is the expected approach to constant extracted power in saturation.
For a closed absorbing two-level system, write , let , and use for the absorption cross section. This differs from the stimulated-emission cross section convention in part (a). With , the rate equations areAt steady state, for . Hence the positive absorption coefficient isFor the given sodium fine-structure transition, the statistical weights are , , so and . The small-signal transmission gives the optical depthTo account for the finite decrease of intensity through the sample, integrate rather than treating the stronger beam as exactly uniform. If , this givesTaking as the incident intensity and ,Using the photon energy then givesIgnoring the small variation of the stronger intensity would instead give about ; the integrated value uses the stated transmissions more consistently. The length is not needed to eliminate , but gives the check .
The resonant absorption cross section is of optical-wavelength-squared size, much larger than a typical atomic geometric collision area. Pressure broadening cross sections involve phase-changing interatomic encounters and need not equal hard-sphere collision areas; their numerical value depends on collision partners and speed. No precise ratio follows from the supplied data. A useful distinction is that resonant absorption involves coherent response on the optical scale, so its cross section need not resemble an chemical atom's geometric area.
For homogeneous spectral broadening, all participating chemical atoms must share the same resonance and single-atom response width. In a dilute sample dominated by natural linewidth, the weak-beam requirement isUsing the sodium atomic mass, equality occurs near , so is a sufficient thermal condition. Spatial Zeeman energy shifts, light shifts and transit effects should also be small, the laser should address the same selected component, and the interaction should last long enough for steady populations. Uniform impact pressure broadening can also be homogeneous, but would change the width and effective cross section; it is not included in the lifetime-only numerical model.
Real sodium also has hyperfine structure and optical pumping among magnetic sublevels. The inferred is the effective cross section of the stated two-level system model, assuming statistical sublevel populations and no loss into unaddressed states. A measurement on a resolved, polarized closed hyperfine structure component requires its actual transition dipole moment and selection rules; a universal polarization-independent sodium cross section cannot be inferred from this simplified model. The dependence of resonant cross sections and saturation intensities on sublevel preparation is tabulated explicitly for the real sodium D lines.[ref]
Sodium D Line Data by Daniel A. Steck. Section on saturation intensity and resonant scattering cross section, and Table 7
The source bundle filename differs from the examination year; the cover identifies Trinity Term 2021. The solutions below retain the printed conventions where consistent and explain the corrections needed elsewhere.
The printed magnetic dipole moment uses dimensionless angular momentum operators: their physical counterparts are and . Choose the magnetic field along . Then the Zeeman energy is , whose characteristic size is . In LS coupling, treating this as a time-independent perturbation theory requires it to be much smaller than the relevant fine structure separation, rather than merely smaller than the Coulomb interaction energy.
For a concrete hydrogen atom estimate take the pair. The hydrogen fine-structure correction iswhere is the fine-structure constant, distinguished from the later potential coefficient. ThusConsequently a suitable weak-magnetic field condition for this pair isThere is no single threshold for every hydrogen energy level: the relevant fine structure separations decrease with . The energy level has no same- spin-orbit interaction partner, so it cannot itself define this comparison. This calculation neglects hyperfine structure; resolving that smaller scale requires an additional weak-magnetic field condition.
Within a fixed angular momentum multiplet, the component of along follows from . The corresponding result for follows by interchanging and . For the quantum state ,These are dimensionless expectation values in the convention of part (a). Substituting into the first-order time-independent perturbation theory formula givesFor , the Landé g-factor becomesThe magnetic sublevels must first be diagonalized within a degenerate multiplet; choosing along the magnetic field already does this. For the displayed fraction is undefined, but all vector expectation values vanish and there is no first-order Zeeman splitting. A higher-order shift may remain.
Write and retain the nuclear Coulomb potential. In spherical coordinates, the total potential energy iswhen ; is a Legendre polynomial. Treat the added term using quadratic anisotropic perturbation of hydrogen and degenerate perturbation theory in the hydrogen energy eigenbasis. It preserves parity and , so it cannot mix with , or distinct magnetic sublevels.
For a hydrogen atom, the hydrogen radial second moment givesThe angular expectation values of are for an orbital, for , and for , obtained by integrating the squared spherical harmonics. Therefore the five lowest spatial quantum states, counting statistical weight, have first-order energy levelsEach spatial quantum state has two electron spin states when spin-orbit coupling and the magnetic field are omitted. Thus these are five spatial states but only three distinct first-order energies; five distinct energies would be an incorrect reading. The accidental equality can be lifted at higher order. Quantitatively the added shifts must also be small compared with neighboring principal-shell gaps; is an asymptotic condition, not a guarantee for arbitrary large .
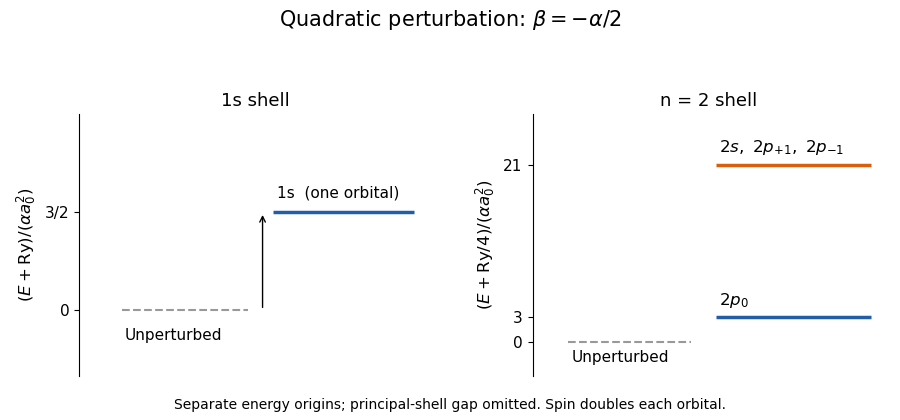
First-order spatial levels of locally perturbed hydrogen; the gap between principal shells is compressed
. Details. For general , the expectation value isIt does not cancel every order. At , couples to orbitals. For example its angular matrix element with is , and the radial matrix element is nonzero. The second-order nondegenerate perturbation theory sum has negative denominators, so this is a concrete nonzero higher-order contribution.
There is also a global limitation: for either negative above, . A localized wavepacket translated far along has fixed kinetic energy but arbitrarily negative potential energy, proving there is no true lowest energy level. The results describe local perturbative resonance poles, or a potential completed to a confining form far from the atomic nucleus. No globally exact ground-state cancellation exists for the potential as printed.
Neglect hyperfine structure, spin-orbit coupling and the diamagnetic term, and set , . The Zeeman energy is . The electron spin is free, so every spatial result below is duplicated with an added .
For a magnetic field parallel to , the old magnetic quantum number remains good and the first-order energy levels areThe two orbitals have zero orbital angular momentum, so only their electron spin contributes at this order.
For a magnetic field parallel to , the expectation of in each individual orbital vanishes. One must still check the degenerate subspace: its projected is zero as well. Thus there is no first-order orbital splitting in a sufficiently weak transverse magnetic field. Spin still contributes .
The transverse orbital Zeeman mixing can be seen without assuming its size. Put , , and . In the real orbital basis, is unaffected while have off-diagonal matrix elements of modulus . Diagonalizing this two-dimensional Hamiltonian gives the spatial energy eigenvaluesWhen , their shifts relative to the earlier levels are , with added for electron spin. The displayed square-root expression also handles comparable to , provided coupling to other principal shells stays negligible. The orbital retains zero orbital correction.
These quadratic orbital estimates keep the enhanced contribution from mixing the split levels. Minimal magnetic minimal coupling also produces ; its expectation value is positive and must be added for a full order- calculation. For example it gives for . It is parametrically smaller than the mixing term when .
Expand the wavefunction in unperturbed energy eigenstates,The are interaction picture probability amplitudes, so are level populations and in this closed two-level system. A classical field supplies the electric-dipole interaction . Choosing the relative phase of the energy eigenstates makes real. This is the resonant Rabi angular frequency.
Combining the field's two exponentials with the unperturbed quantum phases produces terms oscillating at and at . The rotating-wave approximation drops the latter, leaving exactly the supplied equations with coupling . It requires and a field envelope slow enough on the carrier timescale. Spontaneous emission and dephasing must be negligible during the sequence. For a ground-state hyperfine level transition, the analogous direct drive is a magnetic dipole transition; the same equations hold with the magnetic matrix element replacing the electric one.
Assume an initially unexcited chemical atom, two nonoverlapping square pulses starting at , and a common coherent field phase. In the weak-excitation limit , substitute into the second probability amplitude equation. After the second pulse,where the sinc function means , with value one at zero. Squaring givesThe printed prefactor is a factor of four too small for the supplied equations and pulse durations. A direct counterexample is : the exact resonant Rabi oscillation gives for two pulses of duration , because the intervening resonant rotating-frame evolution is trivial. Its small-area limit is , rather than . The latter is the single-pulse result before coherent addition.
The condition printed in the source also does not justify the derivation from a physical carrier field. A consistent rotating-wave approximation pulse regime isIf the supplied equations are treated as the mathematical model regardless, their weak-excitation integral remains valid for any ; subcycle pulses of the original real field require the discarded counter-rotating term and depend on carrier phase. here is the separation of pulse starts, as specified, rather than the dark interval .
The ground electron configuration of caesium has a single electron, with . Addition of angular momentum giving fixes the nuclear spin to . With dimensionless angular momentum operators, write the Fermi contact interaction asThe two hyperfine levels therefore differ by . In the convention and ,This exposes a missing ingredient in a prediction from the supplied data: and alone do not specify the valence electron contact density. Screening effect and penetration of the closed core strongly affect it. If instead denotes the total nuclear moment in units of , it must first be divided by in the displayed convention.
An explicitly crude unscreened hydrogen-like ion model uses and gives , or . It describes neither the shielding nor the radial function of neutral caesium. Using a pure unit-charge hydrogen orbital instead gives only , illustrating why a unique useful estimate cannot follow from an unspecified effective charge.
For physical neutral , the independently known zero-magnetic field clock transition isThis measured frequency, used to define the second, is supplementary physical information, not a consequence of and the rounded alone.[ref]
SI base unit: second (s) by Bureau International des Poids et Mesures. Definition: the unperturbed ground-state hyperfine transition frequency of caesium-133 is exactly 9 192 631 770 Hz
The direct electronic transition is in the microwave range and is a magnetic dipole transition, rather than a direct optical electric dipole transition. An optical implementation needs a two-field Raman transition.Using the corrected weak-pulse Ramsey spectrum of part (b), the peak at has height . The quantum interference fringes have spacing in angular frequency, with zeros at , and the sinc function envelope has its first zeros at . For , the central peak has approximately full width at half maximum in angular frequency. The clock transition between the two states has no first-order Zeeman energy, although a quadratic one remains.
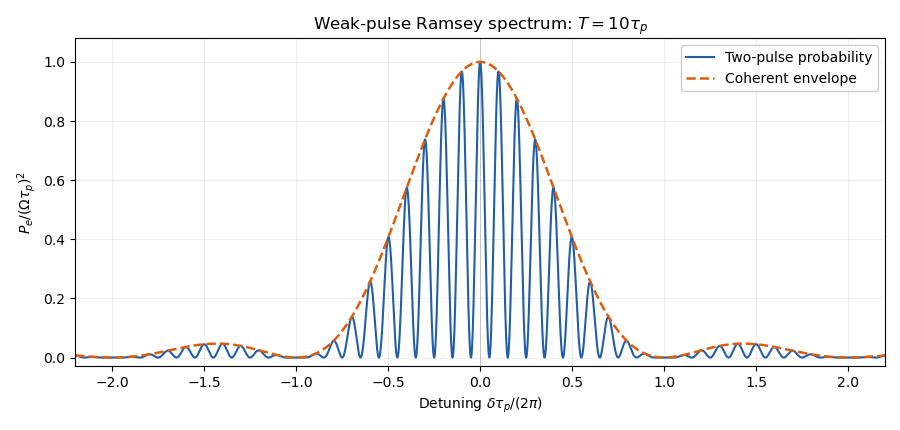


The first-order excitation amplitude is the Fourier transform of the temporal pulse envelope evaluated at the Rabi detuning. Two rectangular time windows therefore give exactly the mathematical structure of double-slit interference: each window gives a sinc function amplitude, and their separation gives a relative quantum phase . The transition probability is a sinc function squared envelope multiplied by .
Pulse duration corresponds to slit width, pulse-start separation to slit-center separation, and Rabi detuning to the transverse wavevector sampled on the screen. The narrow fringes measure the long separation, while the broad envelope measures the short interaction duration. As in spatial interference, adding probabilities instead of coherent probability amplitudes would lose the fringes. Randomizing the relative pulse phase averages the cosine squared to .
Differentiating the specified rotating-frame amplitudes gives the time-independent Schrödinger equationThe characteristic polynomial is , henceFor , corresponding normalized eigenvectors are proportional to . Their arbitrary complex coefficients evolve as ; this solves the amplitude equations for any initial quantum state.
Restoring the common energy origin , the two dressed state energy eigenvalues areThus their positive energy separation increases byEquivalently, the branches connected to and shift by and respectively. Their signed difference changes by , which is the printed expression with a signed convention. For negative Rabi detuning, an actual positive separation still increases; for example at it cannot have the negative change implied by reading the printed expression as an unsigned gap. These opposite AC Stark shifts describe level repulsion.
Specify the radiation normalization first. Let be radiation energy density per ordinary frequency, with SI units . Then is the spontaneous emission rate per upper-state chemical atom, while and are the induced absorption and stimulated emission rates, evaluated at a sharp transition frequency. For a finite spectral line profile, replace by its profile-weighted average.
The Einstein coefficients obeyHere are the statistical weights of the two energy levels, and the field is unpolarized and isotropic. The units and physical dimensions areThe apparently time-independent dimension of follows because a per-hertz density includes an extra unit of time. If instead multiplies the mean intensity , use and . These distinct normalizations must not be mixed in the radiative transfer equation.
Use the energy-density convention of part (a), a normalized common spectral line profile , and isotropic spontaneous emission. For a narrow spectral line centered at ,The factor follows from : angle-integrated absorption removes of energy per volume and time. The emission coefficient is per solid angle, whereas the absorption coefficient has units of inverse length.
The Boltzmann distribution in thermal equilibrium gives . Dividing the two expressions and using the relations between Einstein coefficients,Thus the source function equals the Planck function at the transition frequency, and in the usual narrow-line approximationWith an exactly common finite-width and constant line-center Einstein coefficients, their ratio is constant across the profile; exact equality to a varying at every frequency is only a narrow-line approximation. Exact thermal radiative transfer requires frequency-resolved detailed balance, which supplies the corresponding small profile corrections. This distinction does not alter the intended equilibrium result.
For constant nonzero absorption coefficient, multiply the radiative transfer equation by the integrating factor :Integrating from the entrance to givesThe boundary condition is immediate. Differentiation gives , checking the equation. For positive optical depth , the incident specific intensity is forgotten and . Negative absorption coefficient instead describes gain and does not have this relaxation limit. At use the nonsingular form directly: .
In local thermodynamic equilibrium, write the one-ray radiative transfer equation asLarge total optical depth removes the boundary contribution, but an additional local condition is essential: the temperature and opacity vary little over an absorption length . Iterating once in that small gradient givesThe integrated Planck function and its derivative areUsing the specified Rosseland mean opacity, with measured here in inverse length, the source's definition therefore givesThe printed leading term lacks a factor under that definition. An isothermal optically thick medium is a concrete counterexample: gives exactly, while the printed formula gives .
A physical radiative flux requires an angular integral. For direction cosine , the same gradient expansion gives . Integration over all directions gives the net radiative diffusion fluxThe isotropic equilibrium contribution cancels between opposite directions. For comparison the outward-hemisphere flux alone is at this order. None of these consistent definitions yields the printed mixture of coefficients.
To derive fermionic Einstein-coefficient relations, first consider an idealized emitter that can exchange one ultrarelativistic fermion with the bath, with zero chemical potential. Let be its density of states per volume and ordinary frequency, with populated helicity states; for one neutrino helicity , whereas the corresponding photon count is two. The Fermi-Dirac distribution givesPauli blocking changes the final-state factor in downward emission from the bosonic to . Define positive coefficients and . The upward rate per lower emitter is , and the downward rate per upper emitter isThermal detailed balance, with , yieldsFor example, substituting reduces the balance equation to because .
If one insists on the original additive notation , its formal coefficient is negative:A negative correction means inhibited emission into occupied modes, not a negative total decay rate; ensures positivity. The dimensions are those of part (a), and the factor of two relative to the photon density is a helicity count, independent of the statistical sign.
There is a physical qualification: ordinary electronic energy levels of the same chemical atom have the same fermion parity, and cannot interconvert by emission or absorption of a single neutrino. A realizable single-fermion emitter must change the relevant particle-number and parity sector, with matching conservation laws. For ordinary atomic levels, neutrino-pair processes would require a different calculation. A bath with nonzero chemical potential would also require the corresponding chemical-equilibrium population ratio. The calculation above is the intended single-mode statistical analogy under explicit assumptions.
The closed shells have , leaving the equivalent electrons. Addition of angular momentum and the Pauli exclusion principle permit the atomic terms . Their term symbols use . Hund's rules put the triplet state below the singlet states; for a less-than-half-filled shell the smallest in is lowest. ThusAll have even parity. These assignments agree with independent tabulated carbon energy levels.[ref]
Energy Levels of Neutral Carbon by National Institute of Standards and Technology. 2s²2p² entries: ³P₀,₁,₂ at 0, 16.41671 and 43.41350 cm⁻¹; ¹D₂ at 10192.66 cm⁻¹; ¹S₀ at 21648.02 cm⁻¹
The rounded source values should not be forced to satisfy an exact Landé interval rule; pure first-order LS coupling need not account for every measured fine structure correction.For two decays from a common upper energy level, their photon wavenumber difference equals the separation of the two lower energy levels. Treating both quoted wavelengths as vacuum values initially giveswhose difference is close to . The only matching lower pair among the supplied energy levels is , with the shorter-wavelength photon ending on . The two independent upper energy estimates areThusThe electric-dipole selection rules require odd parity, , and to connect to both singlet lower energy levels. The apparent discrepancy has a concrete convention explanation. The NIST carbon line table lists under vacuum wavelengths and under air wavelengths, reproducing the source's rounded numbers.[ref]
Strong Lines of Carbon by National Institute of Standards and Technology. Vacuum section: C I 1930.906 Å; air section: C I 2478.561 Å
The second number must therefore be converted to a vacuum wavelength before using . For the tabulated upper and lower energy levels, its vacuum wavelength is , corresponding to an air refractive index about . This removes the spurious discrepancy. The first calculation alone, which correctly uses a vacuum wavelength, already gives the intended upper energy level to the accuracy of the rounded data. The independent upper value is .[ref]Energy Levels of Neutral Carbon by National Institute of Standards and Technology. 2s²2p(²P°)3s ¹P°₁ entry at 61981.82 cm⁻¹
In the idealized isolated-carbon chemical atom interpretation, both beams carry approximately per incident particle, close to the threshold. However equal incident energies do not imply equal excitation channels.
A single optical photon cannot strongly excite any of the listed even-parity energy levels from the even-parity ground state through an electric dipole transition. In particular also violates and the restriction. The lowest strong opposite-parity excitations lie well above . Thus one expects no strong resonant absorption or corresponding strong atomic fluorescence from these ground-state chemical atoms at this optical wavelength; weak higher-order processes and scattering are separate effects.
Incident electrons can undergo inelastic scattering and collisional excitation. Their orbital angular momentum and exchange with bound electrons allow transfers excluded for a single electric dipole photon. The energy level at and the low fine structure excitations are energetically accessible. One therefore expects scattered electrons with discrete energy losses, possible weak forbidden spectral lines, and collisional or surface relaxation. For example has wavelength approximately ; spin mixing supplies the weak spin-forbidden radiative channel.
There is a numerical threshold caveat. Literally,Neither quoted beam reaches from the exact ground state if these are taken as exact monoenergetic values. The electron value rounded to two decimal places could represent an actual energy above threshold; if so, is collisionally accessible and its decay to produces the weak electric quadrupole transition at , followed by lower-level relaxation. These emissions require populations and sufficiently low collisional quenching of a metastable line.
An actual solid carbon foil has bands, collective excitations and surface losses, so it need not reproduce the isolated-chemical atom spectrum or selection rules. Its optical and electron-induced emission cannot be predicted quantitatively from the five atomic energy levels alone. This is the limitation of the foil wording in the source.
Take the initial ground-state chemical atom at rest in the laboratory. The incident photon has energy and momentum . The excited chemical atom has rest mass and must carry that momentum. Conservation of energy and conservation of momentum giveSquaring cancels the photon-energy square:ConsequentlyThe extra positive term is the photon absorption recoil correction and supplies atomic recoil energy. The formula is exact for the initially stationary isolated chemical atom; it does not assume . Initial and final chemical atoms cannot both remain at rest during single-photon absorption without another body taking momentum.
Put . Gauss's law for a uniform spherical nuclear charge distribution, with the electrostatic potential matched at , gives the electronic potential energySubtracting the point-nucleus Coulomb potential leaves the nonnegative perturbationThe normalized nonrelativistic hydrogen-like ion wavefunction has . Since , approximate it inside the atomic nucleus by . First-order time-independent perturbation theory then givesDefine , as the paper specifies for the hydrogen-like ion, rather than the neutral-hydrogen value. Using givesto leading order in . For , , , andThe finite nuclear charge radius makes the energy level less bound. The quoted frequency is a level shift divided by : a line frequency shifts by , so a decay into this shifted level alone has the opposite sign.
The formula is a nonrelativistic, small-nuclear charge radius estimate. Keeping the exponential in the first-order integral gives , about a one-percent correction here. More seriously, for lead, so relativistic contraction is substantial. A precise physical prediction needs a finite-nucleus Dirac equation calculation; the boxed estimate is the model requested, not precision heavy-ion spectroscopy.
A nonrelativistic wavefunction vanishes linearly at the origin. Its probability density is therefore proportional to , so its first-order finite nuclear size correction shift scales as , rather than the result for . It is positive but much smaller.
More explicitly, the normalized hydrogen-like ion radial function obeys near the origin. Since the spherical harmonic has unit angular norm,ThusThis is about in the nonrelativistic model. At large , the relativistic Dirac spinor has an -type small component and a much stronger nuclear sensitivity than this nonrelativistic argument predicts; remains much more suppressed. Neglecting spin-orbit interaction corrections would miss that distinction.
The finite nuclear size correction perturbation is unchanged for the same lead atomic nucleus. Its leading effect on one occupied orbital is still proportional to that orbital's contact density,Other electrons screen the nuclear attraction and alter the self-consistent wavefunction, usually reducing its contact density and hence its positive shift relative to the bare hydrogen-like ion. The reduction is modest for a deep orbital: it penetrates well inside most other shells and still samples nearly the full attraction. Replacing by the neutral atom's asymptotic charge one would be inappropriate for this core orbital.
A neutral lead atom has two electrons. Their contributions add in a frozen-orbital estimate of total atomic energy, roughly twice the per-electron shift, with relaxation and correlation corrections. An experimentally observed transition frequency again depends on the difference between the finite-size shifts of both participating many-electron quantum states. A quantitative neutral-lead result requires a screened relativistic many-electron calculation, not the one-electron formula with a single universally valid effective charge.
Articles by others on the same topic
There are currently no matching articles.