The source bundle's filename says 2020, but the paper's cover identifies Trinity Term 2021, C7, A11050W1. All optional answers are included. The unavailable figure in a is explicitly distinguished from deductions supported by the surviving text.
At the initial negative membrane potential, most voltage-gated potassium channels have a closed activation gate. Depolarization moves charged residues in each voltage-sensing domain, particularly the S4 helices, through the electric field. Coupling between the voltage-sensing domains and the pore opens the intracellular gate. A classical voltage-gated potassium channel has four subunits, each with six transmembrane helices; S5, the pore loop and S6 contribute to the central pathway and selectivity filter. The rising open probability produces increasing outward potassium electric current, since the imposed positive membrane potential is above the potassium Nernst equilibrium potential. Here outward electric current is positive: , where is channel number and is single-channel electrical conductance.
The electric current subsequently falls because the rapidly inactivating Shaker form exhibits ball-and-chain inactivation. An intracellular N-terminal domain, tethered to the channel, blocks the open pore. The ion channel can therefore stop conducting while the membrane potential remains depolarized and the activation machinery remains activated. This is ion-channel inactivation, rather than reversal of the voltage step, loss of the ionic driving force, or ordinary gate closure. Repolarization favors removal of the block and recovery of channel availability. Slower C-type inactivation involving the outer pore can coexist, but is not the principal explanation of the rapid decay described here.
Several independent observations connect this functional explanation to structure. Small transient gating electric currents report movement of sensor electric charge before ionic conduction; charge-neutralization studies of S4/S2 change the measured gating charge and activation. Structural studies resolve separate sensor and pore regions.[ref]
Voltage-sensing residues in the S2 and S4 segments of the Shaker K+ channel by S. A. Seoh and others. Gating-charge measurements identify voltage-sensing residues
[ref]Crystal structure of a mammalian voltage-dependent Shaker family K+ channel by Stephen B. Long and others. Voltage-sensor and pore architecture of Kv1.2
Historical N-terminal removal abolished rapid ion-channel inactivation while preserving activation, and the corresponding free peptide restored block from the cytoplasmic side. These results separate the activation gate from the tethered blocker and identify which side of the cell membrane contains the blocking domain.[ref]Biophysical and molecular mechanisms of Shaker potassium channel inactivation by Toshinori Hoshi and others. N-terminal perturbations distinguish rapid inactivation from activation
[ref]Restoration of inactivation in mutants of Shaker potassium channels by a peptide derived from ShB by William N. Zagotta and others. Cytoplasmic peptide restores inactivation to channels lacking the native N-terminal domain
Source-gap TODO: the supplied PDF's figure invokes an undefined image object, so its actual pulse duration, pulse sequence, current scale, peak amplitude, decay time and any tail-current waveform cannot be read. The account above uses only the stated change from to and the stated rise and rapid decay; no numerical kinetic fit or reconstruction of the missing trace is claimed.
For two-state ion-channel gating, write . Gain minus loss of open probability gives the ordinary differential equationThus the ratio of the transition rates determines the final open probability, while their sum determines the response speed. Both given channels have . For an initially closed population, with measured in milliseconds,The mean individual open and closed lifetimes are and : both are for A and for B. With the same channel number, single-channel electrical conductance and driving force, the steady macroscopic electric currents are equal, but B reaches that value ten times faster. If both populations were already at , neither would show a mean relaxation under these fixed rates: a response also requires an initial condition or a change in rates.
The stationary variance of the number open is for independent channels. Its autocovariance is , so B's fluctuations decorrelate ten times faster even though their instantaneous variance is unchanged. Faster gating allows an ion channel population to follow a brief stimulus and contribute to rapid action potential timing. Slower gating integrates over brief changes and can provide a delayed opposing electric current, useful in setting action potential duration or frequency selectivity. Those examples describe consequences of kinetics; the stipulated voltage-independent rates alone do not define a complete excitable neuron.
Add a nonconducting inactivated state to the continuous-time Markov chain:For example, and . Depolarization first increases , then transfers population into by ball-and-chain inactivation. A small recovery rate during the pulse permits a sustained decline of . Repolarization favors recovery, eventually restoring the closed but available population. Several sequential closed states can represent the cooperative movements of four voltage-sensing domains; three states are the minimal explanation of rise followed by decay, rather than a quantitative fit to the missing trace.
Loss of outward potassium electric current weakens repolarization and can broaden an action potential. Reduced residual potassium electrical conductance can reduce the subsequent hyperpolarization and permit faster repeated firing. Conversely, prolonged depolarization can maintain sodium ion-channel inactivation, delay recovery and lower the maximum firing frequency or cause depolarization block. The sign of the frequency change depends on the other ion channels and recovery kinetics; rapid potassium ion-channel inactivation alone does not fix it.
The passive upper limiting membrane potential for a purely sodium-selective pathway is its Nernst equilibrium potential. Equal inside and outside electrochemical potentials, using ideal concentrations, requireThis is the maximum absolute inside-minus-outside voltage under the stated passive assumptions. If the reference resting membrane potential is , the corresponding increase is about .
Actual action potentials usually peak below because voltage-gated sodium channels undergo ion-channel inactivation, while outward potassium and leak electric currents oppose inward sodium electric current. Finite membrane capacitance and finite available time also prevent instantaneous equilibration. At , the sodium driving force itself is zero.
Use the logic approximation for gene regulation, zero basal production and initially vanishing X, Y and Z. Let , , and assume the activated Q level exceeds the X activation threshold. The circuit combines a coherent feed-forward loop with negative autoregulation. While X is below its self-repression threshold it obeys , hence X first crosses atY production begins then. While X remains above , . The AND gate for Z first becomes active when Y reaches , soThe approximation drops the short X build-up time because . Strictly the input must last longer than this onset time to produce a nonzero amount of Z. This requires : at equality the threshold is approached only asymptotically, and below it Z never switches on. No numerical delay follows without the threshold-to-production ratio.
In the strong negative autoregulation limit, X is held close to ; a smooth repression law regularizes the rapid switching of the ideal Boolean rule. After the input disappears at , production of X stops and . It takes for X to fall below . Until then Y remains at and the Z AND gate stays on, so Z stays at . After that Z decays by dilution, taking another to halve. ThereforeMore generally replace the first by . This distinguishes disappearance of the external ligand from disappearance of the persistent X transcription factor.
Put , and . Saturated CheR supplies a constant receptor methylation flux, while CheB removes it with Michaelis-Menten kinetics:At a finite steady state, , givingIt requires and below the total available receptor pool. The steady state is stable because . If , the ideal equation has no finite positive steady state; actual methylatable sites would become exhausted.
Attractant binding reduces receptor signalling, reducing CheA-dependent phosphorylation of CheY and initially reducing tumbling frequency, so smooth runs become longer. A spatially uniform stimulus provides no preferred direction. Perfect adaptation means that after this transient the tumbling frequency returns exactly to its pre-step value despite continued attractant presence.
The equation above alone does not specify attractant occupancy or its influence on activity. If binding simply changes activity from to with , without changing either modifying flux, remains and the output remains : there is no adaptation. Thus the usual fall followed by recovery requires the additional feedback mechanism developed in c or d, rather than following from the stated Michaelis-Menten kinetics alone.
Let attractant reduce signalling activity per methylated receptor by a factor , so the output is . Before stimulation take and . Perfect adaptation requires the final methylated receptor concentration . With , the new steady state condition isEquivalently . Since , this reduces demethylation relative to the initial condition. Immediately after binding, activity and tumbling fall. The excess CheR flux then increases receptor methylation, compensating the activity loss until . The methylated population must have sufficient remaining capacity to reach .
Increasing CheR concentration raises and the baseline activity; increasing CheB concentration raises and lowers it. If the attractant-dependent ratio remains fixed after such a change, the new pre- and post-step outputs areTheir equality requires . Thus a ratio tuned for one generally loses perfect adaptation when either enzyme level changes independently. For a concrete example, initially and require ; changing to requires , so keeping fails. Proportionally scaling both enzyme levels preserves the ratio and the adaptation condition, while changing response speed. This is fine tuning, rather than robust perfect adaptation.
In the simplified Barkai-Leibler model, let be the total methylated receptor concentration and the active, unoccupied part. Ligand-bound methylated receptors contribute zero activity. Because CheB acts only on active receptors,At a finite steady state,The output is independent of attractant concentration : attractant changes the necessary receptor methylation level, while the flux balance fixes activity. This proves perfect adaptation for , and enough methylatable receptors. At fixed , , so this steady state is locally stable. If every receptor is permanently bound, , or if the required exceeds the available pool, compensation fails.
This feedback couples demethylation directly to the output. Lower activity lowers demethylation; continuing CheR action restores it. The structural dependence on active receptors removes the separate adjustment of required in c.[ref]
Robustness in simple biochemical networks by Naama Barkai and Stanislas Leibler. Figure 1b and equation 1: saturated modifying flux and output-dependent reverse modification
Changing CheR or CheB concentration changes and hence the baseline tumbling frequency, but the output still returns to that new baseline after an attractant step within the operating range. Raising CheR raises ; raising CheB lowers it. Adaptation time also changes: linearization gives . Thus robustness of the return property does not mean an invariant tumbling frequency or response time across different enzyme abundances.
If CheR instead has Michaelis-Menten kinetics on the unmethylated pool , its flux is . ThenThe left side depends on ligand through , even at fixed output ; consequently the equilibrium output generally depends on attractant and perfect adaptation is lost. It is recovered approximately when throughout the response, so the CheR flux is effectively saturated.
Dense DNA packaging opposes three main physical effects. The negatively charged DNA backbones repel through electrostatic interactions, moderated by electrostatic screening and counterion condensation. At close approach, hydration forces and excluded volume become important. Bending a semiflexible DNA polymer on a radius comparable with its persistence length stores elastic energy. Confinement also removes configurational entropy, and concentrating mobile counterions can impose an additional osmotic pressure and free energy cost. Their relative importance depends on geometry and ionic conditions.
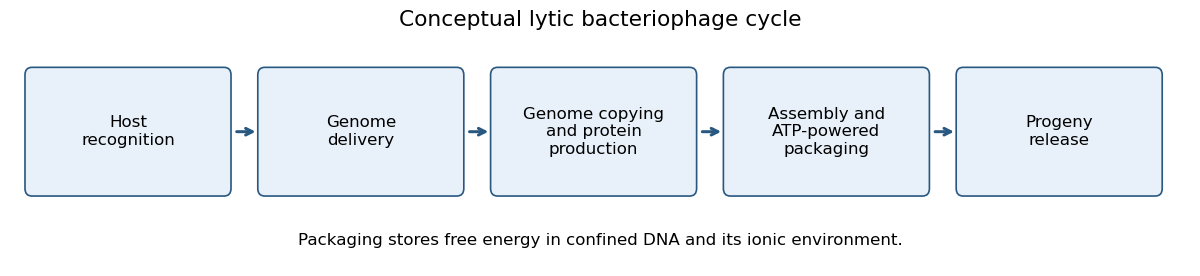
Conceptual lytic bacteriophage cycle and active DNA packaging
. Details. In many double-stranded DNA bacteriophages, a portal-associated molecular motor uses ATP hydrolysis to move DNA into a preformed capsid. Chemical free energy pays for bending, repulsion and confinement, so the opposing force rises as filling proceeds. This is active packaging rather than spontaneous equilibrium compaction. Optical tweezers can infer that opposing force from displacement of a trapped bead and monitor packaging through the changing external DNA extension. Such measurements establish the mechanical principle without requiring any virus-handling procedure. The inverse spool model used below is an idealized packing geometry, not an exact structural claim for every bacteriophage.
For an inextensible worm-like chain, the DNA bending energy iswhere is curvature and is polymer contour length. A circle of radius has and length . Its elastic energy is therefore . Adding the contributions of rings at each radius givesThis ignores short connecting segments, twist, interactions between neighboring DNA segments and end effects. In a cylindrical stack with axial spacing , away from the ends.
Correction of the printed limits: ordinary positive polymer contour length requires integration from to , because and the ring count is positive. For example, taking , and makes the printed outward-to-inward integral negative. Thus its sign cannot be retained as a physical length.
For hexagonally packed strands choose axial ring spacing and radial layer spacing . With and radial layer density ,The same continuum replacement in b givesEliminating the inner radius yieldsThe resisting packaging force is the marginal free energy per added polymer contour length, at fixed , and :The force opposing insertion is positive; its contribution to outward ejection has the same magnitude. The domain is . The divergence as the inner hole closes reflects the singular curvature of ideal rings and marks breakdown of the continuum model. If spacing also relaxes, the full free energy must include interstrand interactions before optimization; simply varying in this bending-only formula is insufficient.
The small binding RNA can serve conceptually as an RNA aptamer recognition probe. Coupling recognition to an optical label would make a specifically bound virus appear as a moving fluorescent spot. Single-particle tracking estimates successive spot positions and fits their displacement statistics. Specificity is assessed by comparison with a nonbinding recognition probe or a sample without the target, while the much faster diffusion of unbound probes distinguishes freely moving label from target-associated signal. Subnanomolar dissociation constant supports appreciable occupancy at low probe abundance, but affinity alone does not determine the dissociation lifetime because . This describes detection principles, not an operational virus preparation or labelling protocol.
For a dilute spherical particle of radius in bulk water, the Stokes–Einstein relation gives, at an explicitly assumed ,A free probe would have , roughly thirty times larger. The recognition probe changes the target's hydrodynamic size only modestly in this estimate.
For isotropic Brownian motion, the mean vector displacement is zero. In , distinct measures of how far it moves areA two-dimensional image instead gives projected root-mean-square displacement and mean projected distance . The projected mean squared displacement is ; independent localization errors of variance in each coordinate add . Finite exposure blurs motion, and nearby walls reduce diffusion; those effects must be separated from particle size when interpreting a trajectory.
The named bacteriophage, Bacillus virus phi29, infects Bacillus subtilis; the paper's identification of Escherichia coli as its host is incorrect. This correction does not alter the idealized geometry or numerical inputs used below.[ref]
The stipulated DNA volume is and the spherical capsid volume is . ThusThe packed DNA occupies a substantial fraction of the available space: the remaining nominal fraction is . Compared with the paper's stated reference, it reaches about of that value. This indicates dense confinement, but does not prove crystalline order or specify interstrand spacing. DNA is a connected elongated polymer, rather than a collection of independent hard spheres.
There is also a numerical qualification: the exact densest equal-sphere packing fraction is , not . Using it changes the comparison to about without changing the calculated volume fraction or its physical interpretation.
The polymer contour length and magnitude of total electric charge areUse a uniform charged-sphere model for the packed DNA, a homogeneous dielectric with , and initially neglect electrostatic screening. Gauss's law gives inside the sphere and outside. Integrating the electrostatic energy both inside and outside,For an illustrative aqueous relative permittivity and ,The question does not specify permittivity; the result scales as . Using vacuum instead would increase it eightyfold and would not describe an aqueous interior.
Strictly, the work of packaging is a difference of electrostatic energies, rather than the packed self-energy alone. An extended charged rod also has self-energy. With a microscopic cutoff of order the DNA radius, the thin-rod approximation isOne derivation integrates the pair Coulomb potential along the rod, replacing separation by : the exact regularized expression is . The diagram's transverse label suggests , giving . Hence this explicit reference-state estimate is . The cutoff and precise initial conformation are additional model assumptions, but their correction is small compared with the unscreened sphere value.
The bare packed-sphere estimate exceeds by approximately ; subtracting the estimated extended-rod self-energy still leaves a factor of about . The dominant missing effect is electrostatic screening: mobile positive counterions accompany the negatively charged DNA, greatly reducing long-range repulsion. Counterion condensation can further reduce its effective electric charge. Treating the genome's bare charge as uncompensated throughout and outside a sphere therefore grossly overestimates its long-range electrostatic energy.
An optical tweezers measurement integrates an imposed force over a displacement. A reversible extraction or packaging comparison concerns a free energy difference relative to the unconfined DNA and ionic reservoirs. It can include entropy, bending and hydration contributions; dissipative pulling can add further work. Extraction can also be assisted by stored internal free energy, so the sign of externally supplied work requires the experimental convention. Here the supplied positive value is compared as an energy magnitude.
Adding monovalent counterions makes the combined genome-plus-ion electric charge nominally zero. It does not make the local electric field or free energy vanish: opposite charges have spatial distributions, and confining mobile ions costs entropy.
A definite screened continuation of b replaces the bare Coulomb potential by , with and . In the Debye–Hückel approximation, a uniformly distributed fixed charge density satisfies inside, with zero fixed charge outside and matching potential and radial field at . Its interior solution isThe screened charging free energy is thereforeFor , the bracket approaches one: only charges within roughly a screening length contribute appreciably, yielding the local estimate . With the same as before, gives bracket andabout times smaller than the unscreened packed-sphere self-energy. This is a specified continuum estimate of packed-state charging free energy; a full packaging free energy also subtracts the screened unpacked reference and includes ion redistribution. It should not be presented as a unique exact packaging work fixed by the problem's few inputs.
The supplied counterion count also permits an independent estimate of the entropic contribution. For ideal confined ions, , so compression from an accessible reference volume to costsFor example, approximate the original screening atmosphere as an annulus of thickness around a rod. With , . Taking gives . This useful order-of-magnitude ion-entropy model is distinct from the screened charging model; the two estimates must not simply be added as independent terms without a consistent ionic free energy functional. The rod cutoff, fixed DNA volume and screening-atmosphere geometry are rough geometric assumptions, not a microscopic reconstruction of the source sketch.
There is a further consistency limit. The nominal ion number density is . For a uniform ideal monovalent screening gas, would give at , rather than ; added salt would shorten it further. Thus the stated ion inventory and screening length cannot both be interpreted as a self-consistent dilute uniform screening gas with this dielectric. Bound counterions, nonuniform distributions and an effective phenomenological screening length can change that interpretation. Linear screening is itself only an estimate near highly charged DNA.
The specified screened-sphere charging model gives , about times the supplied experimental magnitude. It is much closer than the unscreened result. The separate ideal counterion-compression estimate gives , within about of that magnitude, but this numerical proximity does not validate its rough geometry.
Remaining differences can arise from the unpacked reference self-energy, counterion condensation, ion exchange with the surrounding reservoir, nonuniform charge density, nonlinear electrostatic screening, finite DNA thickness and the actual capsid shape. DNA bending energy, hydration forces and configurational entropy also contribute. The experiment measures a process-dependent work or inferred free energy, whereas the simple screened sphere computes just one model contribution. Neither a zero-net-charge argument nor agreement of one approximate number establishes an exact total energy.
For ideal freely mobile counterions, their gross osmotic pressure isIf the DNA volume is excluded from the ion-accessible volume, use instead, giving . These are ideal-ion estimates; bound counterions contribute less like independent mobile particles, and the net outward osmotic pressure also subtracts the external osmotic contribution.
Stored free energy associated with this pressure, together with bending and interstrand repulsion, can favor initial genome release when a delivery pathway becomes available. This explains a possible mechanical benefit of dense packaging at a conceptual level. It does not imply that internal pressure necessarily supplies all the work of complete genome delivery into a host, whose own environment and interactions also matter.
A major source of ATP is chemiosmosis: an electron transport chain uses downhill electron transfer to establish a proton-motive force, and ATP synthase converts that electrochemical gradient into ATP formation. In aerobic oxidative phosphorylation, reduced carriers supply electrons, ultimately reducing oxygen to water; selected respiratory complexes couple the favorable oxidation-reduction reaction to uphill proton translocation. In a mitochondrion, this takes place across the inner membrane, creating a relatively positive, acidic exterior side and a relatively negative, alkaline matrix. Bacteria use their cell membrane. In photosynthetic systems, light-driven electron transfer can supply the gradient instead. Substrate-level phosphorylation is another ATP-producing route, so the description is not universal to every organism and condition.
The membrane region of ATP synthase contains a rotating ring of c-subunits next to subunit a. Two offset aqueous half-channels expose the ring's proton-binding sites to opposite membrane sides. Proton binding and release bias ring rotation down the electrochemical gradient. The central shaft transfers torque to the catalytic head, while a peripheral stator prevents the head from simply co-rotating. This is the ATP synthase rotary mechanism, rather than unrestricted proton leakage through a pore.
The three catalytic beta subunits cycle through different binding states as the shaft turns: ADP and inorganic phosphate bind, bound ATP is favored, then ATP is released. The binding-change mechanism uses mechanical work to alter binding affinities and product release as well as the chemical step. A full coupled revolution produces three ATP molecules; a ring with c-subunits transfers approximately protons per revolution in the tightly coupled model, or per ATP.[ref]
Structure of a bacterial ATP synthase by Hui Guo and others. Proton-translocation half-channels and rotor architecture
[ref]The overall cycle must have a favorable Gibbs free energy change. If the proton-motive force becomes too small, synthesis stops; under suitable conditions the same molecular motor can reverse, using ATP hydrolysis to pump protons uphill. A catalyst changes kinetics, not the thermodynamic condition for synthesis.
Use the supplied as a standard transformed Gibbs free energy of ATP hydrolysis, rather than replacing it with a different commonly quoted standard value. Approximate thermodynamic activities by concentrations divided by , at an explicitly assumed . The dimensionless reaction quotient isConsequentlyand the reverse synthesis cost is . Substituting bare millimolar numbers in this quotient without the standard state would give the wrong logarithm.
For outside-to-inside proton movement, the membrane potential is and , so . The molar electrochemical potential change isThus the available proton-motive force magnitude is , supplying per transported proton. Assuming one proton per c-subunit per revolution, three ATP per revolution and no leakage, slippage or transport overhead,The smallest integer permitting synthesis is therefore in this ideal budget. A ring of thirteen gives only a small driving margin; inefficiency or sustained finite output could require a larger ring. The result is an energetic lower bound and near-reversible estimate, not a unique structural determination from these data. Reasonable temperatures near – leave the minimum integer thirteen. Additional membrane transport costs have been excluded explicitly.
The given probability density function is the completion-time density for two independent sequential irreversible steps, with exponential distributions of waiting times at rates . Let . The convolution isIt is a two-stage hypoexponential distribution. The density vanishes at zero, initially rises as , peaks, and then decays with a tail governed by the slower rate. Setting its derivative to zero givesNormalization follows either from the convolution of normalized densities or directly from . In the equal-rate limit , the apparent singularity is removable andThis is an Erlang distribution with shape two. If one rate is much faster, its short waiting time becomes hard to resolve and the distribution resembles a single exponential distribution after its initial rise.
The observation supports at least two kinetically resolved stages before the recorded event, rather than one memoryless rate-limiting event. Possibilities include substrate binding followed by a conformational or catalytic step. It does not identify which stage is binding, chemistry, product release or rotation. The density is unchanged by exchanging and , so temporal order and chemical identity cannot be recovered from this fit alone. Other hidden-state mechanisms can also mimic the same density. Dependence of one fitted rate on ATP concentration, for example, would provide additional evidence about assignment, rather than following from the distribution by itself.
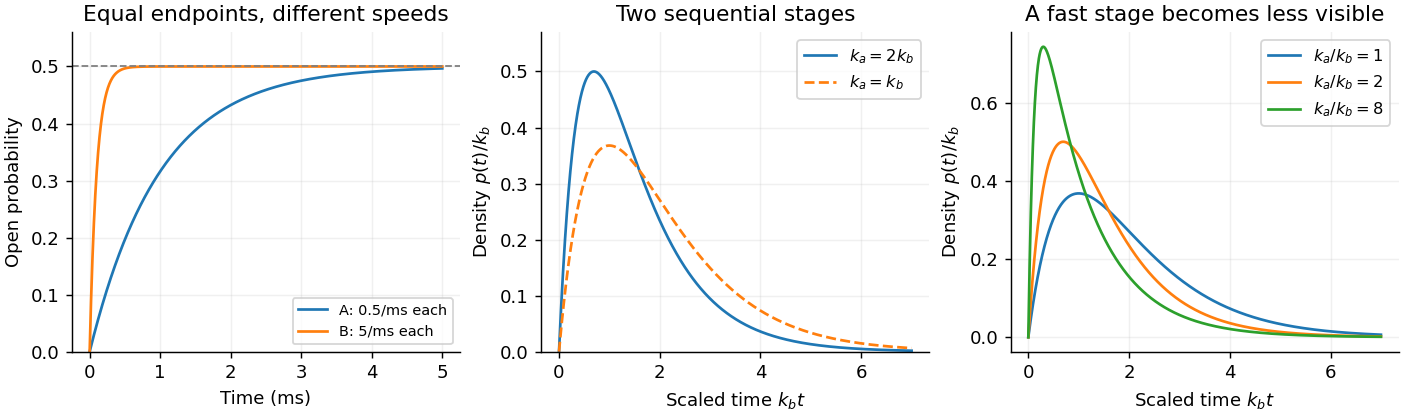


The left panel illustrates b for an initially closed population. The other panels are dimensionless examples of the distributions derived here; none reconstructs the missing source trace.
Magnetic tweezers apply force and controlled turns to a single torsionally constrained DNA tether joining a surface and a magnetic bead. A magnetic-field gradient pulls the bead; rotation of the field rotates the bead and changes imposed turns. Imaging measures bead height, hence DNA extension. At sufficient excess twisting and suitable force, DNA supercoiling creates writhed loops that shorten the tether. The extension-versus-turn curve supplies a calibration relating the removal or addition of supercoiling to vertical motion. Multiple attachments at both ends are needed conceptually to constrain rotation; freely swivelling DNA would relax the signal.
During transcription initiation, RNA polymerase binds a promoter in a closed complex and then opens a local DNA bubble, exposing template for RNA synthesis. In bacterial initiation a sigma factor helps recognize the promoter. Opening changes local helical twist. Because the total linking number is constrained,Unwinding the bubble means , so the remainder acquires a positive turn increment. A negatively supercoiled tether therefore loses some negative writhe and becomes longer; a positively supercoiled tether gains positive writhe and becomes shorter. Ordinary DNA compaction instead shortens both tethers. Comparing both signs separates local unwinding from compaction rather than assigning every height change to opening.
If is the number of turns unwound, the magnitude of the extension change per imposed turn, and the compaction, the symmetric-slope approximation predicts and . HenceMeasured unequal slopes can be retained in the two separate calibration equations. Multiplying by the helical repeat converts turns to an estimated number of unwound base pairs, subject to the mechanical model and the distinction between twist and writhe.
Repeated extension transitions reveal open-complex lifetimes, closed-complex lifetimes and their dependence on mechanical torque. Initial short RNA synthesis can occur while RNA polymerase remains associated with the promoter, including abortive initiation and DNA scrunching. Addition of nucleotides allows progression to promoter escape; an unwound bubble can then persist with elongation, so loss of the bubble is not the definition of escape. Successive cumulative unwinding events can identify renewed access to the same promoter after clearance. These are the measurement principles demonstrated by the paired-sign experiments.[ref]
Promoter unwinding and promoter clearance by RNA polymerase: Detection by single-molecule DNA nanomanipulation by Andrey Revyakin and others. Figures 1–4: extension calibration, opposite-sign opening responses and promoter-clearance readout
The experiment observes mechanics rather than directly imaging individual opened base pairs. Drift, nonspecific binding, tether integrity, force-dependent buckling and limited time resolution can mimic or obscure steps. A control lacking productive promoter recognition and agreement of the opposite-sign responses help identify the origin of a signal.
A point mutation changes an individual nucleotide position, introducing heritable sequence variation. Replication errors and spontaneous chemical damage are possible origins; mutation does not occur because a future change would be useful. Natural selection changes the frequencies of variants through their effects on reproduction, while genetic drift causes random frequency changes, especially in small populations. A sequence difference fixed between lineages is a substitution; many new mutations disappear and never become substitutions. Under a simple neutral diploid model, new neutral variants per generation times fixation probability gives substitution rate , motivating a molecular clock for suitable neutral sites.
To infer a phylogenetic tree, compare homologous, preferably orthologous sequences and align corresponding sites. A nucleotide substitution model converts observed sequence differences into estimated evolutionary branch lengths, allowing for multiple changes at one site. For example, in the equal-frequency, equal-rate Jukes-Cantor model, a site changes to any alternative with rate , where is its total substitution rate. The probability of matching its initial state after elapsed time is . Thus for an evolutionary separation with expected substitutions per site , the observed mismatch fraction is andFor small , ; near saturation , large evolutionary distances become difficult to distinguish. The correction does not rescue fully saturated data. Real analyses may allow unequal nucleotide frequencies, transition biases and variation of rates between sites.
Tree topology and branch lengths can be estimated using maximum likelihood estimation or Bayesian phylogenetic inference, with support assessed from resampling or posterior uncertainty. A gene tree need not equal the species tree, because incomplete lineage sorting, genetic recombination and horizontal gene transfer can produce different histories. Rooting with an appropriate outgroup establishes ancestral direction; pairwise differences alone do not supply a root.
Branch lengths measured in substitutions per site are not yet calendar times. Under a strict molecular clock with rate per site per year, a branch of duration has length ; two contemporaneously sampled descendants separated at time have total path length , hence . Calibration comes from independently dated constraints such as fossil calibration, geological information or sufficiently informative sampling dates. A dated common ancestor is not automatically identical in age to the earliest observed fossil. A relaxed molecular clock permits branch-specific rates while imposing a statistical model on them.
Without an external temporal calibration or an independently known rate, multiplying all times by a constant and dividing all rates by that constant leaves expected sequence divergence unchanged. Absolute dates are therefore unidentifiable from sequence divergence alone. A useful time-calibrated phylogenetic tree reports uncertainty from sequence sampling, substitution models, rate variation, topology and calibration bounds, rather than displaying an unjustified exact date for each ancestor.
An ion channel supplies a water-accessible pathway through the otherwise strongly insulating, hydrophobic lipid bilayer. Moving a hydrated ion directly into that low-permittivity region is energetically expensive. The channel's polar pore and coordinating groups reduce the barrier while retaining selectivity. Transport is passive: the electrochemical potential difference drives net movement, and the channel does not itself pump an ion uphill. For an ideal species of charge number ,The net electric current of a pathway permeable only to that species reverses at its Nernst equilibrium potential. An open channel can carry zero net electric current at that voltage even while individual ions cross both ways.
The selectivity filter balances loss of the hydration shell against favorable coordination within the pore. In a typical potassium channel, backbone carbonyl oxygen atoms create appropriately arranged binding sites for a substantially dehydrated potassium ion. The smaller sodium ion is not automatically preferred simply because it fits geometrically: its hydration is stronger and the filter does not compensate its dehydration in the same way. Geometry, flexibility, coordination chemistry and ion occupancy all matter. Sodium-selective and calcium-selective channels use different arrangements, often involving charged side chains; a single rigid-sieve explanation is inadequate.
Rapid passage coexists with transient binding because several nearby sites have comparable free energies and relatively low barriers between them. Multiple ions occupying a pore influence one another through electrostatic interactions, enabling correlated exchange and displacement rather than irreversible trapping at one deep site. High throughput therefore requires favorable passage kinetics as well as strong discrimination against unwanted species. A more tightly bound species can even block a pathway instead of being its fastest permeant.
Gating and selectivity are distinct. A voltage-sensing domain couples membrane potential to activation-gate conformation; ligand-gated ion channels couple binding to opening; mechanosensitive ion channels couple membrane stress to opening. Ion-channel inactivation introduces additional unavailable states. None of these mechanisms alone identifies which ion the open pore prefers.
For similar independent channels, a local Ohmic approximation with outward-positive electric current isHere open probability controls availability, single-channel electrical conductance controls current per open pore, and is the electrical driving factor in this approximation. Nonlinear current-voltage relations and mixed-ion permeation may require a fuller model. Single-channel recording resolves discrete openings; reversal shifts when ionic concentrations change test selectivity; gating electric currents test sensor motion; structural evidence identifies the pore and its coordinating groups. Together these distinguish rapid selective permeation from the separate process of opening the gate.
A small swimming member of the bacteria samples too little spatial separation across its own body to compare weak chemical gradients accurately. Temporal sensing in bacterial chemotaxis instead compares recent receptor activity with an adapted memory. Straight runs carry it between locations; tumbles reorient it. When conditions improve, tumbling becomes less frequent and favorable runs last longer. An unfavorable direction is abandoned sooner. Repetition creates a biased random walk without requiring the cell to identify the best direction at each tumble. Rotational diffusion and environmental obstacles eventually destroy the benefit of maintaining a direction indefinitely.
An order-of-magnitude transport estimate compares the time to swim a distance with the time for an attractant to diffuse that distance. Their crossover is a Péclet number of order one:This estimates a useful run scale: over much shorter distances, nutrient diffusion rapidly replenishes the local environment compared with the organism's displacement. Runs of order a second also permit comparison along a substantial path before reorientation. The given and determine this scale, not an exact biological mean run length or a complete sensory optimization. If a one-coordinate root-mean-square diffusion length is equated with , the resulting prefactor is two; a three-dimensional root-mean-square convention gives six. Such constants cannot be fixed by a scaling argument alone.
For a cilium of length , compare its advective time with the local diffusion time . Significant stirring relative to diffusion requires , soThis is about times the stated swimming speed. If the tip travelled on a circular path of radius , that speed would correspond to an illustrative frequency . That geometric conversion is not a specified beat pattern. At microscopic scales the flow has small Reynolds number; useful transport depends on the actual nonreciprocal motion and resulting flow, and rapid motion alone does not guarantee increased uptake. The scaling explains why simply stirring with a very short appendage is demanding when molecular diffusion is already fast.
Articles by others on the same topic
There are currently no matching articles.