Jack W. Szostak is a molecular biologist whose research connects chromosome maintenance, laboratory selection of functional molecules and experimental models of early life. His joint 2009 Nobel Prize in Physiology or Medicine recognized discoveries concerning chromosome protection by telomeres and telomerase.[ref] That recognition concerns chromosome ends, a distinct contribution from his work on molecular selection.
With Richard W. Roberts, Szostak coauthored the 1997 experiment that covalently attached an encoded peptide to its mRNA, providing a basis for mRNA display. Synthetic messages carried puromycin at the 3′ end; translation produced RNA–peptide fusions. The authors demonstrated selective recovery using an epitope and immunoprecipitation.[ref]
RNA-peptide fusions for the in vitro selection of peptides and proteins by Richard W. Roberts and Jack W. Szostak. Abstract and Introduction; RNA–peptide fusion and selection demonstration
The experimental question was how to recover the sequence encoding a selected protein after selecting its physical binding behavior.The connection between genotype and phenotype is what makes iterative selection possible. As an original bookkeeping example, if an intact linkage survives each of five processing steps with stipulated probability 0.9, independent losses leave of the initially linked molecules. Even a useful covalent design does not by itself guarantee complete recovery through a multistep experiment. Retention of encoding information and the selected molecular function must both be checked.
His university research account also describes work toward copying RNA without enzymes and coupling replication to simple membrane-bounded systems in origin-of-life experiments. It distinguishes experimental progress from unresolved problems in repeated replication and chemical compatibility.[ref]
Jack W. Szostak research profile by the University of Chicago Department of Chemistry. The Origins of Life; Nonenzymatic RNA Replication; Model Protocells
These laboratory models explore possible mechanisms rather than directly observing the historical origin of life.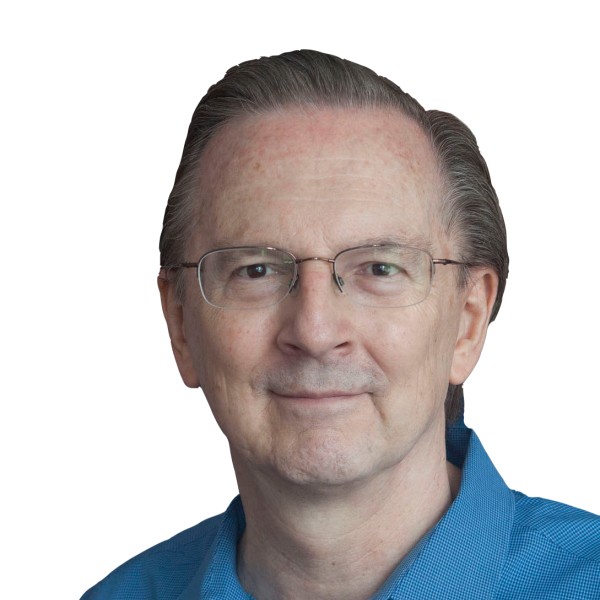
Jack W. Szostak in his University of Chicago faculty portrait
. Richard W. Roberts researches protein engineering and molecular recognition, including methods that select peptides and proteins while retaining their encoding nucleic acid. His official chemistry profile identifies the same full byline as the 1997 research paper with Jack W. Szostak and describes a research focus on mRNA display.[ref]
Their 1997 experiment addressed a practical information problem: after recovering a protein with a useful property, how can its encoding sequence be recovered and amplified? A synthetic mRNA carried a 3′ puromycin group that accepted the growing peptide during translation, producing a stable RNA–peptide linkage. Selective immunoprecipitation of fusions carrying the chosen epitope provided an experimental demonstration.[ref]
RNA-peptide fusions for the in vitro selection of peptides and proteins by Richard W. Roberts and Jack W. Szostak. Abstract and Introduction, recovery of encoding information and selection of epitope fusions
An original yield calculation illustrates why fusion formation and binding selection are separate experimental questions. Let a preparation contain encoding molecules, let fraction form the intended fusion, and let fraction of those fusions survive a stipulated recovery step. The recovered count isRaising to 0.50 doubles that count without changing . Increased recovered material alone therefore does not prove that the encoded peptide acquired greater binding affinity. Appropriate comparison distinguishes expression or fusion efficiency, nonspecific recovery and the function under selection.
The broader methodological contribution is a physical route from selected phenotype back to recoverable genotype. mRNA display supports experiments on molecular recognition without requiring each candidate protein to remain inside a living cell. The resulting selection still reports performance under the actual assay conditions; it does not by itself establish activity in an organism or therapeutic effectiveness.

L. Gold, also published as Larry Gold, is a molecular biologist associated with the development of SELEX, a method of repeated selection and amplification of nucleic-acid ligands. Gold's own retrospective connects the abbreviated 1990 byline with the work carried out with Craig Tuerk at the University of Colorado.[ref] His university profile also documents the laboratory's bacteriophage research and identifies him as Larry Gold.[ref]
The 1990 primary experiment randomized eight positions in an RNA region interacting with a DNA polymerase. Repeated binding selection and amplification recovered two sequences with equivalent binding constants, including a variant differing from the original at four positions.[ref]
Systematic evolution of ligands by exponential enrichment: RNA ligands to bacteriophage T4 DNA polymerase by C. Tuerk and L. Gold. Abstract, randomized region and selected-sequence comparison
This showed that sequence variation could be explored experimentally through measured molecular function.The distinction between enrichment and absolute yield is important. In an original two-class example, begin with 1,000 desired molecules and 99,000 others. Suppose one selection retains 20% of the desired class and 1% of the others. There are then 200 desired and 990 other molecules: the desired fraction rises from 1% to , even though most desired molecules were lost. Equal amplification of both surviving classes restores material without changing that fraction.
Thus a selection cycle can improve population composition while decreasing the absolute number of every class. Amplification makes another round possible; it does not retrospectively recover a sequence lost completely. This explains the experimental importance of both discrimination and population survival when interpreting in vitro selection.
Gold's firsthand account situates the work in a change of perspective: functional RNA can be investigated as a folded binding molecule rather than only as an information carrier.[ref] Selection provides evidence about performance under the assay conditions; an aptamer recovered there still requires separate characterization for another environment.

C. Tuerk, identified as Craig Tuerk in L. Gold's firsthand account, coauthored the 1990 experiment establishing SELEX. The retrospective describes his investigation of the RNA binding site involved in regulation of a bacteriophage DNA polymerase and attributes the process name to him.[ref] The fuller name is supported by that participant account; no middle name or life dates are inferred from the abbreviated research byline.
The primary paper selected RNA ligands after randomizing an eight-base region. There are possible strings for that region. Two recovered sequences differed at four positions yet had equivalent binding constants in the reported comparison.[ref]
Systematic evolution of ligands by exponential enrichment: RNA ligands to bacteriophage T4 DNA polymerase by C. Tuerk and L. Gold. Abstract, sequence-pool size and binding-constant comparison
The observation distinguishes a difference in nucleotide sequence from a difference in the measured molecular function.An original geometric count sharpens the scale of the experiment. Around one fixed eight-base string, the number of strings differing at exactly four positions isThe binomial coefficient chooses which four positions change; each chosen position has three alternative bases. A selected four-change sequence is consequently one of thousands at that same sequence distance. Sequence distance alone does not identify the functional substitute; the binding experiment supplies that information.
The theoretical count also does not prove that a physical preparation contains every possible sequence at equal abundance. Synthesis, survival during selection and subsequent polymerase chain reaction amplification can affect representation. Conversely, equivalent binding affinity to one target does not imply equal behavior toward every other target or equal stability in every environment.
Tuerk's contribution illustrates how a focused question about an RNA regulatory binding site could motivate a reusable experimental search procedure. A candidate's sequence can be recovered after selection and its binding measured separately, connecting the population-level search to an individually testable molecular result.
Titus H. J. Huisman investigated hemoglobin variation and the different globin chains composing it. His university laboratory's history identifies him as a leader of hemoglobin research at the Medical College of Georgia.[ref]
Titus H. J. Huisman coauthored the 1968 chemical study distinguishing glycine and alanine at position 136 in gamma-globin material. That observation supported the presence of multiple sequence forms within human fetal hemoglobin.[ref]
Evidence for multiple structural genes for the gamma chain of human fetal hemoglobin by W. A. Schroeder and others. Primary byline; page 537, opening paragraph
It was evidence from protein chemistry, rather than direct sequencing of the modern HBG1 and HBG2 genes.In a 1970 report with W. A. Schroeder and colleagues, Titus H. J. Huisman followed G-gamma globin and A-gamma globin proportions in seven infants after birth. The reported G-gamma-to-A-gamma ratio changed from approximately 3:1 at birth toward 2:3 by five months.[ref]
Changes in the Nature of Human Fetal Hemoglobin During the First Year of Life by Titus H. J. Huisman and others. Abstract and primary byline
This study connects molecular discrimination with a time-dependent biological observation, rather than treating all gamma chains as one unchanging category.An original accounting example shows why a ratio is not an absolute production rate. A 3:1 ratio corresponds to G-gamma and A-gamma fractions and ; a 2:3 ratio corresponds to and . If a stipulated total pool falls from 100 to 20 units, the A-gamma amount falls from 25 to 12 even though its fraction rises from 0.25 to 0.60. These totals are invented for the calculation, not measured values from the paper.
The scientific distinction is useful whenever developmental changes are described by a normalized molecular signal. Sequence identity, relative abundance, absolute amount and assembly pairing remain different questions. Gamma-chain pairing bounds from bulk composition further shows why chain fractions alone do not identify every fetal hemoglobin assembly.

W. A. Schroeder, also identified as Walter A. Schroeder in a primary publication, investigated hemoglobin structure and the chemical differences among its globin chains. A contemporary Caltech account describes his work on fetal hemoglobin sequences and its distinction from adult hemoglobin.[ref]
Retirements 1986 in Engineering and Science. Printed page 31, Walter Schroeder subsection
The 1968 study on which W. A. Schroeder is first author reported glycine and alanine at position 136 in human fetal gamma-chain material. This supplied chemical evidence that the label gamma-globin encompassed distinguishable sequence forms, motivating the inference of multiple structural genes.[ref]
Evidence for multiple structural genes for the gamma chain of human fetal hemoglobin by W. A. Schroeder and others. Page 537, opening paragraph
The historical observation concerned protein chemistry; it was not direct sequencing of the modern HBG1 and HBG2 loci.A 1970 report with Titus H. J. Huisman followed the changing proportions of G-gamma globin and A-gamma globin after birth. Its primary author information explicitly spells Walter A. Schroeder and gives the Caltech affiliation.[ref]
Changes in the Nature of Human Fetal Hemoglobin During the First Year of Life by Titus H. J. Huisman and others. Abstract; Authors and Affiliations
That documented connection supports the expanded-name alias while preserving the original initial-based identifier.An original analytical example explains why chemical discrimination and quantitative abundance are separate achievements. Suppose sequence-selective signals are and , with responses per unit amount and . The corresponding amounts are 60 and 20, so the G-gamma chain fraction is , rather than the raw signal fraction 0.60. These stipulated numbers illustrate response correction; they are not measurements reported by Schroeder.
Even correctly measured chain proportions leave another question open: which forms share the same fetal hemoglobin assembly? Gamma-chain pairing bounds from bulk composition shows why that requires additional molecular information. Distinguishing sequence, abundance and assembly makes the scientific contribution of the chemical studies more precise.
A labeled portrait appears on page 31 of Caltech's 1986 retirement notice.
R. Fourme coauthored the 1984 high-resolution structure of human deoxyhemoglobin with G. Fermi, Max Perutz and B. Shaanan. The primary record describes X-ray diffraction collected on film at room temperature using synchrotron radiation, followed by refinement at resolution.[ref]
The crystal structure of human deoxyhaemoglobin at 1.74 A resolution by G. Fermi and others. Primary byline and Abstract, experimental apparatus
This establishes a contribution to experimental structural biology; the byline alone does not assign each instrument or calculation to one coauthor.The physical chain is illuminating: a crystal scatters the incident radiation, a detector records diffraction intensity, and a model connects the measurements with an electron density distribution. Improving incident flux can help collect weak reflections, but flux and spatial resolution remain different quantities.
For an original counting example, suppose an ideal detector records 400 independent photons in one reflection and background is negligible. A Poisson distribution then gives standard uncertainty counts, or 5% relative uncertainty. Four times as many recorded photons gives 1,600 counts and relative uncertainty. The square-root improvement describes the chosen counting model; it is not a measured uncertainty for the historical film detector.
Increasing exposure also need not preserve the specimen unchanged. A comparison should keep track of radiation-induced changes, detector response and background rather than assuming that every additional count is interchangeable. Those experimental controls complement the molecular constraints used in X-ray crystallography. Fourme's verified coauthorship connects the biological question to the apparatus needed to obtain its structural evidence.
B. Shaanan determined a human oxyhemoglobin structure by single-crystal X-ray crystallography in a 1983 paper bearing that exact single-author byline. The experiment used an Arndt–Wonacott camera at and reported a resolution of .[ref]
Structure of human oxyhaemoglobin at 2.1 A resolution by B. Shaanan. Byline; Abstract, diffraction apparatus and resolution
It supplied an experimental arrangement against which molecular explanations of hemoglobin oxygen binding could be tested.The paper distinguished the local environments in the alpha-globin and beta-globin subunits and specified iron positions relative to defined porphyrin planes. Shaanan also coauthored the later deoxyhemoglobin study with G. Fermi, Max Perutz and R. Fourme.[ref]
The crystal structure of human deoxyhaemoglobin at 1.74 A resolution by G. Fermi and others. Primary byline
Comparing the two ligation states requires consistent geometric definitions; distances from different reference planes are different observables.An original plane-fitting example makes that distinction concrete. A point at coordinate angstrom is 0.30 angstrom from the plane , but only 0.10 angstrom from the parallel plane angstrom. The atom has not moved. Changing the atoms used to define a fitted plane can change a reported displacement.
For a unit normal and point on a reference plane, signed distance from position isReversing the normal reverses the sign; translating both the model and its plane by the same vector leaves unchanged. These are geometric checks for interpreting a structural comparison, not additional measurements from the paper.
The value of the structural work is therefore more specific than an image of a protein: it turns questions about oxygen-binding geometry into quantities with stated reference conventions and experimental support. The published initials are retained rather than assuming a full identity from a similarly named crystallographer.
G. Fermi is the first author of the 1984 study refining human deoxyhemoglobin at resolution, and is named with Max Perutz as a depositor of its primary structure record, 4HHB.[ref]
The crystal structure of human deoxyhaemoglobin at 1.74 A resolution by G. Fermi and others. RCSB 4HHB, deposition authors, primary citation and Experimental Data Snapshot
The published initial-only identity is retained.The model allowed the investigators to examine local geometry around iron in the oxygen-binding groups of hemoglobin. Their analysis also tested whether conclusions about that geometry persisted when selected model restraints were removed.[ref]
The crystal structure of human deoxyhaemoglobin at 1.74 A resolution by G. Fermi and others. Primary Abstract, unrestrained refinement from modified geometry
Such a comparison asks how strongly the observations constrain a feature independently of assumptions imposed during fitting.An original one-parameter calculation illustrates this issue without recreating the historical refinement. Suppose a scalar coordinate is constrained by an observed value with uncertainty , and by a preferred value with strength . MinimizeFor , , and in compatible units, . Removing the restraint gives . A fitted value can therefore depend on both observations and a modeling choice. This toy weighted least squares problem is not the algorithm used for 4HHB.
Agreement with data should consequently be examined together with restraint sensitivity and independent physical evidence. A precise-looking coordinate is not established merely by printing many decimal places. Fermi's documented work exemplifies using X-ray crystallography to make a molecular arrangement quantitatively testable.
Max Perutz, fully Max Ferdinand Perutz, investigated protein structure through X-ray diffraction. The Nobel nomination archive explicitly pairs his full name with the initials M. F. Perutz and records his 1962 Chemistry prize.[ref]
Max Ferdinand Perutz in the Nobel Prize nomination archive. Nomination Chemistry 1962, number 64-0, nominee name and comment
His structural research connected the organization of protein atoms with the behavior of oxygen-binding hemoglobin.With J. A. Frier, Perutz coauthored the structure of human fetal deoxyhemoglobin. The associated primary record, 1FDH, identifies alpha and gamma chains and a diffraction resolution of .[ref]
Structure of human foetal deoxyhaemoglobin by J. A. Frier and M. F. Perutz. RCSB PDB 1FDH, Literature, Biological Assembly and Experimental Data
The study relates structural differences to interaction with 2,3-Bisphosphoglycerate. A structural explanation is strongest when its proposed interaction can also be tested through functional measurements.An original diffraction example shows why obtaining a molecular structure is not equivalent to photographing atoms directly. In a reduced model, a complex scattering amplitude iswhere are positions and scattering strengths. Measured intensity is proportional to . Translating every atom by multiplies by without changing that intensity. Intensities therefore do not by themselves fix every phase or coordinate convention. Recovering an electron density model requires appropriate phase information and structural constraints, connecting the measurement with the phase problem.
The fetal structure also illustrates a separate limitation: identifying a deoxygenated conformation does not directly measure the entire oxygen-binding curve. Structural comparisons, ligand-binding experiments and quantitative analysis supply complementary evidence. Perutz's contribution lies in connecting those molecular questions with experimentally determined three-dimensional organization.
J. A. Frier coauthored the 1977 study of human fetal deoxyhemoglobin with Max Perutz. The primary structure record 1FDH reports a human alpha/gamma tetramer examined by X-ray diffraction at resolution.[ref]
Structure of human foetal deoxyhaemoglobin by J. A. Frier and M. F. Perutz. Primary citation, Biological Assembly, Macromolecules and Experimental Data
This is a verified structural-biology contribution; the published initials are retained without inferring an expanded identity from a deposition-name variant.Comparing fetal and adult hemoglobin structures tests how changes in subunit composition alter local interactions. The paper's indexed abstract discusses differences at the binding region for 2,3-Bisphosphoglycerate.[ref]
Structure of human foetal deoxyhaemoglobin by J. A. Frier and M. F. Perutz. Indexed primary publisher abstract
Such a comparison can suggest why an effector binds differently. It does not make a static crystal structure a direct measurement of all physiological response rates or affinities.An original coordinate example distinguishes spatial resolution from uncertainty in a derived distance. Suppose two modeled positions along one axis differ by , with independent standard uncertainties each. The uncertainty of their signed separation isIf the coordinate errors have covariance , the variance becomes . Shared model errors can therefore change the uncertainty of a relative separation. These are illustrative assumptions, not coordinate uncertainties reported for 1FDH.
A quoted diffraction resolution alone should not be substituted for either number in this example. It describes the experimental detail available to the model, while coordinate uncertainty also depends on data, occupancy, disorder and refinement. Frier and Perutz's structural result is valuable because it offers a molecular arrangement against which those broader mechanistic questions can be tested.
Allan M. Maxam, with Walter Gilbert, developed a chemical approach to DNA sequencing reported in 1977. Their primary paper describes partial cleavage of terminally labeled DNA at selected bases, then size separation of labeled fragments by polyacrylamide gel electrophoresis.[ref]
A new method for sequencing DNA by Allan M. Maxam and Walter Gilbert. PNAS 74, pp. 560–564, original full byline and Abstract
Maxam-Gilbert sequencing converts chemical selectivity into a position-dependent pattern along a pre-existing strand.The word partial is central to the measurement logic. Different molecules contribute fragments ending at different positions, while a common labeled end supplies a reference. Extensive repeated cleavage of one molecule can remove information about more distant sites, because only the fragment still containing the label is visible in the corresponding readout.
An original probability model illustrates the tradeoff without prescribing experimental conditions. Suppose the number of relevant breaks in one molecule follows a Poisson distribution with mean . ThenAt stipulated , about 81.9 percent have no break, 16.4 percent have one and 1.75 percent have multiple breaks. At , those fractions are about 13.5, 27.1 and 59.4 percent. More cleavage does not translate monotonically into a more interpretable set of singly terminated fragments. Actual sequence-dependent chemistry need not follow this simplified independent-break model.
The measured gel is also not the sequence itself. Reading requires calibrating fragment size, identifying the labeled end and comparing the base-selective channels. Overlapping bands or an incorrect orientation can corrupt an otherwise real chemical signal. Maxam's contribution demonstrates how a carefully designed molecular measurement can expose sequence information that bulk composition alone cannot recover.
Pamela M. Pollock investigates molecular alterations associated with cancer. She is first author of a 2003 study that analyzed BRAF mutations in microdissected samples classified as melanocytic nevus or melanoma.[ref]
High frequency of BRAF mutations in nevi by Pamela M. Pollock and others. Nature Genetics 33, pp. 19–20, primary byline and Abstract
The study found the substitution labeled V599E in its original numbering in 63 of 77 nevi. The presence of that alteration in benign lesions made it important to distinguish a pathway-associated initiating event from sufficient evidence of malignant transformation.Microdissection connects a molecular result with a selected tissue region. It can enrich the cells relevant to the question, while leaving the sample's actual composition and sequencing sensitivity as separate measurement issues. A mutation detected in a mixed tissue sample is not automatically present in every constituent cell.
An original allele-count model illustrates the distinction. Suppose 30 percent of the sampled cells carry one altered BRAF copy and one unaltered copy, while all remaining cells have two unaltered copies. With equal DNA contribution per cell, no copy-number alteration and no measurement bias, the expected variant allele fraction isA 15 percent variant fraction is therefore compatible with 30 percent mutation-bearing cells in this model. It would be incorrect to identify those two percentages without the copy-count assumptions. Different purity, copy number or recovery changes the relationship.
The published paper's selected-lesion frequencies answer another question again: how many sampled lesions had the detected alteration. A lesion-level proportion such as is neither a within-lesion allele fraction nor the probability that an individual lesion will progress. Pollock's team's contribution links molecular pathway evidence with those distinct biological levels of interpretation, rather than making mutation presence a self-sufficient classification of malignancy.
Benno Müller-Hill, with Walter Gilbert, isolated the lac repressor in work published in 1966. The research connected a genetically inferred control function with an experimentally detectable molecular component. Their primary paper developed an assay based on binding an inducing small molecule, detected through equilibrium dialysis against radioactive Isopropyl β-D-1-thiogalactopyranoside.[ref]
Isolation of the lac repressor by Walter Gilbert and Benno Müller-Hill. PNAS 56, printed p. 1891, assay description and Design of the Experiment
That choice was logically important. The opening discussion considered a model in which repressor binds the lac operator, but the assay sought inducer interaction without requiring every detail of the repression mechanism to be settled first. Measuring binding to an inducer is not itself the same experiment as demonstrating binding to operator DNA.
An original one-site model explains the dialysis signal. A membrane permits free ligand to equilibrate between two compartments while retaining protein on one side. If the common free ligand concentration is , total binding-site concentration is and dissociation constant is , the retained bound concentration isThe protein-containing compartment has total ligand concentration , while the comparison compartment has under this ideal model. With stipulated , , the totals are 2.5 and 2.0 micromoles per litre. Their excess measures binding even though free ligand equilibrates.
These values are illustrative, not fitted constants from the 1966 experiment. Nonspecific adsorption, unequal accessible volumes and other binding components require controls. Protein purification can then be followed with a functional assay rather than by the mere presence of protein material. This is the experimental bridge between a gene-control hypothesis and a molecular mechanism; the joint paper does not justify assigning every experimental operation to Müller-Hill alone.
Walter Gilbert helped turn questions about gene regulation and DNA sequence into experimentally accessible molecular measurements. He moved from physics toward molecular biology at Harvard; the Cold Spring Harbor Laboratory archive records that transition and preserves his research collection.[ref]
Walter Gilbert in the Cold Spring Harbor Laboratory archives. Institutional biography and Scope and Content

Walter Gilbert, portrait identified by Cold Spring Harbor Laboratory and credited to Harvard News Service
. Source. With Benno Müller-Hill, Gilbert reported isolation of the lac repressor in 1966. Their assay detected binding to an inducing small molecule using equilibrium dialysis, providing a biochemical handle on a genetically inferred control protein.[ref]
Isolation of the lac repressor by Walter Gilbert and Benno Müller-Hill. printed p. 1891, assay description
The distinction between a measurable binding activity and a fully established regulatory mechanism remains important: the assay did not assume that every detail of operator recognition was already known.With Allan M. Maxam, he later developed the chemical DNA sequencing method now called Maxam-Gilbert sequencing. Their 1977 paper described partial cleavage at selected bases in terminally labeled DNA, followed by polyacrylamide gel electrophoresis to resolve fragment lengths.[ref]
A new method for sequencing DNA by Allan M. Maxam and Walter Gilbert. PNAS 74, pp. 560–564, Abstract
A ladder of labeled fragment lengths converts chemical specificity into information about positions along a strand.An original schematic inference illustrates the readout. Suppose calibrated fragment-length patterns assign A to positions 1 and 6, G to positions 2 and 5, C to positions 3 and 7, and T to positions 4 and 8. Combining the position sets gives AGCTGACT in the stated reading direction. A G-selective measurement alone would leave six positions unresolved; the different chemical channels supply complementary information.
The example abstracts away migration calibration, cleavage-site offsets and band overlap. Real interpretation requires identifying the labeled end and relating each band to the appropriate base position. This sequencing approach interrogates a pre-existing labeled strand through cleavage, so its measurement logic is different from inferring sequence from terminated synthesis products. Gilbert's work exemplifies how designing a recoverable physical signal can transform an otherwise indirect molecular question.
B. J. Hoffman coauthored the 1983 cDNA library method with U. Gubler. Their procedure connected first-strand copying of messenger RNA with an enzymatic second-strand synthesis step, simplifying recovery of double-stranded DNA copies.[ref]
A simple and very efficient method for generating cDNA libraries by U. Gubler and B. J. Hoffman. Primary byline and Abstract
The paper establishes this collaborative contribution; it does not assign every laboratory step to an individual author.A particularly informative validation used a previously sequenced bovine transcript and tested the recovered copies by sequencing. The abstract reports faithful full-length products for that test.[ref]
A simple and very efficient method for generating cDNA libraries by U. Gubler and B. J. Hoffman. Abstract, known bovine preproenkephalin RNA comparison
Sequence comparison addresses what the method preserved, rather than merely showing that DNA material was produced.An original probability example separates two quality questions. Let mean that a recovered copy is full length, and mean that its sequence meets a stated correctness criterion. ThenIf a stipulated preparation has and , the fraction satisfying both criteria is 0.76. Multiplying marginal probabilities would require an additional independence assumption; the conditional expression does not.
For a cDNA library, a product can therefore have plausible length but sequence changes, or correct sequence over a shorter region while lacking an end. Validation should state which region was read and which criterion was tested. An illustrative 100-copy sample with 80 full-length products and 76 meeting both conditions has 95 percent correctness among the full-length products but 76 percent joint success across all recovered copies. These numbers explain the distinction and are not measurements from the original study.
Hoffman's initials remain as printed. The verified molecular-method contribution is sufficient to describe the research without inventing personal dates, an expanded name or a portrait.
U. Gubler is first author, with B. J. Hoffman, of a 1983 method for generating cDNA libraries from small amounts of messenger RNA. The method combined first-strand synthesis with second-strand synthesis mediated by Ribonuclease H and DNA polymerase I.[ref]
A simple and very efficient method for generating cDNA libraries by U. Gubler and B. J. Hoffman. Gene 25, pp. 263–269, primary byline and Abstract
This contribution made information in an RNA population accessible through recoverable DNA copies. The published initial is retained without assuming an expanded identity.A cDNA library represents the RNA molecules successfully copied and recovered, rather than automatically representing every gene equally. Reverse transcription, second-strand synthesis and subsequent recovery can change representation. A useful library therefore requires more than a high total number of clones: rare transcripts and full-length products must also be considered.
An original sampling calculation isolates the rare-transcript question. Suppose a target transcript has probability in each independent recovered clone. The probability that examined clones contain no target isFor , this is about 0.448. Requiring at least 95 percent probability of finding one target givesso at least 299 clones are needed under the stated independent-sampling model. Copying bias that lowers the recovered target fraction changes that calculation even if the starting RNA fraction was known.
The primary method paper also tested fidelity against a known transcript sequence.[ref]
A simple and very efficient method for generating cDNA libraries by U. Gubler and B. J. Hoffman. Abstract, full-length sequence validation
Representation, product length and sequence agreement are distinct measurements. A large library can still undersample a rare transcript or contain truncated copies; neither conclusion follows from clone count alone.Daniela Teixeira is a molecular biologist whose published research investigated P-body assembly and the relationship between messenger RNA organization and biological translation. She was first author of a2005 study with Roy Parker's group showing that yeast P-bodies depend on RNA for their assembly and contain nontranslating messages.[ref]
Processing bodies require RNA for assembly and contain nontranslating mRNAs by Daniela Teixeira and others. Primary byline and Abstract, RNA dependence and nontranslating mRNAs
The study combined cellular observations with experiments on enriched P-body preparations. Digestion with a ribonuclease disrupted the preparations, testing whether RNA contributed to their integrity. In cells, perturbing translation initiation and trapping messages in polysomes produced contrasting P-body responses. Those comparisons distinguished effects on RNA availability from the vague statement that translation had been inhibited.[ref]
Processing bodies require RNA for assembly and contain nontranslating mRNAs by Daniela Teixeira and others. Abstract; Figure6 and associated Results, RNA digestion and translation-state comparisons
An original image-analysis example illustrates a limit of such observations. Suppose a hypothetical cell contains ten visible foci with four arbitrary fluorescence units each, while a second condition contains five foci with eight units each. Both have40 units assigned to the foci. Counting foci alone would miss that equivalence; conversely, equal total signal would not establish identical molecular composition or RNA decay activity. The numbers are stipulated, not measurements from Teixeira's paper.
The Parker group's institutional alumni directory records her training period as2001–2005.[ref]
Alumni of the Parker Research Group. Grad Students section, Daniela Teixeira entry
Her scientific contribution here is the experimental connection among RNA, translation state and the assembly of a visible cytoplasmic structure.Ujwal Sheth is a molecular biologist associated with experimental studies of messenger RNA localization and turnover. A 2003 paper with Roy Parker, listing a University of Arizona research affiliation, investigated the relationship between RNA decay and discrete cytoplasmic structures named P-bodies.[ref]
Decapping and Decay of Messenger RNA Occur in Cytoplasmic Processing Bodies by Ujwal Sheth and Roy Parker. Primary byline, affiliation and Abstract.
The study combined localization with functional perturbations. Factors involved in decapping accumulated in cytoplasmic foci, and RNA degradation intermediates localized there. Interfering with different steps of RNA turnover changed the abundance of the foci. These comparisons helped distinguish a compartment associated with processing from a fluorescent aggregation seen without a functional test.[ref]
Decapping and Decay of Messenger RNA Occur in Cytoplasmic Processing Bodies by Ujwal Sheth and Roy Parker. Abstract and Figures1–2: localization and changes following pathway perturbations.
Sheth also coauthored a 2005 study linking P-body assembly to nontranslating RNA. Its experiments connected assembly to RNA integrity and showed contrasting responses to inhibition of translation initiation and elongation.[ref]
Processing bodies require RNA for assembly and contain nontranslating mRNAs by Daniela Teixeira and others. Abstract and Figure6: translation-state comparisons and RNA-dependent integrity.
The work made RNA's participation a testable part of the explanation, beyond treating the visible focus as a container for proteins.The broader scientific lesson is to connect three kinds of evidence: where molecules localize, what happens when a pathway changes, and whether the proposed components are needed for assembly. A change in spot number alone does not identify the direction or rate of RNA processing. These questions connect cell organization with the lifetime and use of a genetic message.
A captioned laboratory photograph of Sheth beside Roy Parker appears on PDF page 9 of HHMI Bulletin September 2003 by Howard Hughes Medical Institute.
Roy Parker is a molecular biologist studying how cells produce, translate and degrade messenger RNA. His University of Colorado Boulder profile describes this research program, and his institutional curriculum vitae records a genetics doctorate from the University of California, San Francisco in 1985.[ref]
Roy Parker profile at the University of Colorado Boulder. Research description; institutional CV p.1, education.
His early work helped establish a sequence-recognition role for U2 small nuclear RNA. In a 1987 collaborative study, compensating changes in U2 suppressed corresponding changes in the yeast intron branch region. The matching suppression connected RNA sequence to base pairing in RNA splicing, rather than merely associating U2 with the machinery.[ref]
Recognition of the TACTAAC box during mRNA splicing in yeast involves base pairing to the U2-like snRNA by Roy Parker and others. Abstract, branch-site mutations and allele-specific compensatory U2 changes.
A later study with Ujwal Sheth connected RNA decay to cellular location. Functional fusions to green fluorescent protein placed several decapping factors in discrete cytoplasmic foci, termed P-bodies. Perturbing steps of the decay pathway changed those foci, and RNA decay intermediates also localized there.[ref]
Decapping and Decay of Messenger RNA Occur in Cytoplasmic Processing Bodies by Ujwal Sheth and Roy Parker. Abstract and Figures1–2: localization and pathway perturbations.
The combination of imaging and perturbation is scientifically important. A fluorescent spot alone shows where a marked component accumulates; it does not establish what reaction happens there. Comparing locations with the consequences of changing particular processing steps tests a mechanistic explanation. Nor does an accumulation of decay intermediates automatically mean that decay has become faster: material can accumulate when an exit step is blocked. This connects the chemistry of messenger RNA processing to the organization and dynamics of the cytoplasm.

Roy Parker, portrait on his University of Colorado Boulder BioFrontiers profile
. - Decapping and Decay of Messenger RNA Occur in Cytoplasmic Processing Bodies by Ujwal Sheth and Roy Parker
- Processing bodies require RNA for assembly and contain nontranslating mRNAs by Daniela Teixeira and others
- Recognition of the TACTAAC box during mRNA splicing in yeast involves base pairing to the U2-like snRNA by Roy Parker and others
Umesh Varshney is a molecular biologist whose research investigates mechanisms of biological translation and DNA repair. His Indian Institute of Science profile identifies studies of protein synthesis and DNA repair using bacterial model systems, and lists him as an honorary professor in its Department of Microbiology and Cell Biology.[ref]
Umesh Varshney research profile at the Indian Institute of Science. Profile heading and Research section.
A 1991 collaborative paper provides a specific contribution to understanding transfer RNA identity. Varshney and colleagues altered an initiator transfer RNA in Escherichia coli and tested its function in both initiation and chain elongation. Internal amber stop codon suppression measured elongator activity; one variant could also initiate at an upstream UAG and suppress a downstream UAG. The result showed that the two activities can coexist in an engineered carrier.[ref]
Mutants of initiator tRNA that function both as initiators and elongators by Umesh Varshney and others. Abstract and pp.18018–18024: acceptor-stem variants and the dual-function test.
This distinguishes two questions that an amino-acid label alone cannot answer: what chemical cargo a transfer RNA carries, and which step of biological translation recognizes that carrier. Testing the same RNA at an initiating position and an internal codon probes a functional distinction, rather than merely comparing two RNA sequences.
His departmental research account also describes ribosome recycling and the recognition features that distinguish initiator transfer RNA from elongator transfer RNA.[ref]
Umesh Varshney research profile at the Indian Institute of Science. Protein Synthesis section, ribosome recycling and initiator-tRNA structure–function work.
These connect molecular recognition to the continued use of the translation machinery, beyond delivery of a single amino acid.
Sebastian M. Fica is a molecular biologist studying how the spliceosome carries out RNA splicing. His work connects biochemical tests of chemical mechanism with structural analysis of the RNA–protein machine. His Oxford laboratory biography records a doctorate in cell and molecular biology in 2013 and a move into structural postdoctoral work in 2014.[ref]
Sebastian M. Fica laboratory biography at the University of Oxford. “Short biography”: doctoral work on splicing catalysis and subsequent training in electron cryo-microscopy.
In a 2013 collaborative study, Fica and colleagues used substitutions in U6 small nuclear RNA and metal-rescue assays in assembled yeast spliceosomes. Their results connected RNA ligands to catalytic metals in both splicing reactions. The paper marks equal contributions by its first two authors; the result belongs to the collaborating team.[ref]
The scientific question is sharper than asking whether RNA is present near a reaction. An RNA component might hold other molecules in place, or its own chemical groups might help build the catalytic center. Perturbing a candidate group and testing a compensating chemical change links structure to function. Structural snapshots complement such tests by locating molecular contacts, while reaction assays ask whether those contacts matter for catalysis.
Oxford's departmental profile describes his group's combination of biochemical and structural approaches to splice-site selection and catalytic dynamics.[ref]
https://www.bioch.ox.ac.uk/node/1889921. Departmental research profile, “Mechanisms and molecular structure of the spliceosome during pre-mRNA splicing”

Sebastian M. Fica, portrait identified on his University of Oxford departmental profile
. Researcher in plant gene silencing. Herr and collaborators investigated RNA polymerase IV in silencing endogenous DNA, connecting a specialized transcription enzyme with regulation of genomic sequences.[ref]
RNA polymerase IV directs silencing of endogenous DNA by A. J. Herr and others. Title, author list and reported study
Researcher in small-RNA biology. Malone and collaborators investigated specialized piRNA pathways in germline and somatic tissues of the Drosophila ovary, connecting tissue context with genome defense.[ref]
Researcher in nucleic acid-guided interference. Swarts and collaborators investigated DNA-guided DNA interference by a prokaryotic Argonaute, connecting sequence recognition by a guide with selective action on a target.[ref]
DNA-guided DNA interference by a prokaryotic Argonaute by Daan C. Swarts and others. Title, author list and reported study
Researcher in epigenetics. Zilberman and collaborators investigated ARGONAUTE4 control of small-RNA accumulation and DNA and histone methylation, connecting RNA-guided targeting with localized chromatin regulation.[ref]
Researcher in epigenetic inheritance. Ashe and collaborators investigated multigenerational memory triggered by piRNAs in the Caenorhabditis elegans germline, connecting a small-RNA signal with a persistent inherited state.[ref]
Researcher in small-RNA regulation. Watanabe and collaborators investigated endogenous siRNAs derived from double-stranded RNA in mouse oocytes, connecting naturally paired transcripts with post-transcriptional control.[ref]
Researcher in genome defense. Aravin and collaborators investigated a piRNA pathway linked to new DNA methylation in mice, connecting individual transposable elements with small-RNA-guided genomic regulation.[ref]
Researcher in piRNA biology. Brennecke and collaborators investigated inherited small RNAs and transposon silencing, connecting a maternal molecular contribution with genome defense in offspring.[ref]
Researcher in developmental gene regulation. Lee and collaborators identified small RNAs encoded by lin-4 microRNA with complementarity to lin-14, establishing a molecular basis for a new form of developmental regulation.[ref]
Researcher in gene silencing. Fire and collaborators demonstrated potent sequence-specific interference by double-stranded RNA in Caenorhabditis elegans, providing an experimental foundation for RNA interference.[ref]
Researcher in anaerobic cell biology. With Donald G. Lindmark, Müller characterized the hydrogenosome and its role in pyruvate metabolism, connecting an unusual organelle with energy production without oxygen.[ref]
Researcher in RNA processing. With Claire L. Moore, Sharp investigated accurate cleavage and polyadenylation of an experimentally supplied RNA substrate, connecting sequence recognition with formation of a mature transcript end.[ref]
Researcher in enzyme evolution. Orencia and collaborators combined directed evolution with structural analysis to investigate the emergence of antibiotic resistance, connecting sequence changes with altered enzyme behavior.[ref]
Researcher in enzyme structural biology. Langan and collaborators used neutron and X-ray crystallography to investigate a conserved glutamate in beta-lactamase, connecting atomic positions with a proposed catalytic role.[ref]
Researcher in organelle biology. Mi-ichi and collaborators identified a sulfate-activation pathway in Entamoeba histolytica mitosomes, connecting a reduced organelle with a retained metabolic function.[ref]
Mitosomes in Entamoeba histolytica contain a sulfate activation pathway by Fumika Mi-ichi and others. Title, author list and reported study
Researcher in reduced mitochondrial organelles. Tovar and collaborators investigated iron–sulfur protein maturation in Giardia organelles, connecting a mitochondrial remnant with a specialized cellular requirement.[ref]
Researcher in anaerobic metabolism. With Miklós Müller, Lindmark described the hydrogenosome and its involvement in pyruvate metabolism, helping explain energy conversion in organisms with unusual cellular organization.[ref]
Researcher in live-cell imaging. Sakaue-Sawano and collaborators developed fluorescent visualization of multicellular cell cycle progression, connecting changing reporter signals with the timing and location of cell-cycle states.[ref]
Researcher in mitochondrial dynamics. Smirnova and collaborators investigated the requirement for Drp1 in mammalian mitochondrial division, connecting a dynamin-related protein with changes in organelle shape and number.[ref]
Researcher in mitochondrial dynamics. Chen and collaborators investigated coordinated action of Mfn1 and Mfn2 in mitochondrial fission-fusion cycle, connecting organelle remodeling with embryonic development.[ref]
Researcher in RNA processing. Zarkower and collaborators investigated the requirement for the AAUAAA signal sequence in transcript cleavage and polyadenylation, connecting a short RNA signal with formation of a mature transcript end.[ref]
Researcher in RNA processing. With Phillip A. Sharp, Moore reconstructed accurate cleavage and polyadenylation of an external RNA substrate, helping make transcript-end formation experimentally accessible.[ref]
Researcher in structural biology. Campbell and collaborators determined the structure of a bacterial RNA-polymerase sigma factor, connecting molecular architecture with selective promoter recognition.[ref]
Researcher in chloroplast gene regulation. With Prakitchai Chotewutmontri, Barkan investigated how photosystem damage triggers psbA messenger RNA translation through an assembly-linked circuit, connecting protein production with repair demand.[ref]
Researcher in translational control. With Krishna M. Vattem, Wek investigated ATF4 regulation through translation reinitiation at upstream reading frames, connecting transcript architecture with a cellular stress response.[ref]
Researcher in developmental RNA localization. With Kevin M. Forrest, Gavis investigated diffusion and entrapment of nanos messenger RNA in Drosophila, distinguishing selective retention from directed transport along an entire route.[ref]
Researcher in cellular stress signaling. Inglis and collaborators investigated activation of GCN2 by the ribosomal P-stalk, connecting the translation machinery with a regulator that responds to amino acid availability.[ref]
Activation of GCN2 by the ribosomal P-stalk by Alison J. Inglis and others. Title, author list and reported study
Researcher in epigenetic inheritance. Ragunathan and collaborators experimentally separated inheritance of a chromatin state from sequence-specific recruitment, testing whether maintenance can persist after the initiating targeting signal changes.[ref]
Epigenetic inheritance uncoupled from sequence-specific recruitment by Kaushik Ragunathan and others. Title, author list and reported study
Articles were limited to the first 100 out of 280 total.
Articles by others on the same topic
There are currently no matching articles.