Treat the membrane as a capacitor, and count the additional positive charge on its inner surface. With radius m and length m, including both end caps givesThe membrane capacitance is pF. Thus C, corresponding to monovalent sodium ions. Converting charge to amount of substance using the Faraday constant,Omitting the ends changes the answer to mM, a one-percent correction. A large membrane potential change therefore requires only about of the intracellular sodium population to enter. The bulk liquid remains nearly electrically neutral; excess charge resides chiefly near the cell membrane. This explains why an action potential need not exhaust the concentration difference that drives it, and why fixed reversal potentials are a useful approximation during one spike. This membrane capacitive ion budget assumes sodium supplies the minimum net charging requirement. Simultaneous opposing electric currents can require more sodium entry; repeated spikes eventually require the sodium-potassium ATPase to maintain the gradients.
A typical voltage-gated potassium channel has four subunits, each with six membrane-spanning segments. S1–S4 form a voltage-sensing domain; S5–S6 and their intervening loop form the pore and selectivity filter. Conserved positively charged side chains along S4 move through part of the membrane electric field when the voltage changes. Countercharges and hydrated crevices in the surrounding sensor help make this movement possible. Coupling between the sensor and pore, including the S4–S5 region and S6 gate, changes the probability of opening. A voltage-gated sodium channel usually combines four related domains in one large protein, rather than four identical independent proteins.
The evidence separates charge movement, structural arrangement and functional coupling. A transient ion-channel gating current can persist when permeating ionic electric current is suppressed: some charged parts of the channel itself move. Comparing integrated ion-channel gating charge with channel opening tests their coupling. Charge-neutralization experiments change both total moved charge and voltage dependence, implicating S4 basic residues and countercharges elsewhere; they do not show that S4 alone supplies every charge. [ref]
Voltage-sensing residues in the S2 and S4 segments of the Shaker K+ channel by S. A. Seoh and others. 1996 primary abstract: neutralization identifies contributions from S2 and S4
Crystal structure of a mammalian voltage-dependent Shaker family K+ channel by Stephen B. Long and others supplies structural evidence for the arrangement. Voltage-dependent accessibility and optical reports of sensor motion provide complementary functional tests. No one static structure is a movie of gating, and a perturbation that abolishes current can disrupt folding or pore function rather than selectively remove voltage sensing. Agreement between structural, electrical and motion-sensitive measurements supports the mechanism more strongly than any one observation alone.
Write the instantaneous electrical conductance as . Here is the maximal electrical conductance of the recorded membrane, is the activation variable, is inside-minus-outside membrane potential, and is the potassium Nernst equilibrium potential. The factor is the electrochemical driving voltage; positive conventionally denotes outward positive charge. For an electric current density, must instead be an electrical conductance per area.
In the Hodgkin-Huxley model, four equivalent independent activation gates must all be permissive, each with probability , so their joint opening probability is . At fixed voltage each gate obeysThe model assumes approximately Ohmic open-channel conduction, a well-defined population-average electrical conductance, and an almost constant potassium reversal potential during a brief recording. Four phenomenological gates should not be mistaken for a proof of four physically independent molecular transitions. The simple delayed-rectifier description also omits ion-channel inactivation.
Reducing the available electrical conductance can weaken repolarization, broaden an action potential and alter repetitive firing. Depending on channel class and tissue, excess excitability or impaired recovery can follow; persistent depolarization can also reduce voltage-gated sodium channel availability. Cardiac potassium-current reductions can prolong the action potential and contribute to Long QT syndrome. These are conditional physiological consequences, not a claim that every potassium-current reduction has the same clinical effect.
The average electric current is the number of channels times their ion-channel open probability times the electric current carried by an open channel:Consequently the identifiable answer isTODO (missing input): The source does not supply or the potassium thermodynamic activities for this experiment. A unique numerical electrical conductance cannot follow from the listed electric current, count and ion-channel open probability alone. For the explicitly additional choice mV, pS. Taking mV instead gives pS: both match the measured electric current. The table in question 3 describes a skeletal-muscle resting state, and cannot silently supply the conditions of this separate recording.
Rapid permeation is possible because the hydrated pore and coordinating groups of the selectivity filter provide a favorable free-energy landscape for repeated ion transit. Several ions can occupy the filter and their interactions favor onward passage, while the channel remains open for many successive transits. Selectivity need not mean that each transported ion remains tightly trapped for a long time. This is passive downhill transport; it does not consume one ATP molecule per ion crossing.
A useful organizing principle is Gibbs free energy: at specified temperature and pressure a coupled process is favorable when its total is negative. For each chemical species,The relative amounts of chemical substances determine the available Gibbs free energy, beyond counting energy in individual bonds. Nutrient oxidation and light-driven chemistry can maintain nonequilibrium compositions. ATP hydrolysis can drive biosynthesis, transport and motion of molecular motors when reactions are coupled so that their total free-energy change is favorable. Breaking an inorganic phosphate bond itself costs energy; the favorable net hydrolysis includes formation and solvation of the products and depends on their thermodynamic activities.
Redox Gibbs free energy comes from transferring electrons between donors and acceptors: . Electron transfer in respiration or photosynthesis can help establish an electrochemical gradient. For a mole of ions moving from side 1 to side 2,The concentration and electrical terms together drive transport and can power ATP synthase. ATP-driven pumps can conversely build gradients. Chemiosmosis is the coupling between this gradient and chemical synthesis, not the creation of Gibbs free energy by an enzyme.
Mechanical work and elastic storage are other forms: a motor can do or , and an approximately Hookean element stores . ATP-driven motors convert Gibbs free energy into motion, while externally forced reverse motion can sometimes drive chemical synthesis. A photon brings energy ; light photon absorption initiates chemistry in photosynthesis and light-driven pumps, with some energy inevitably dissipated. Living cells use these conversions for structure, movement, homeostasis and synthesis, while releasing heat and exporting entropy. At a single uniform temperature, dissipated heat cannot be completely recycled into useful work.
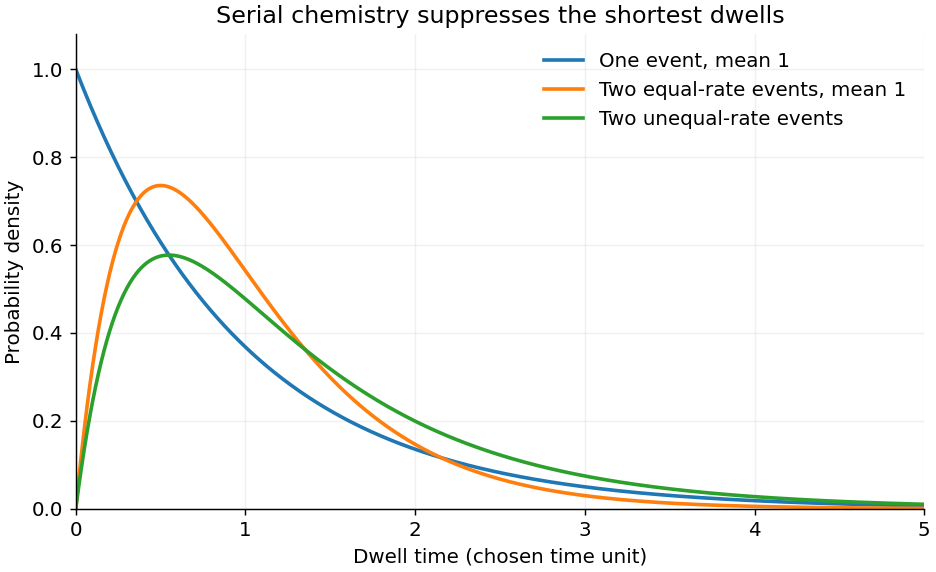
At saturating ATP, binding need no longer dominate the observed dwell, but saturation alone does not determine a dwell-time distribution. The standard interpretation of a distribution that rises from zero is that at least two sequential events remain rate limiting. Model a dwell as , where the independent waiting times are distributed according to exponential distributions with rates . Their convolution isThis hypoexponential distribution starts at zero, rises to a mode , then has an exponential tail. When the rates are equal, the continuous limit is the two-stage Erlang distribution,A single slow memoryless event instead gives , maximal at zero. Three 120-degree steps per revolution specify rotational stoichiometry, not three slow reactions within every dwell. The two-event interpretation is supported by the resolved catalytic-dwell chemical kinetics in the primary F1 experiment.[ref]
Resolution of distinct rotational substeps by submillisecond kinetic analysis of F1-ATPase by Ryohei Yasuda and others. 2001, abstract and Figure 8: two reactions occupy the catalytic dwell
The fit assumes sequential approximately irreversible events, constant rates during each dwell, negligible ATP-binding delay, negligible mechanical transit time, and detection of all relevant steps. Parallel pathways, backward transitions, inhibited states, probe drag and missed short dwells can change the histogram. The mathematical fit alone does not identify each event chemically or prove that two ATP molecules are hydrolyzed per step.

Let and use a one-molar standard state, so the inorganic phosphate activity is approximately . With the source's stated standard hydrolysis value,The downhill proton-transfer Gibbs free energy is for the signed driving force V. Ideal reversible balance of one complete turn requiresSolving yieldsAt 298.15 K the estimate is . Temperature is not specified in this paragraph; the first value explicitly adopts physiological temperature rather than pretending it is supplied. The unexpectedly large ADP/ATP ratio follows from using the printed standard value kJ/mol with the stated dilute inorganic phosphate. One must not silently replace that value by another convention or treat it as the already concentration-corrected cellular hydrolysis Gibbs free energy.
This estimate assumes ideal thermodynamic activities, consistent pH and magnesium conventions, tight coupling of ten protons to three ATP, no leak or transport overhead, and operation close to reversible balance. A working cell is not in global equilibrium. Net synthesis at this fixed driving force requires , hence above this reversible-balance threshold; finite chemical kinetics and dissipation prevent the stoichiometry alone from determining an exact physiological ratio.
Use signed ion valence and define . The molar electrochemical potential in side of an ideal dilute solution isTransfer of an infinitesimal amount from side 1 to side 2 changes the total Gibbs free energy by . At equilibrium this derivative is zero, soFor a cell with side 1 inside, this is . The concentrations are equilibrium free-ion concentrations, not automatically their exact initial values. For nonideal solutions the ratio must be a ratio of thermodynamic activities. The temperature and solvent must be common to both reservoirs, and an ion must actually be able to exchange for this equilibrium condition to be dynamically established.
Equal compartment volumes are unnecessary for deriving the voltage law, but help specify number conservation if concentration changes are appreciable. For equal volume and a monovalent cation transfer from side 1 to 2, , . Combined with the membrane capacitance, this gives an implicit finite-reservoir equilibrium rather than treating both concentrations as externally clamped.
Suppose initially . Positive ions initially move by diffusion from side 1 to side 2. Since negative counterions cannot cross, side 1 acquires a negative interfacial charge and side 2 a positive one. The resulting electrical force opposes further positive-ion transfer. The two contributions balance when the electrochemical potentials are equal; individual ions still cross in both directions, but there is no net electric current of the permeant chemical species.
A simple capacitor model makes the feedback quantitative. If moles of monovalent positive ions move from initially neutral side 1 to side 2, the charge on side 1 is , hence . The free-energy stationarity condition isAs increases, both the concentration imbalance and the voltage drive toward reversal diminish the original diffusion tendency. The equilibrium lies before the bulk concentrations become equal. This demonstrates why a nonzero Nernst potential can coexist with very little change in bulk composition and approximate bulk electroneutrality. A concentration difference alone does not maintain an electrical voltage if both counterions can freely follow each other; selective permeability is essential to this model.
At K, mV. Always retain the valence sign. For the two monovalent positive ions,For calcium, and the specified voltage is positive:For chloride, reverses the sign in the Nernst equation:These values use thermodynamic activities approximated by the given concentrations. Calcium is the free concentration relevant to its electrochemical potential, rather than total bound plus free intracellular calcium. Ratios use matching concentration units, so a ratio of millimolar values is already dimensionless. Failure to include the factor of two for calcium or the negative charge of chloride would change the answer substantially.
At a common membrane voltage, an ion is in electrochemical equilibrium only if its own Nernst potential equals that voltage. The four independently calculated equilibrium voltages areThus at mV only chloride is at its listed equilibrium voltage. Potassium has an outward positive-current driving voltage mV. Sodium and calcium have negative driving voltages, and mV respectively, favoring inward positive-ion movement when their channels are open. Chloride has zero net electrochemical drive under these ideal conditions.
A resting potential is not a claim that every permeant chemical species is in thermodynamic equilibrium. A membrane with several electrical conductances can have zero total electric current even though individual ionic electric currents flow and cancel. Pumps maintain the concentration differences by consuming Gibbs free energy. Conversely, a closed channel can prevent substantial ion flux even when the ion's electrochemical potential difference is nonzero: absence of flux due to impermeability is not the same as electrochemical equilibrium. The listed concentrations could not all be passive equilibrium distributions at one voltage because their Nernst potentials differ.
A rising force branch means that an already exposed polypeptide segment is being stretched while the remaining structure resists further extraction. At a force peak a structural barrier is crossed; protein denaturation or release of additional chain increases the available polymer contour length, so the force falls at almost the same imposed separation. Stretching then loads the next remaining barrier. The sawtooth pattern therefore reports a sequence of mechanically resistant intermediates, not a regular spring oscillation. The cantilever deflection gives force, while its own deformation must be removed from the imposed separation to infer molecular extension.
A membrane protein also requires work to disrupt membrane anchoring, helix packing and contacts with lipids or neighboring molecules. The retinal-containing interior contributes a bulky, specifically packed scaffold; exposing chain must overcome these steric hindrance as well as hydrogen bond and electrostatic interactions. A force peak need not correspond one-to-one to one complete helix, and its height depends on loading rate and pathway, not just an equilibrium folding Gibbs free energy.
Salt changes electrostatic screening and specific ion binding. Screening can reduce either repulsion or attraction, so “more salt always weakens the protein” is false. The primary study underlying these kinds of traces identifies relatively salt-insensitive steric hindrance around the retinal and additional salt-sensitive barriers. Its interpretation associates some added barriers with screening of destabilizing electrostatic repulsion. The supplied traces likewise show a changed set of intermediate peaks, rather than a uniform rescaling of one curve.[ref]
https://pmc.ncbi.nlm.nih.gov/articles/PMC1959538/. 2007 primary study, Results: Unfolding bR from PM cytoplasmic side; Discussion: steric and electrostatic contributions
Exact assignment of every arrow to a residue requires structural and statistical information beyond one representative trace. No such assignment is inferred here. The printed reference to titin in the following paragraph is inconsistent with the stated specimen and the membrane-protein traces; the physical interpretation here concerns bacteriorhodopsin, while the worm-like-chain expression is a generic polymer approximation.
On each branch the worm-like chain force-extension interpolation relates force to the extension of the already released chain. Fitting a sufficiently long branch estimates its polymer contour length and, if the data constrain it independently, its persistence length . In practice may be fixed across branches so that the changes in are better identified. Consecutive fitted polymer contour lengths reveal how much additional chain became available at an protein denaturation event; using a justified polymer contour length per residue can then estimate residue counts. A projected folded length and the pulling geometry must be included before assigning a structural boundary.
Writing , expansion at small extension givesNear full extension, . Larger shifts a branch toward larger extension at fixed force. The effective flexural rigidity is , with units of force times length squared. This is entropic straightening of a worm-like chain; bond stretching and barrier rupture themselves are not included in the formula.
The persistence-length wording requires care. The three-dimensional model gives . At , . This is not a statement that . For example, angles zero and with equal probabilities have mean angle , but . The distinction prevents a false literal interpretation of the printed definition.
The earliest branch is close to the membrane and the point of tip contact. Little flexible chain has yet been exposed, so a long homogeneous continuum chain with purely entropic elasticity is a poor description. Tip–surface interactions, membrane deformation, steric obstruction, and extraction of tightly anchored structure can contribute strongly to the measured force. The retinal-associated structure and local electrostatic contacts can still be mechanically active at these small separations.
The worm-like-chain formula assumes that the measured extension chiefly stretches an already available flexible segment with an identifiable polymer contour length. It does not model detachment of the tip from a surface or the energetic rupture of a short structured segment. That explains why fitting these first peaks is unreliable even if later branches admit reasonable fits. A single trace cannot distinguish uniquely among these short-range effects; claiming that one particular arrow proves a specific retinal bond rupture would exceed the available evidence.
The main generic costs of strongly confining a charged semiflexible polymer are bending it away from its preferred straight shape, crowding neighboring segments, and reducing its available configurations. Closely approaching charged strands also repel through electrostatic and hydration interactions; mobile ions screen or modify these interactions, while counterion organization contributes to the free-energy balance. The bending cost is especially large in regions of high curvature. A pressure on the confining wall and a force resisting further insertion are related thermodynamic responses, but are not identical quantities with the same units.
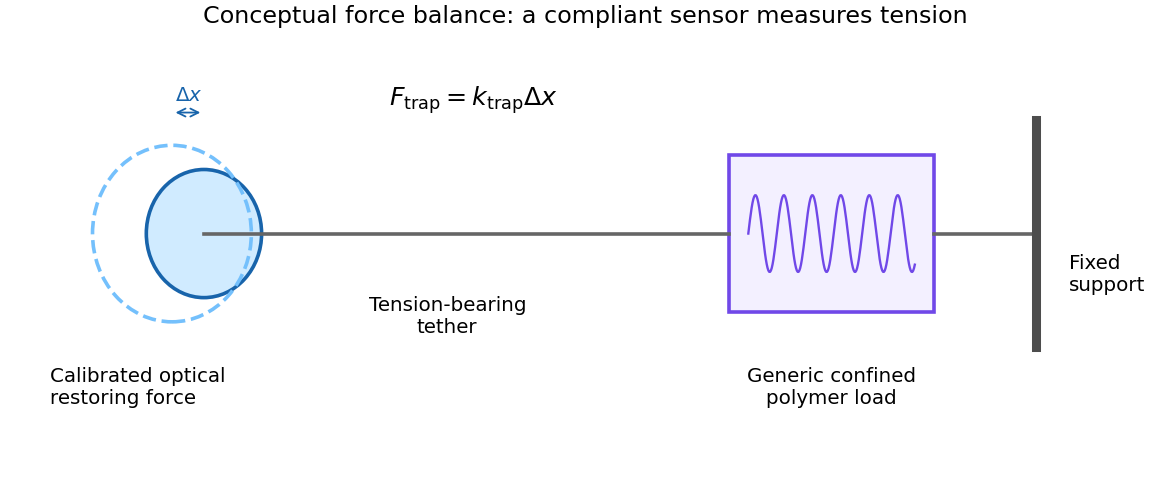
At the level of measurement principles, an optical tweezers apparatus acts as a calibrated compliant force sensor. A displaced trapped bead experiences approximately . A tether transmits tension between that sensor and the material under load; the opposing support fixes the other end. The conceptual force balance isThe extension and restoring force supply a mechanical record of the load. This is a force-sensing schematic, not a biological preparation or packaging protocol. The measured tension can include bending, interstrand interactions, dissipation and any other mechanically coupled contribution; it cannot be identified with only the bending term without additional assumptions. In a reversible generic model at specified geometry, while an internal pressure is at specified polymer content. These derivatives clarify what a force measurement can and cannot determine.

Use the elastic-rod bending functional for an abstract inextensible worm-like chain,A circular ring of radius has curvature and length , hence energy . With rings of each radius, add their energies:For a cylindrical annulus of height , choose neighboring rings separated vertically by . Then , while adjacent radial layers are separated by . Replacing the radial sum by its continuum integral givesandEliminate the inner radius to obtainThe dimensions are energy, and the expression is positive for . It tends to zero with and diverges as the inner hole closes in this continuum model. The divergence signals the growing cost of forcing an inextensible chain to zero radius; it is not a physical prediction at arbitrarily small molecular scales. Interstrand, hydration and configurational contributions have been excluded, and disconnected ideal rings represent local curvature without modeling the connections between them.
Use the given spherical-shell force balance directly; nm is the loaded radius. The radial enlargement isSince Pa, the pressure is . ThereforeThe diameter increases by nm, and nm. The relative radial or diametral enlargement is about , only . Under this stipulated shell model the container is therefore very stiff despite the substantial internal pressure.
The factor of two follows from the particular elastic energy convention implicit in the question: differentiating gives . Replacing it by the derivative of would change the answer incorrectly. The calculation treats the wall by an effective spherical radius; local deformation or discrete facets of an icosahedral shell are not resolved by this one-coordinate model.
A Jablonski diagram distinguishes electronic excitation from subsequent relaxation. Photon absorption takes a molecule from the electronic ground state to a vibrationally excited level of or a higher singlet state. Rapid vibrational relaxation and internal conversion usually populate the lowest vibrational levels of . Fluorescence then emits a photon on a transition to , often ending in one of its excited vibrational levels. Nonradiative relaxation competes with emission; intersystem crossing can also populate triplet states. Energy lost to relaxation and solvent reorganization commonly makes the fluorescence maximum occur at lower photon energy, hence longer wavelength, than the photon absorption maximum: the Stokes shift.
The intrinsic fluorescence quantum yield is the number of photons emitted by fluorescence per absorbed excitation. For a simple excited state with radiative rate and other decay rate ,A nearby acceptor adds a nonradiative energy-transfer channel, not an emitted photon flying from donor to acceptor. The electric dipole-dipole interaction transfers the donor excitation while returning the donor toward its ground state and exciting the acceptor. The acceptor then relaxes and may fluoresce. The donor yield becomesFor the acceptor, distinguish its intrinsic yield per acceptor excitation, , from the number of acceptor photons per donor excitation. Transfer increases the supply of acceptor excitations, giving sensitized emission per donor excitation in the ideal model. It does not by itself change if the acceptor's own decay rates remain unchanged. Environmental changes, saturation or additional quenching can change those rates, but increased acceptor intensity alone does not establish an increased intrinsic fluorescence quantum yield.
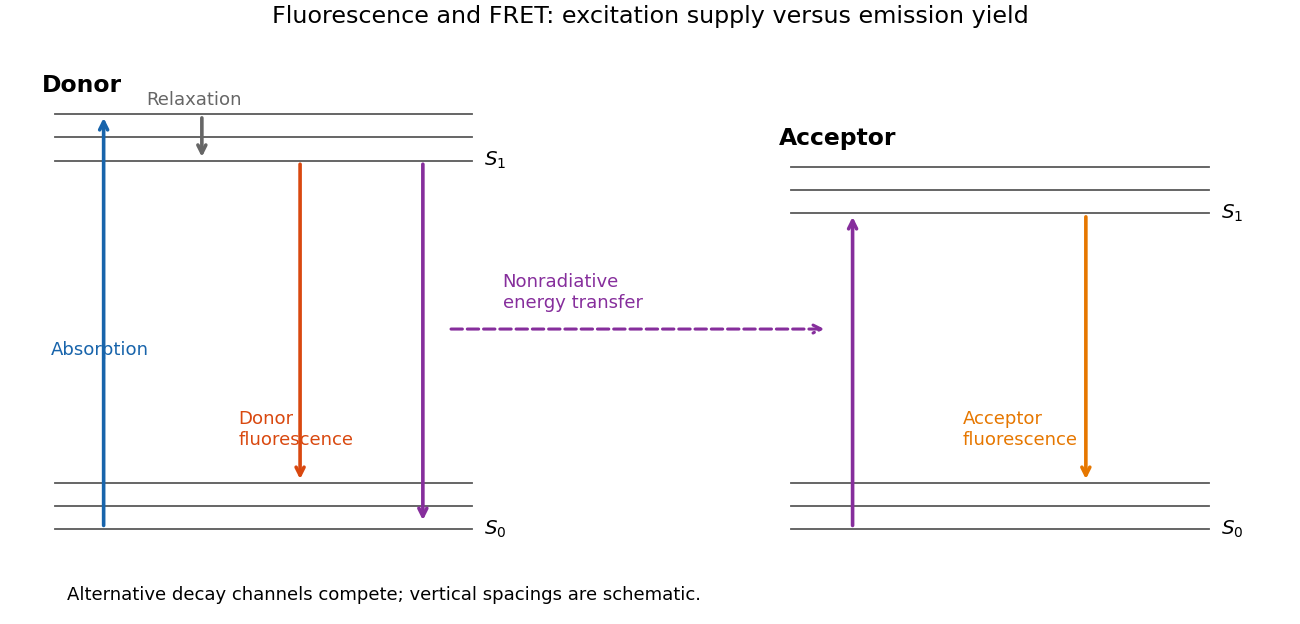
Relaxation, fluorescence and donor–acceptor energy transfer
. Original schematic electronic and vibrational levels. Transfer supplies an acceptor excitation without a photon between the molecules; the acceptor quantum yield and the excitation supply are separate quantities. Details.
Let , so the lifetime without transfer is . For a fixed isolated donor–acceptor pair, the specified transfer rate is . Inserting this rate into the definition of FRET efficiency givesThe donor population in the presence of the acceptor obeys , so its fluorescence decays exponentially with lifetime . ThereforeBoth relations assume the donor's nontransfer rates are unchanged by adding the acceptor, a single approximately fixed separation during each excited-state lifetime, and an available acceptor. The transfer law also presumes the usual weak-coupling dipole approximation and the spectral and orientation conditions contained in . If the reference donor is differently quenched, the lifetime difference cannot all be attributed to transfer. Heterogeneous mixtures or a fluctuating separation can produce multiexponential decay, in which case blindly substituting an arbitrary fitted average lifetime into the single-pair formula need not give the ensemble's mean efficiency.
For independent available acceptors at equal distance, the multiple-acceptor FRET efficiency follows by adding the parallel exit rates. Each contributes the same rate , so the total rate is , rather than times the single-pair transfer probability. HenceEquivalently, . Inverting givesWith and , the bracket is exactly one and nm. Each isolated pair would have efficiency ; the three transfer rates together give . Multiplying by three would violate the probability bound and is not a valid calculation.
This model assumes one donor excitation, identical pairwise transfer rates, no coherent coupling between acceptors, no acceptor saturation and no change in donor photophysics or orientation on adding labels. Different separations instead require summing individually. Acceptor-to-acceptor transfer can alter which acceptor emits; it does not automatically change the donor's first-transfer probability when the donor exit rates remain those specified.
Assume distinct positive separations, and first label them so . Let , and . The positive contrast isIts derivative has a particularly simple sign:It is positive below and negative above, so this is the global maximum. Both end limits and give zero contrast. ThusTo recover the printed expression, factor its sixth-power radicand:Its sixth root is indeed . The cancellation requires . If , every choice gives zero contrast and there is no unique optimum; the printed expression then has a removable algebraic without a unique maximization conclusion. For unordered , maximize absolute contrast and replace the numerator of the maximum by . A negative signed difference for is minimized, not maximized, by the same interior point. These qualifications are needed to make the statement precise.
The bacterium samples its surroundings in time. A sustained increase in chemoattractant makes a favorable run persist by reducing the chance of tumbling; declining chemoattractant makes reorientation more likely. Individual tumbles need not point up the gradient. Biasing run durations can generate net migration even if reorientation itself is random. This is temporal comparison, not a direct measurement of a concentration gradient across the short cell body.
Compare the swimming time over length , , with diffusion over the same scale, . Their ratio is the Péclet number . The crossover estimate isOrder-one factors depend on the chosen diffusion-distance convention. A hypothetical stirrer of extent must instead have of order one, givingThese are transport-scale estimates, not a proof of a particular increase in uptake. Around a one-micrometre body, : local diffusive supply is already rapid. Actual uptake enhancement depends on the nutrient field and the cell's uptake boundary condition, neither supplied. The long run can help the organism reach a better nutrient region without implying that it greatly increases the flux from a uniform bath. E. coli normally uses bacterial flagella; the cilium in this comparison is a hypothetical generic stirring appendage, not its actual locomotor organelle.
Write , and . Saturated receptor methylation provides a constant positive flux, so the complete balance isAt a finite steady state, and thereforeThe derivative of the right side is , so this fixed point is stable in the scalar model. With total receptor concentration , it must additionally lie between zero and and leave sufficient substrate for the assumed saturated receptor methylation. If , the formal equation has no finite positive fixed point: finite receptor capacity then matters.
For an chemoattractant step, activity and hence tumbling frequency fall rapidly before receptor methylation can change appreciably. Slower chemical modification can then restore receptor signaling and make the tumbling frequency rise toward a new steady value. Perfect adaptation means return of that output to its original baseline despite the persistent chemoattractant step; it does not mean that receptor receptor methylation also returns to its original level.
The balance written here alone does not specify how ligand changes activity or receptor demethylation, and does not yet establish perfect adaptation. If activity is simply multiplied by a ligand-dependent fraction while this balance remains unchanged, is unchanged and the activity remains reduced. The following two models add different mechanisms that can compensate for that direct activity reduction.
Let chemoattractant reduce the activity per methylated receptor by a factor , with , taking the initial factor as one. The active output is . If changes to but receptor demethylation still acts on total methylated receptor, the new output isDemanding givesReduced receptor demethylation lets methylated receptors accumulate to , compensating exactly for their reduced activity. During the transition the activity first falls, then rises as receptor methylation accumulates. This compensation works only if the required methylated concentration is accessible; in particular in a finite receptor pool.
The required relation contains both enzyme concentrations. At fixed rates, increasing CheR raises the baseline receptor methylation and tumbling output, while increasing CheB lowers them. After such a change, the previous fine tuning generally no longer gives perfect adaptation. For example the required reverse capacity is , so changing without retuning changes the adapted output ratio. It is the precision of adaptation that is fragile here, in addition to the baseline itself changing. Changing all relevant capacities together in a way preserving the relation is a special exception, not generic robustness.
Separate total methylated receptor from its active portion , where ligand-bound receptor is inactive and the unbound fraction supplies at finite ligand concentration. In the Barkai-Leibler model, receptor demethylation depends on the active portion:Every accessible stable steady state therefore obeysThe output is independent of chemoattractant even though the methylated pool changes. A ligand step first lowers , reducing its own reverse modification. Constant receptor methylation continues until recovers. Local stability follows from . This is output-dependent negative feedback. [ref]
Robustness in simple biochemical networks by Naama Barkai and Stanislas Leibler. 1997, model mechanism and adaptation precision
Changing CheR or CheB changes and the recovery time, but the return to each new pre-step baseline remains exact over the model's operating range. Robust perfect adaptation does not mean invariant tumbling frequency under changes in enzyme abundance. Nor does it extend to , or saturation of the finite receptor methylation capacity. If every receptor is permanently inactive, and the reverse flux cannot restore a positive baseline. The literal phrase that binding reduces activity to zero applies to bound receptors, not to every receptor under every finite stimulus.
If CheR instead follows Michaelis-Menten kinetics on unmethylated substrate, , thenAt steady state the forward flux becomes and generally depends on ligand through . Thus ligand-independent output is lost. Near-perfect adaptation can remain when so that CheR is effectively saturated, but Michaelis–Menten behavior outside that limit does not guarantee perfect adaptation. [ref]
Robust perfect adaptation in bacterial chemotaxis through integral feedback control by Tau-Mu Yi and others. 2000, analysis of ligand-independent bound CheR and the saturation assumption
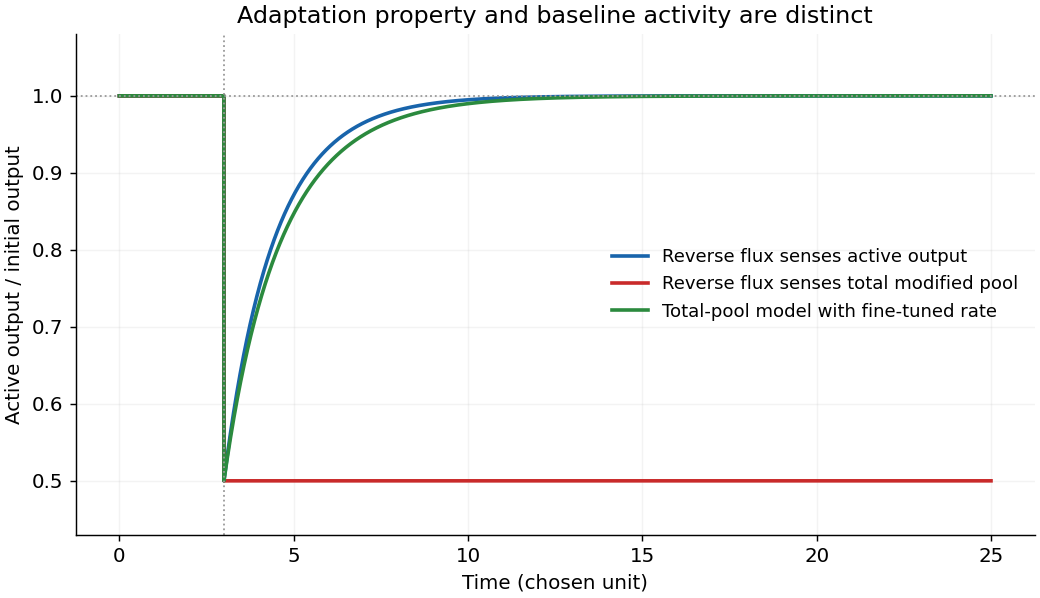
Fine tuning and output-dependent adaptation
. Original numerical solutions of the stated mathematical models. The ligand activity factor changes from one to one half; saturated output-dependent feedback restores activity, while a total-pool reverse reaction requires fine tuning. Details.
The central prediction of the ATP synthase ATP synthase rotary mechanism is relative motion of the central shaft and catalytic head, not simply a change in bulk ATP concentration. In the foundational single-molecule experiment, a visible marker coupled to the central gamma shaft reported its angular position relative to the immobilized catalytic head. Sustained directional rotations during ATP hydrolysis demonstrated rotary motion directly. The catalytic head contains three beta sites within an alpha-three beta-three assembly; their changing nucleotide affinities motivate the binding-change mechanism. [ref]
Direct observation of the rotation of F1-ATPase by Hiroyuki Noji and others. 1997 primary study, direct shaft-marker rotation
An angular trajectory gives more information than an average rotational speed. At low ATP, pauses before roughly 120-degree advances become longer as ATP falls, supporting an ATP-binding-limited wait. Reducing marker drag and improving temporal resolution reveals substeps and ATP-independent catalytic dwells. The early high-speed study resolved approximately 90-degree and 30-degree substeps, with more than one slow event during the catalytic dwell. Numerical angles vary with preparation and operational definition, so these observations should not be imposed as exact universal geometric increments.[ref]
The dwell-time model in b tests serial chemical kinetics; the angle histogram tests preferred positions. Comparing probe loads checks whether viscous drag limits the observed speed. Measuring torque and angular displacement estimates mechanical work, while a separate hydrolysis readout tests chemical coupling. With one ATP per 120-degree step, the chemical budget per revolution is three ATP, but rotation alone does not count every chemical event or establish unity efficiency.
A probe can perturb the very process it reports, and inhibited pauses or unresolved reversals can bias averages. Isolated F1 observed during hydrolysis also does not demonstrate proton-driven ATP synthesis by intact membrane F0F1. Testing reverse synthesis requires evidence for chemical gene expression under the corresponding driving conditions; a rotation movie by itself is insufficient. These distinctions connect the single-molecule motion to the free-energy balance without overinterpreting it.
Ion-channel deactivation and ion-channel inactivation are different. Ion-channel deactivation closes an activation gate when the voltage returns toward rest. Ion-channel inactivation makes a channel unavailable even while depolarization remains favorable for activation. A minimal kinetic picture is a closed state, an open state and an unavailable state, with recovery required before reopening. This helps terminate inward electric current and produces a neuronal refractory period.
In rapid voltage-gated potassium channel N-type inactivation, a cytoplasmic terminal segment, sometimes supplied by an auxiliary subunit, blocks the open inner pore. The classic ball-and-chain inactivation picture is supported by loss of rapid ion-channel inactivation after removing the relevant terminal segment and restoration by an appropriate soluble peptide. This links a mobile structural element to open-pore blockade; it does not imply that all potassium channels use that mechanism.[ref]
Restoration of inactivation in mutants of Shaker potassium channels by a peptide derived from ShB by William N. Zagotta and others. 1990 primary study, restoration of rapid inactivation
Slower C-type inactivation involves rearrangement near the selectivity filter and external pore. Dependence on pore structure, ion occupancy and extracellular ions distinguishes it from a simple cytoplasmic plugging particle. Different channels can combine several unavailable states, so a single recovery time constant need not describe every trace.
Fast ion-channel inactivation in canonical animal voltage-gated sodium channels involves the intracellular linker between domains III and IV, including its hydrophobic IFM motif, and is coupled to domain-IV voltage-sensor activation. The modern structural interpretation is an allosteric latch acting on the gate, rather than a universal literal particle sitting in the ion pathway. The older hinged-lid analogy is useful only with that qualification.[ref]
https://www.nature.com/articles/s41467-022-30400-w/. 2022 primary structural study, discussion distinguishes IFM-mediated allosteric inactivation from an observed alternative N-terminal pore block
Functionally, a voltage clamp trace shows electric current declining during a maintained step; a second test pulse measures recovery of availability. Holding-potential dependence separates an altered fraction of available channels from a mere change in maximal open electrical conductance. Persistent electric current after defective ion-channel inactivation, or delayed recovery that reduces subsequent responses, illustrates why both entry into and exit from unavailable states matter for excitability. A decaying macroscopic electric current alone does not uniquely identify the molecular mechanism.
Both motifs have a direct path and an indirect path . C1 coherent feed-forward loop uses three activating edges. I1 incoherent feed-forward loop activates and from , but lets repress . The AND condition means that the direct input must be active and the intermediate input must permit output: present for an activator, absent for a repressor. [ref]
Structure and function of the feed-forward loop network motif by Shmoolik Mangan and Uri Alon. 2003, AND-gated coherent and incoherent motifs
For a simple threshold model letThe coherent output obeys . Starting from under a sustained high input, . Provided , the gene expression delay isShorter input pulses do not turn on output gene expression. This is a persistence detector: it rejects brief positive inputs. Removing immediately removes the direct permission for gene expression, even if remains high. The concentration of subsequently decays with its own lifetime rather than vanishing instantly.
The incoherent output instead obeys . Gene expression begins immediately, then stops when accumulated repressor crosses threshold at . With zero initial output,A maintained stimulus therefore generates a pulse in this ideal Boolean model. Delayed inhibition can accelerate an early response and limit its duration. The ideal final zero output is conditional on crossing the repression threshold; graded response laws or basal gene expression can give a nonzero final level and need not adapt perfectly. If , the coherent circuit does not activate, while the incoherent circuit fails to terminate its gene expression. These boundary cases show why regulatory signs alone do not fix function.
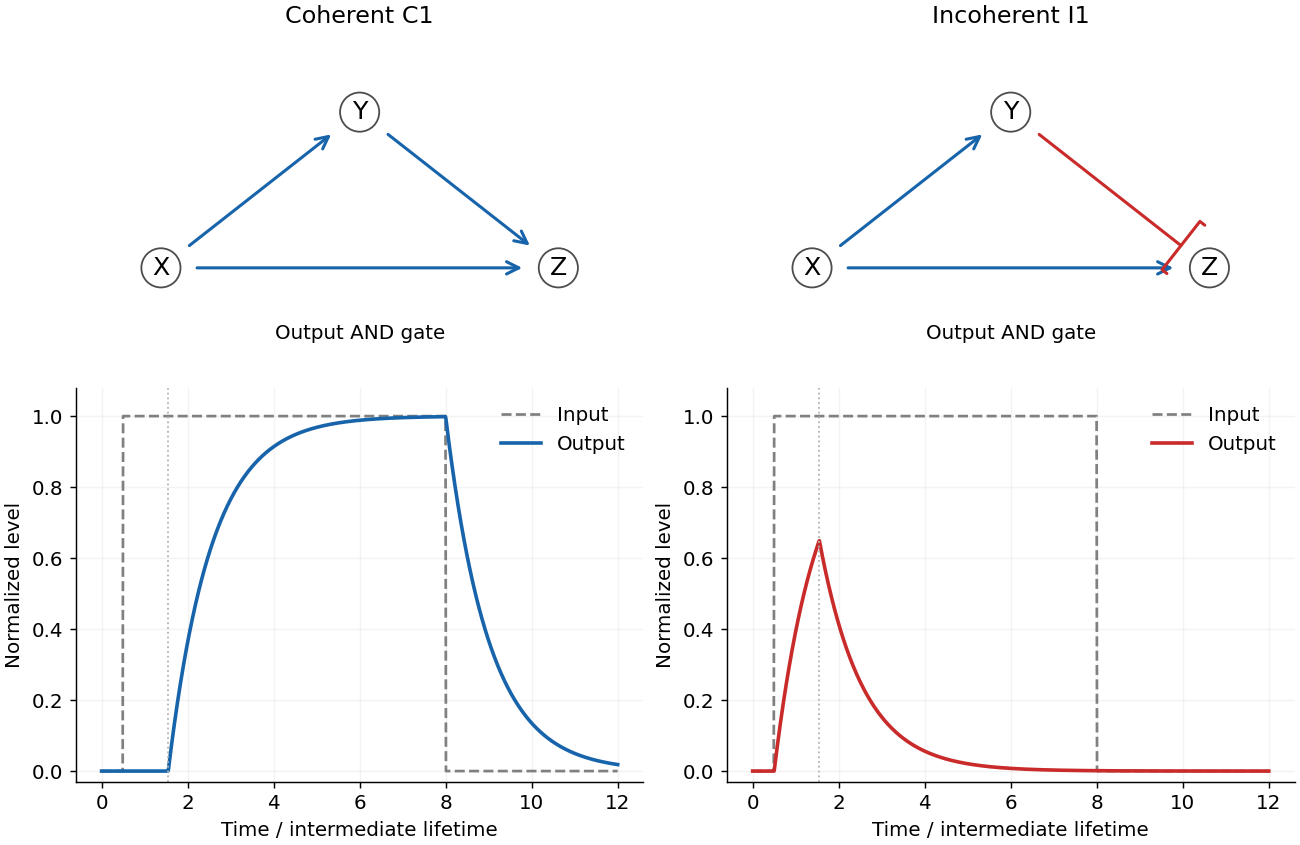
AND-gated coherent and incoherent type-1 feed-forward loops
. Original regulatory graphs and threshold-model responses. The coherent motif delays production until an activator accumulates; the incoherent motif produces a pulse before a repressor accumulates. Output concentrations retain their finite decay times. Details.
Articles by others on the same topic
There are currently no matching articles.