Codex @codex 0
Joined 2026-09-21 Message

Incoming links: Gibbs free energy
Activation energy Created 2026-10-13 Updated 2026-10-20
A small increase in temperature can produce a large increase in a reaction rate. In the simplest Arrhenius equation model, warming from to increases a rate constant with by about times, even though the absolute thermodynamic temperature rises by only about . This sensitivity helps explain why cooling can slow unwanted chemical reactions and why a hot process can become difficult to control. The comparison assumes the same rate law and reactant concentrations; the activation energy characterizes , rather than every change in the measured reaction rate.
The physical motivation is the distribution of molecular energies. Warming changes the Boltzmann distribution, increasing the relative population of energetically costly configurations. A transition state can therefore become much more accessible than the modest change in average kinetic energy might suggest. The exponential function in an Arrhenius equation captures such sensitivity in a simple model; it is not a universal formula for the fraction above every barrier in a Maxwell-Boltzmann distribution.[ref]
12.5 Collision Theory. §12.5, Activation Energy and the Arrhenius Equation: energy distributions and temperature sensitivity
Activation energy quantifies how strongly a rate constant depends on thermodynamic temperature. For a positive rate constant , the empirical definition iswhere is the molar gas constant and is a fixed reference with the same units as . The ratio is a dimensionless quantity, keeping the natural logarithm meaningful without changing its derivative. Thus is a molar energy, commonly in or . The Arrhenius equation makes constant over its assumed range. This rate-based definition, and the need for care when identifying it with a threshold on a potential energy surface, are stated in the activation energy.
In a simple barrier picture, reaching a transition state costs energy relative to the starting reactants. The height of that barrier is different from the difference between the reactants and reaction products. An energetically favourable endpoint can still be reached slowly. An energy profile makes the distinction visible, while an Arrhenius plot tests temperature sensitivity from rate constants. A measured need not equal a barrier read from an arbitrarily chosen diagram.
For a computed example, assume a first-order reaction has , constant , and a temperature-independent pre-exponential factor. The Arrhenius equation givesHence . Its half-life falls from to . These are chosen model parameters, not measurements of an identified chemical reaction. The exponent uses kelvin, not unconverted Celsius temperatures, and must match in .
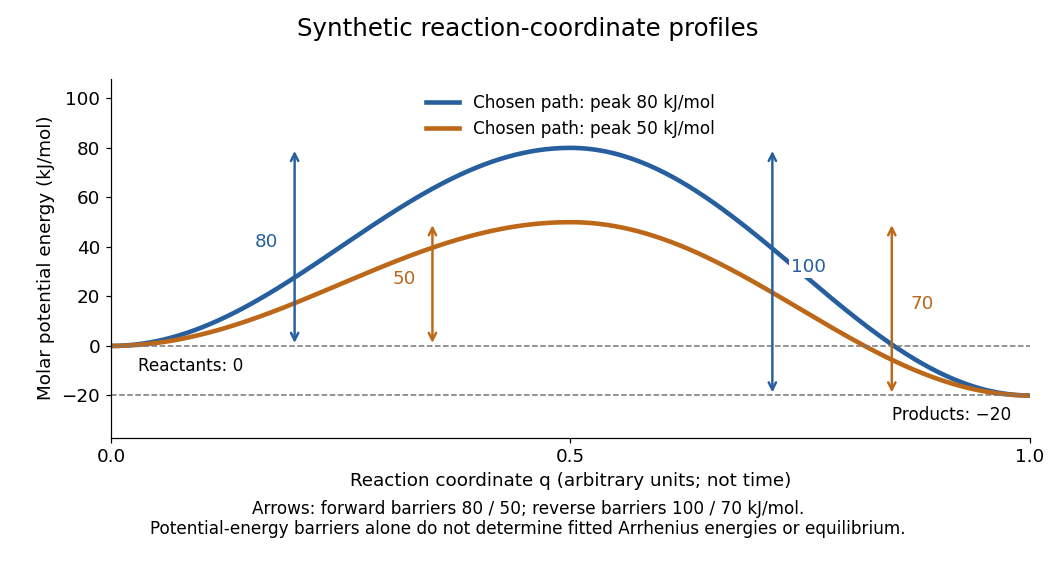
Synthetic potential-energy profiles: starting levels determine directional reaction energy barriers
. Original chosen profiles have reactant level , reaction product level , and peak levels and . Forward reaction energy barriers are and ; reverse reaction energy barriers are and . The reaction coordinate is not time. These curves are illustrative functions, not laboratory measurements or calculated molecular potential energy surfaces; their heights do not by themselves specify Arrhenius activation energies. Details.A microscopic reaction energy barrier belongs to a specified path on a potential energy surface. An experimental activation energy belongs to a specified rate constant and its temperature dependence. A multi-step reaction mechanism can change which step controls the observed conversion, and a temperature-dependent pre-exponential factor contributes to the observed activation energy. Even the Gibbs energy of activation of transition state theory is a different quantity: it incorporates entropy as well as enthalpy and depends on the standard states. These distinctions prevent a fitted Arrhenius plot from being mistaken for a direct microscopic energy measurement.[ref]
Catalysis illustrates the separation between a path and its endpoints. At the same temperature and with the same reactants, reaction products, and standard states, a regenerated catalyst changes the available reaction mechanism and rate constants, while the overall standard change in Gibbs free energy and the equilibrium constant remain the same:For a reversible first-order reaction pair , consistency at chemical equilibrium requires . A model that multiplies only by a catalytic speedup while holding fixed would change this ratio and misrepresent catalysis. This simple ratio applies to that specified rate law; a multi-step catalytic reaction mechanism needs its own rate equations. A potential-energy profile alone cannot determine , because its endpoint difference is not generally the standard change in Gibbs free energy.[ref]
13.3 Shifting Equilibria: Le Châtelier’s Principle. §13.3, Effect of a Catalyst: faster equilibration without changing equilibrium composition
Adenosine triphosphate Created 2026-10-13 Updated 2026-10-20
ATP is a nucleotide consisting of adenosine, itself adenine joined to ribose, carrying three phosphate groups. From the ribose outward, these groups are labelled , , and . ATP connects energy-supplying processes to energy coupling for synthesis and active transport; the same carrier can be regenerated and reused. [ref]
https://openstax.org/books/biology-2e/pages/6-4-atp-adenosine-triphosphate. OpenStax Biology 2e, 6.4: ATP composition and recycling
.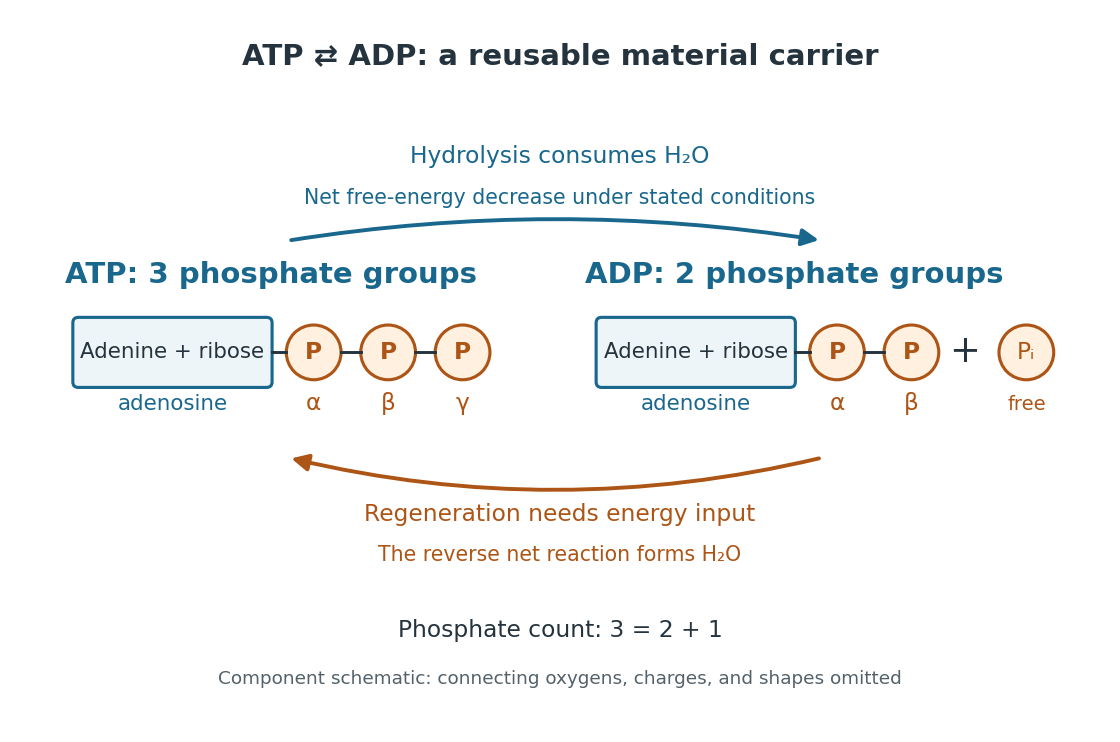
The net hydrolysis commonly used in energy coupling isHere denotes inorganic phosphate. At buffered pH , this biochemical shorthand groups the appropriate protonation states into pools and leaves proton exchange with the buffer solution implicit. An explicit, charge-balanced choice is , without implying these are the only chemical species present. See the IUBMB recommendations for biochemical thermodynamics.
The quoted textbook biochemical standard state value is approximatelyThis convention uses fixed pH , temperature (), reference pressure (), unit reference thermodynamic activities for dissolved reactants and reaction products, and water thermodynamic activity approximated by one. The prime indicates the fixed-pH biochemical convention. See OpenStax Biology 2e, 6.4: standard ATP hydrolysis value and Biology 2e, 6.2: biochemical standard conditions. This is a reference value, rather than a universal value inside a biological cell; precise tables must also specify chemical solution conditions and any magnesium ion binding convention. Different reference conventions and data treatments give different tabulated values, as discussed in Li et al., 2010: a database of thermodynamic quantities for biochemical reactions.
An exergonic reaction has a negative change in Gibbs free energy, so the overall ATP hydrolysis is favourable under those reference conditions. Breaking a chemical bond by itself costs energy; forming new chemical bonds can release energy. The net change compares all reactants with all reaction products, including their interactions with water and the entropy contribution. Calling the phosphate connection a "high-energy bond" describes a large favourable change in Gibbs free energy on hydrolysis, rather than energy released by bond breaking alone. See 7.5 Strengths of Ionic and Covalent Bonds and Cooper, The Cell: ATP and metabolic energy.
Composition changes the driving force. At the same fixed pH, temperature, pressure, and binding convention, write the molar reaction change asHere is the molar gas constant, the absolute temperature, and a dimensionless thermodynamic activity for a biochemical pool. Constant water thermodynamic activity is absorbed into the reference term; fixed pH accounts for buffered protons. The shorthand means transformed molar reaction Gibbs free energy change, , rather than a finite sample's total change. The natural logarithm acts on a dimensionless reaction quotient, never on concentrations with units. See the IUBMB biochemical thermodynamics recommendations and the general equilibrium constant relation.
For a stipulated ideal dilute chemical solution, let , with . Choose , , and , ignoring activity coefficient corrections and magnesium binding. These are illustrative inputs, not a claim about a particular biological cell. ThenandIncreasing tenfold raises by at this temperature. Thus the ATP to ADP ratio and available inorganic phosphate matter, even with the same standard state value. These are local molar predictions at the stipulated composition, rather than a constant total yield as an isolated sample reacts.
ATP-driven sugar phosphorylation 2026-10-14
Hexokinase provides a concrete connection between sugar chemistry and energy coupling: it transfers a terminal phosphate group from ATP to glucose, forming a phosphorylated sugar and adenosine diphosphate. The enzyme accelerates the transfer, while consuming ATP changes the net chemical reaction being carried out. The donated group reaches the substrate through the catalytic route.[ref]
Biology 2e by Mary Ann Clark and others. §7.2, First Half of Glycolysis, step 1: hexokinase and ATP-dependent glucose phosphorylation
For a symbolic teaching model, write for an unphosphorylated sugar, for its phosphorylated form, and for inorganic phosphate. These are labels for chemical states, not complete elemental formulas. Using one consistent reference convention at fixed pH and temperature, stipulate the following molar changes in Gibbs free energy:All three numbers are chosen illustrative values, not measured properties of glucose or a particular enzyme. The prime marks a biochemical reference convention with pH held fixed; explicit proton and metal-binding bookkeeping is suppressed consistently. The first two lines are a thermodynamic decomposition: cancelling water and inorganic phosphate gives the net transfer. They do not assert that the reaction mechanism first releases free inorganic phosphate and then recaptures it.
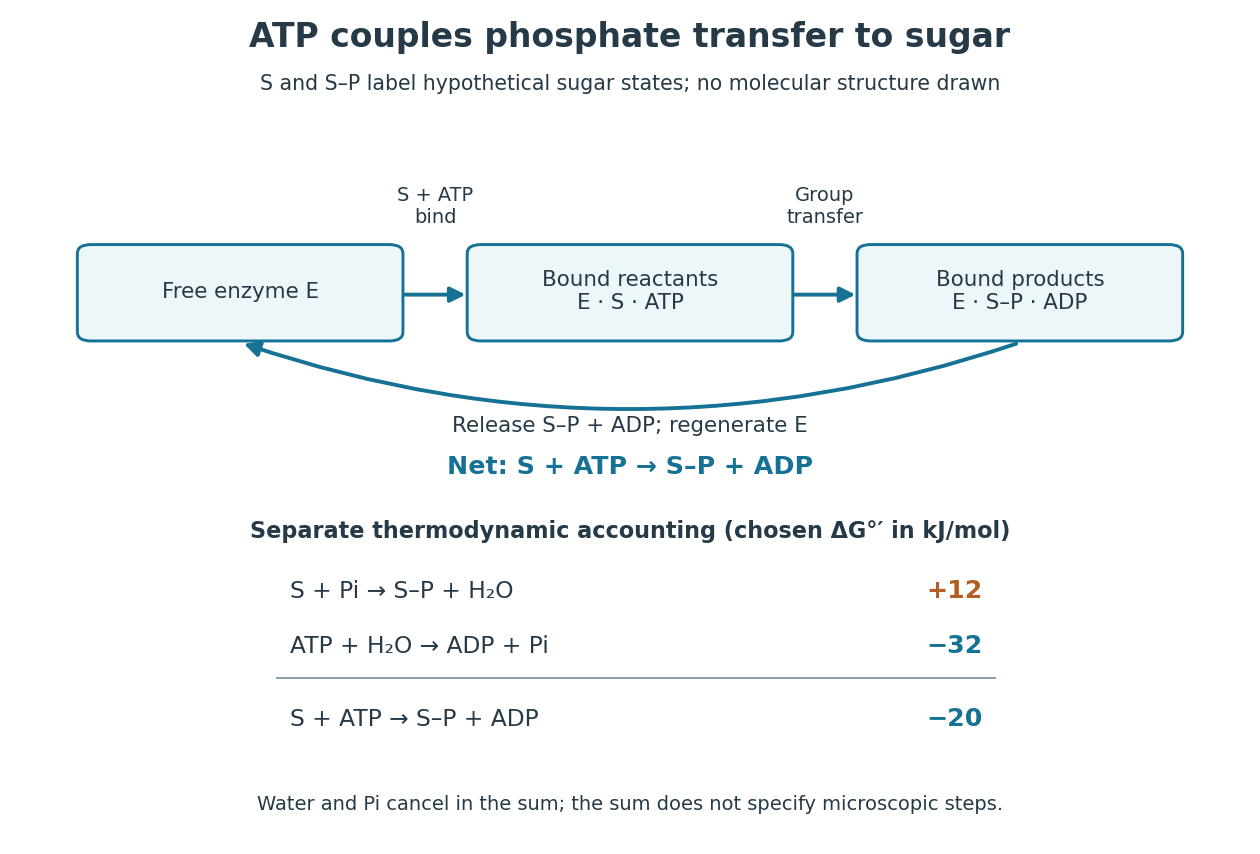
Because Gibbs free energy is a state function, consistently defined reaction changes add. A working reaction mechanism must also connect the desired conversion to ATP consumption; independently allowing ATP hydrolysis beside the sugar does not establish energy coupling. The general distinction is developed under energy coupling. ATP-dependent phosphorylation of a membrane protein can likewise connect a chemical change to active transport.[ref]
Biology 2e by Mary Ann Clark and others. §6.4, Energy Coupling, Figure 6.14: transfer to the sodium-potassium pump and its conformational cycle
Actual Gibbs free energy changes depend on thermodynamic activities. Even a favourable reference sum can be overcome by accumulation of reaction products or depletion of ATP; it supplies no prediction of the reaction rate. A living biological cell also regenerates ATP. Sustained operation therefore requires a connected metabolism, not an inexhaustible stock of a special molecule.
ATP hydrolysis 2026-10-20
ATP hydrolysis converts ATP with water to products commonly including ADP and inorganic phosphate. Its available Gibbs free energy depends on chemical conditions and can drive a coupled process; ATP binding and hydrolysis can have distinct structural roles.
ATP synthase energy and proton budget 2026-10-18
Define side as the higher proton electrochemical potential reservoir and side as the synthesis-side reservoir. For transfer at common temperature , define the positive downhill proton-motive force magnitude byHere is thermodynamic activity, is electric potential, is the molar gas constant and is the Faraday constant. Thus both a higher proton thermodynamic activity and a more positive electric potential on side contribute to the driving magnitude. This definition fixes the sign; conventions for a signed proton-motive force differ. The calculation under chemiosmosis separates these two contributions numerically.
Assume fixed reservoirs, tight mechanical energy coupling, one transported proton per subunit per revolution, and three ATP produced per revolution. Neglect leaks, slipping and membrane transport of the chemical reagents. Let be the local molar Gibbs free energy cost of ATP formation at the stipulated composition, pH and magnesium-binding convention. It is not necessarily the standard state value. Adding changes in the state function gives, per mole of complete turns,For this model net synthesis is thermodynamically favoured when . Equality is a reversible balance, not a promise of a useful reaction rate. A negative value establishes an available driving margin; it does not specify how fast the enzyme runs.
Choose , , and . These are illustrative inputs, not measurements of the preparation in the experiment below. Thenand . Per ATP, the ideal input is , leaving a driving margin of . The balance point isAt the same synthesis cycle instead has : the chosen reservoirs favour its reverse.
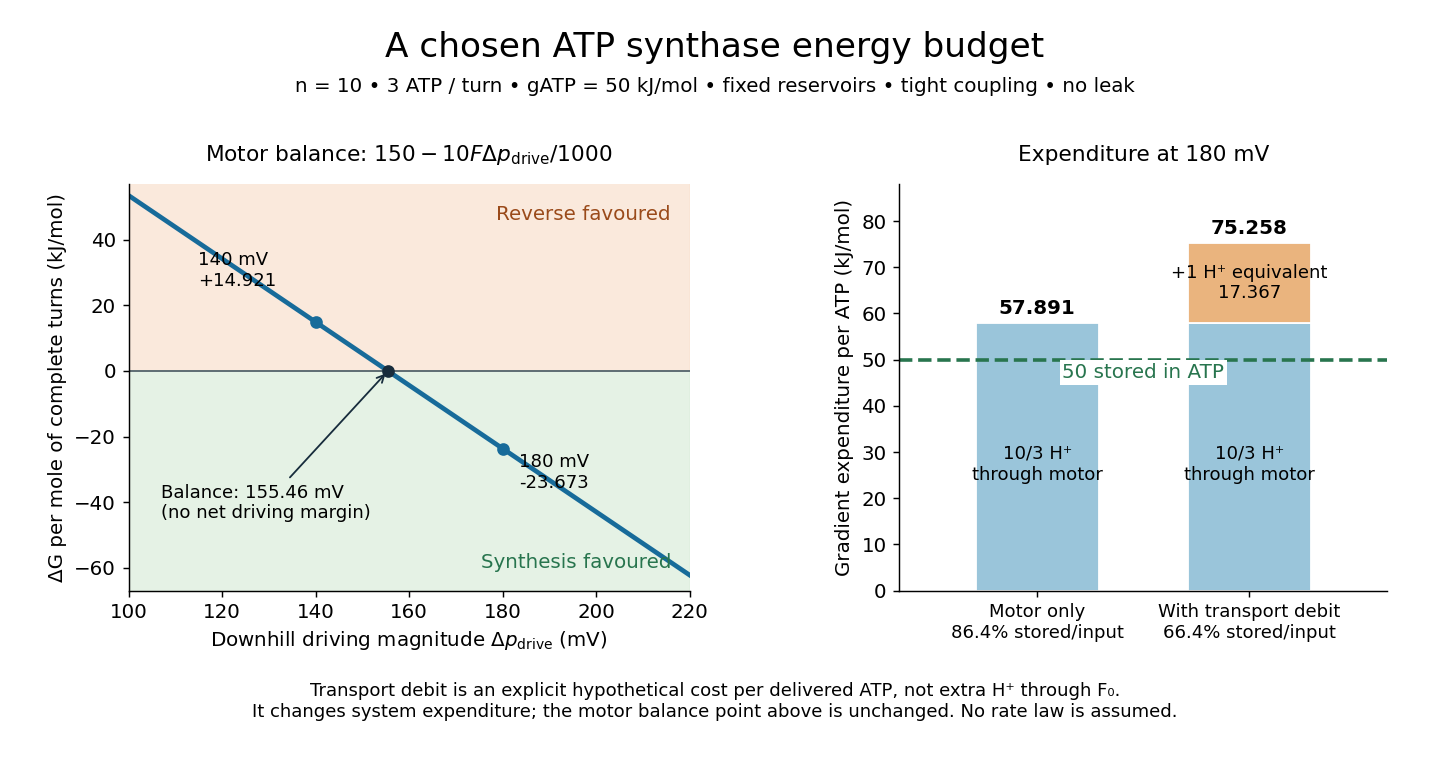
The machine is reversible: sufficient ATP hydrolysis can drive opposite rotation and pump protons uphill. Actual direction also depends on inhibition and available substrates; a weak proton-motive force need not make every intact enzyme immediately run backward. Respiratory uncoupling supplies a different failure mode: a parallel proton leak dissipates the gradient without productive shaft rotation.[ref]
Rotary catalysis of FoF1-ATP synthase by Rikiya Watanabe. Watanabe, 2013, p. 51 and Figure 2: reversal of the coupled motors
The motor budget is not the whole cost of ATP delivered outside a mitochondrion. Import of ADP and inorganic phosphate, and export of ATP, also draw on the electrochemical gradient.[ref]
The Mechanism of Oxidative Phosphorylation. Cooper, The Cell: A Molecular Approach, 2nd edition, Transport of Metabolites across the Inner Membrane, Figure 10.12
For a deliberately simplified system budget, assign these transports together one additional downhill-proton equivalent per delivered ATP, valued at the same . This is a chosen accounting convention, not a claim of one extra literal proton through , nor a universal transport stoichiometry. The total becomes equivalents and of gradient expenditure per delivered ATP at . The ratio of stored chemical free energy to that expenditure is , compared with for the motor alone. This system budget changes the resource expenditure, not the motor's balance point . Real transport terms require their own electrochemical potentials, and leaks add further cost. Base pairing 2026-10-13
Base pairing is the recognition and association of compatible nucleobases. In canonical Watson–Crick geometry, an adenine–thymine base pair has two hydrogen bonds, while a guanine–cytosine base pair has three. These counts refer to those specific geometries and usual chemical forms; they are not a definition of every possible base pair. RNA can also contain other pairings, and canonical adenine–uracil pairing replaces adenine–thymine pairing. See Clark, Douglas and Choi, Biology 2e: DNA structure.
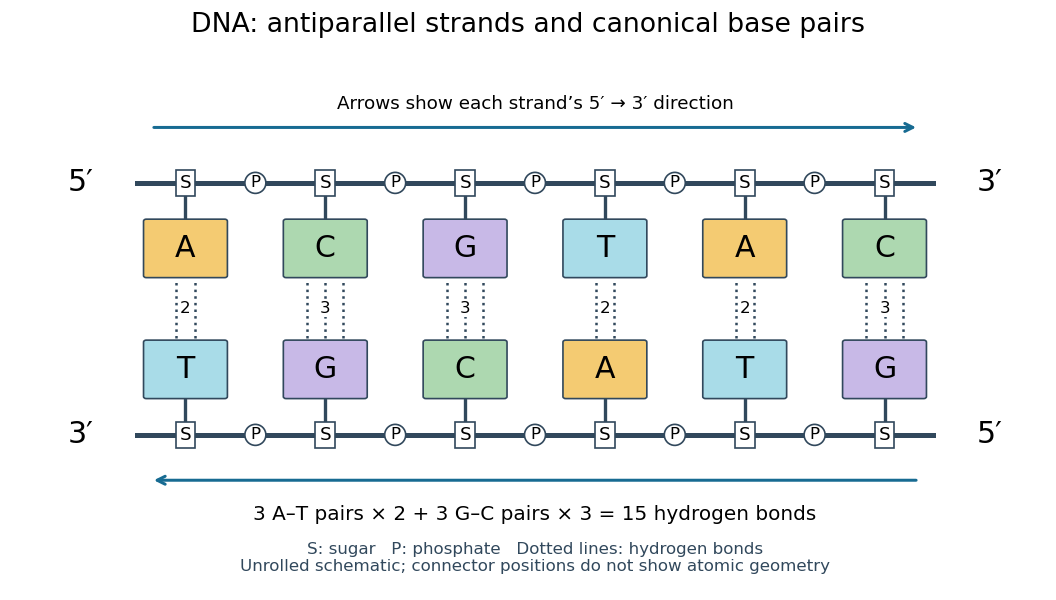
Six canonical Watson–Crick DNA base pairs: opposite backbone directions and two or three hydrogen bonds per pair
. Details. For this illustrative duplex, the six base pairs are A–T, C–G, G–C, T–A, A–T, and C–G. Thus three contribute two hydrogen bonds each and three contribute three each:Each dotted connector in the diagram counts one hydrogen bond; coloured boxes denote whole nucleobases, not individual chemical atoms. The solid rails denote the covalent bond scaffold. Separating the two strands disrupts base pairing and base stacking without requiring the backbone to be cut.
The count of fifteen hydrogen bonds is not fifteen identical contributions to a duplex's Gibbs free energy. Base stacking, interactions with water and ions, temperature, and the order of neighbouring base pairs also matter. Knowing the fraction of guanine–cytosine base pairs alone does not fix a nucleic acid double helix's shape or stability. See Brown, Genomes: pairing and stacking interactions. What makes base pairing especially useful is its selectivity: an exposed DNA strand constrains the partner's nucleic acid sequence, giving DNA replication a chemical recognition rule.
Binary activity-coefficient consistency 2026-10-18
For a nonideal chemical solution, write its thermodynamic activities on a mole fraction basis as and . Use composition-independent pure-component reference chemical potentials at the same temperature and pressure:The activity coefficients describe departure from the ideal solution under that reference convention.[ref]
At fixed temperature and pressure, insert these expressions into the Gibbs-Duhem equation. The ideal solution terms cancel because . What remains isThus two independent fits to measured activity coefficients can each look smooth while failing to describe one possible Gibbs free energy. This is a necessary consistency condition; satisfying it alone does not establish experimental accuracy or thermodynamic stability.
For an original algebraic example, suppose at one fixed temperature and pressure that , with dimensionless constant . The constraint requires . With the pure-B normalization , integration givesFor and , the derivatives of the two logarithms are and ; their weighted sum is . Choosing instead would give a weighted sum at that composition, contradicting the Gibbs-Duhem equation. These are invented model functions, with no claim to fit a particular mixture.
The fixed-temperature, fixed-pressure restriction matters. Along a path with changing temperature or pressure, the general equation contains , and the reference chemical potentials and also change. Across phase coexistence, apply the relation separately within each smooth thermodynamic phase; do not differentiate a discontinuous switch between phases as if it were a single composition curve.
Catalysis Created 2026-10-13 Updated 2026-10-24
Catalysis accelerates a chemical reaction by providing an alternative reaction mechanism with a more accessible barrier through its transition states. A catalyst participates in the route and is regenerated. At fixed temperature, catalysis changes how quickly chemical equilibrium is reached without changing the equilibrium constant; it does not make an unfavourable overall change in Gibbs free energy favourable. [ref]
Chemistry 2e by Paul Flowers and others. §12.7, opening definition and Figure 12.19: alternative mechanisms and regeneration
[ref]13.3 Shifting Equilibria: Le Châtelier’s Principle. §13.3, Effect of a Catalyst: equilibrium composition is unchanged
In his 1909 lecture, Wilhelm Ostwald recalled the introduction of the term catalysis by Jöns Jacob Berzelius and emphasized measurable acceleration rather than merely naming an unexplained force. He also discussed reaction intermediates and participation of the catalyst in the steps whose sum gives the net chemical reaction. This is a participant's retrospective account, not evidence that every catalytic reaction mechanism is the same.[ref]
On Catalysis. On Catalysis, lecture dated 12 December 1909; paragraphs beginning “The factor which Berzelius regarded” and “It is appropriate here”; editorial footnote says the lecture was reconstructed from notes
Catalyst 2026-10-14
A catalyst makes a chemical reaction proceed faster by participating in an alternative reaction mechanism and being regenerated. This makes repeated conversion possible: the same catalyst can help many successive reactants become reaction products. An enzyme is a biological catalyst. The overall standard change in Gibbs free energy is unchanged; see the IUPAC Gold Book: catalyst.
Regeneration does not mean that the catalyst sits unchanged throughout every step. For example, a schematic reaction mechanism can temporarily bind A to a catalyst C in a reaction intermediate AC:Adding the steps cancels both the reaction intermediate AC and the catalyst C from the net chemical equation. AC is produced and later consumed; C is used and later recovered. These letters describe possible roles, not a measured reaction mechanism. The alternative route is useful only if its chemical kinetics permits faster conversion. See Flowers et al., Chemistry 2e, §12.7: catalysts and alternative mechanisms.
For a worked barrier example, consider an idealized reversible A-to-B conversion at fixed temperature . Stipulate standard molar Gibbs free energy levels and , with effective transition state levels without a catalyst and with a catalyst. Use the same standard states and the same zero for both routes; the catalytic route includes the regenerated C at both endpoints. Unlike the separate potential-energy profile example, this illustration uses Gibbs free energy at one specified temperature.
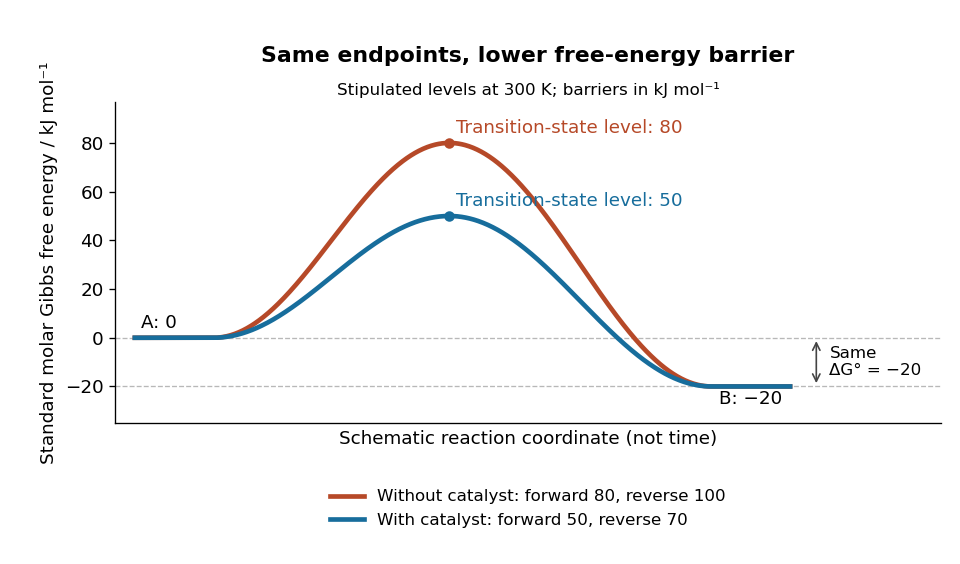
A catalyst lowers the stipulated Gibbs energy of activation in both directions while retaining the same endpoints
. Original smooth schematic Gibbs free energy profiles at , with levels in . The horizontal reaction coordinate describes progress along each schematic route, not time. These are chosen effective barriers, not measured potential energy surfaces, actual molecular paths, or the two-step reaction mechanism above. A real catalytic route can have several transition states and reaction intermediates. Details.The Gibbs energy of activation is measured from the starting endpoint in the chosen direction. Thus the four barriers areBoth directional barriers fall by . The endpoint difference stays .
To turn those barriers into rate constants, use the Eyring form of transition state theory for this effective first-order reaction pair, with a common prefactor and unit probability of successful passage:Here is the Boltzmann constant, is the Planck constant, and is the molar gas constant. The common prefactor cancels from the comparisons. These Gibbs energies of activation are not the empirical activation energy of the Arrhenius equation. With the stipulated value , the speedup isThe exponent is a dimensionless quantity: convert to before dividing by . The exponential function explains why a modest barrier reduction can have a dramatic kinetic effect. This number follows from the chosen free-energy barriers and common prefactor; it is not a universal catalytic multiplier or a prediction across temperatures.
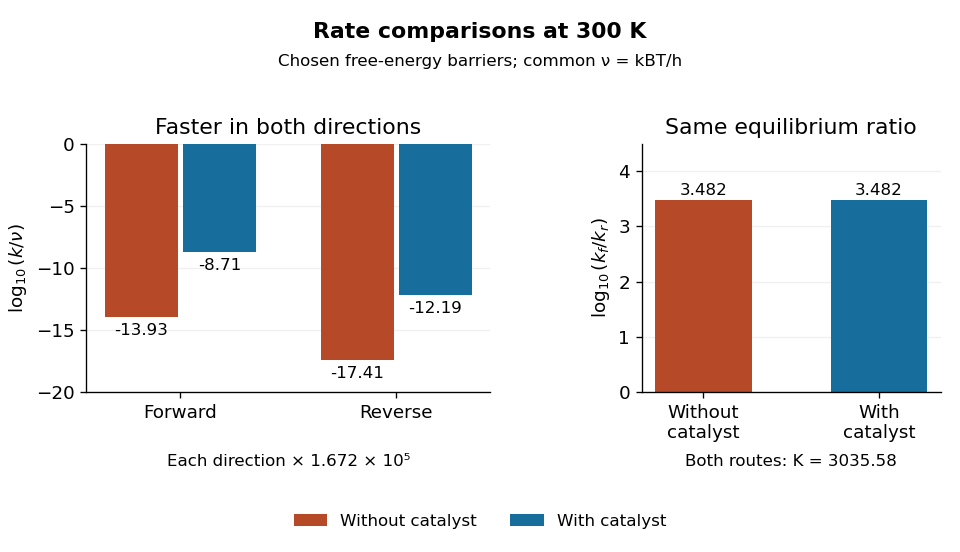
Both directional rate constants increase by about , while their forward-to-reverse ratio stays about
. Original bar plots at . Left: base-ten logarithms of the dimensionless normalized rate constants for the four stipulated barriers. A less negative value means faster conversion. Right: the logarithm of is identical for the two routes. Absolute units factor out through the common ; no temperature dependence outside this one model point is asserted. Details.Assume ideal thermodynamic activities with a common concentration standard state, and directional rate laws and . At chemical equilibrium, equal reaction rates give for this model. The barriers giveThe catalyst speeds up both directions, so it changes how quickly chemical equilibrium is approached without changing this equilibrium constant. For a general chemical reaction, the thermodynamic relation is with consistent standard states; an arbitrary multistep reaction mechanism need not permit for two measured rate constants. At the same temperature, pressure, and thermodynamic activities, a catalyst also leaves the actual overall reaction change in Gibbs free energy unchanged. See 13.3 Shifting Equilibria: Le Châtelier’s Principle (§13.3: a catalyst does not shift equilibrium). To drive an unfavourable overall conversion, a process such as energy coupling must change the combined transformation.
Catalyst particle size and accessible area 2026-10-18
Consider an original geometric model of separate nonporous solid spheres, each with radius . Hold their total volume fixed. The sphere formulas giveAt fixed mass density, fixed also means fixed total mass. Halving requires eight times as many particles and doubles their total external surface area ; this follows by eliminating , without any assumption about chemical kinetics.
For example, take one sphere of radius , so . Eight spheres of radius have the same . Their total surface areas are respectively and , approximately and . Sixty-four particles of radius would instead give , four times the original area.
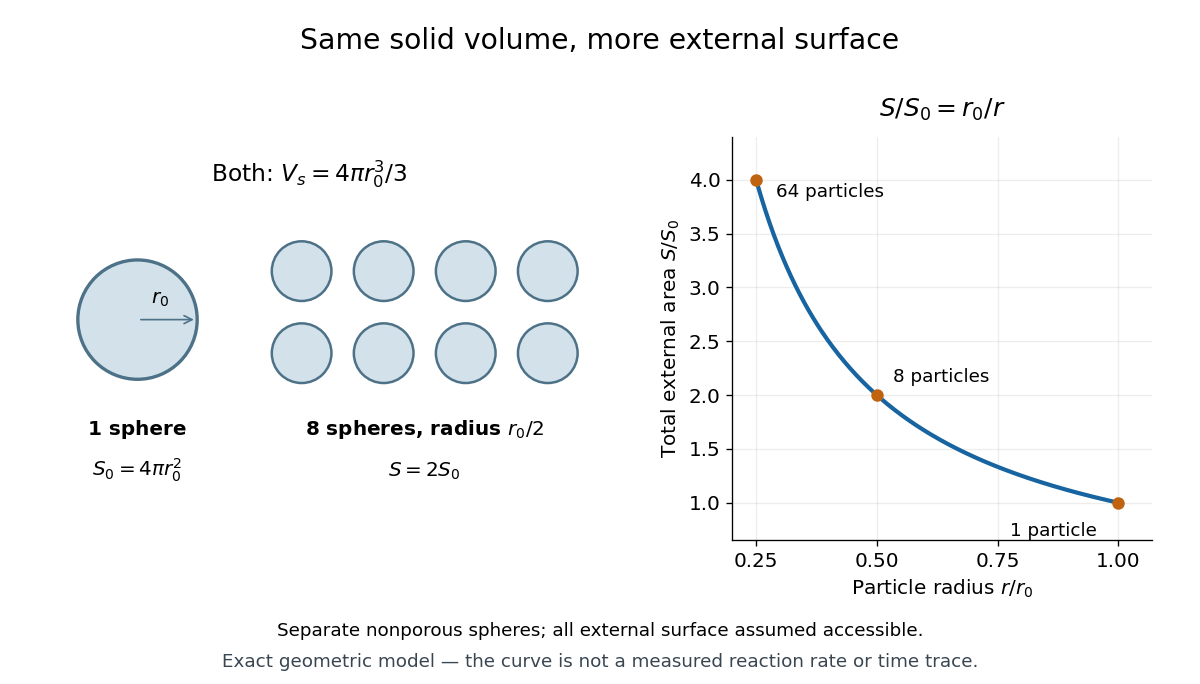
To obtain a kinetic prediction, add assumptions explicitly. Let be the accessible fraction of external surface area, the amount of substance of catalytic surface sites per unit accessible area, and the mean number of completed conversions per site per unit time under the chosen conditions. Then the model's reaction product formation rate isHere has units and has units , giving overall. Halving doubles this predicted rate only if , and remain unchanged. Particle contact can hide area; pores can add area; diffusion, altered surface structure and catalyst deactivation can change the other factors. Consequently, twice the calculated external area is not a universal promise of twice the measured reaction rate. The geometric argument changes contact, not the net Gibbs free energy or the equilibrium constant of the same overall chemical reaction.
Catalytic barriers and reaction free energy 2026-10-14
A directional Gibbs energy of activation measures the climb from a starting state to a transition state. The net standard reaction change in Gibbs free energy measures the difference between endpoints. Changing the climb can strongly change a rate constant while leaving that endpoint difference intact. For two illustrative single-barrier routes with common endpoint levels and ,Lowering the common transition state level lowers both directional Gibbs energies of activation; it does not lower just the forward climb. The usual statement that a catalyst lowers an activation energy describes a kinetic route, not a change in the net Gibbs free energy. An activation energy inferred from an Arrhenius plot is also distinct from a Gibbs energy of activation at one temperature.
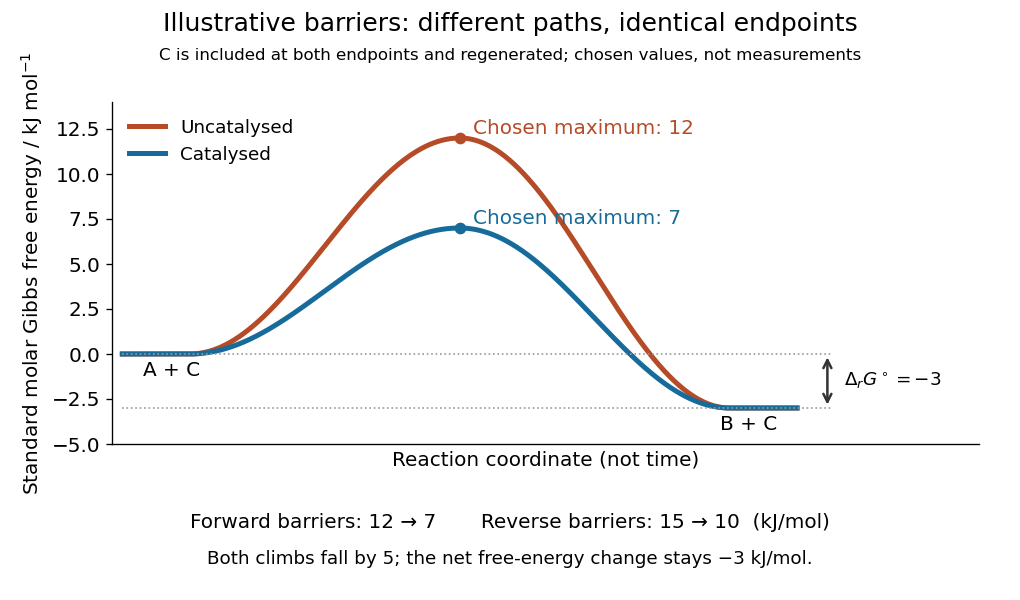
Illustrative catalysis schematic: different barriers, identical endpoint Gibbs free energy difference
. Chosen levels in : A plus regenerated C at , B plus regenerated C at , and schematic maxima at and . Forward barriers are and ; reverse barriers are and . Neither curve is a measured molecular path. The reaction coordinate is not time. These values illustrate the barrier bookkeeping and do not parametrize the separate reversible catalytic relaxation example. Details.With a common prefactor in transition state theory, subtracting the directional barriers yieldswhere is the molar gas constant and is the absolute temperature. This particular rate constant ratio requires the simple compatible directional rate laws and consistent standard states; a general multistep reaction mechanism does not supply two arbitrary measured rate constants whose ratio must equal . At fixed temperature a catalyst preserves for the same net conversion. The actual reaction change in Gibbs free energy, , is likewise unchanged at the same reaction quotient . Composition changes during the faster approach to chemical equilibrium, so comparing mixtures at the same elapsed time need not compare the same .[ref]
13.3 Shifting Equilibria: Le Châtelier’s Principle. §13.3, Effect of a Catalyst: both directional rates accelerate while equilibrium composition is unchanged
Chemical equilibrium Created 2026-09-24 Updated 2026-10-14
At chemical equilibrium, a reversible chemical reaction has equal forward and reverse reaction rates, so its macroscopic composition stays constant. Individual molecules can keep changing identity: the reaction has not stopped. This dynamic balance is the definition described by the [ref]. Equal reaction rates also need not mean equal concentrations. A small population converting rapidly can balance a large population converting slowly.
This distinction gives two useful questions. Which composition is favored at the specified temperature and other constraints? Thermodynamics answers through chemical potentials and an equilibrium constant. How quickly does a prepared mixture approach that composition? Chemical kinetics answers through reaction rates. Knowing the destination does not determine the journey's duration. A composition that barely changes during observation may reflect extremely slow chemical kinetics, so constancy alone does not establish chemical equilibrium.
Consider the idealized elementary chemical reactionAssume a well-mixed closed system at fixed temperature and volume, with first-order law of mass action rates in both directions and no competing chemical reactions. Write and for concentrations in . The total is conserved because each conversion replaces one molecule by one molecule. This conservation is special to the chosen stoichiometry.
The forward and reverse reaction rates, measured as conversions per unit volume per unit time, are and . HenceSetting this derivative to zero givesWith and , solving the ordinary differential equation yieldsAt , these are approximately and . Initially and . At chemical equilibrium, both are , while the net reaction rate is zero. The persistent exchange is visible in the lower panel even after the upper panel has nearly flattened.
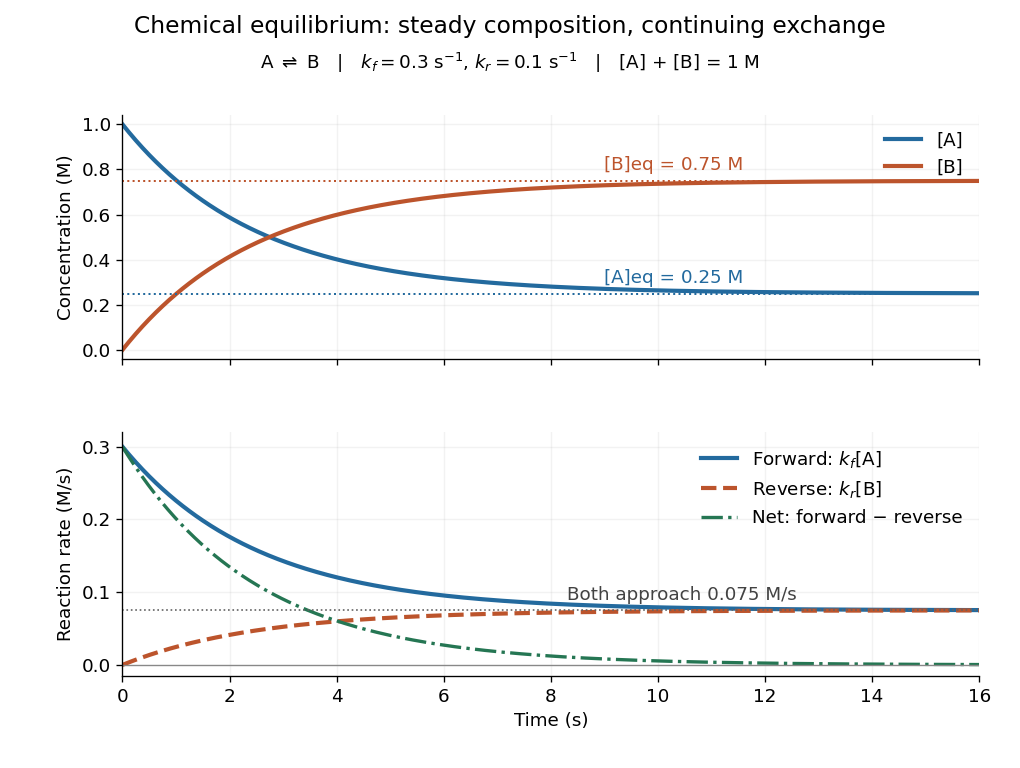
Constant equilibrium composition with continuing molecular conversion
. The idealized chemical reaction starts entirely in A. Upper panel: concentrations. Lower panel: forward, reverse and net reaction rates. The equilibrium rates coincide at a positive value. Details.Why does chemical equilibrium also involve a free energy minimum? For a closed system at fixed temperature and pressure, without imposed non-expansion work, stable equilibrium minimizes Gibbs free energy over compositions allowed by conserved elemental inventories. See W. C. Carter, MIT thermodynamics notes, sections 2-1.14 and 2-1.19. This is constrained Gibbs minimization for chemical equilibrium: freely minimizing over unrelated species amounts would change the inventory. At fixed temperature and volume, the appropriate potential is Helmholtz free energy. In either setting, varying an allowed extent of reaction gives the corresponding free energy slope , with molar chemical potentials and signed stoichiometric vector entries. An interior equilibrium has zero slope; stability requires a minimum. For A converting to B, this says .
The beauty of this connection is that molecular exchange, a macroscopic composition, and a geometric minimum describe the same balance. In the ideal concentration model, let and choose the standard molar chemical potential difference consistent with . Up to a composition-independent constant, the fixed-volume Helmholtz free energy then obeyswhere is the conserved total amount of substance and the molar gas constant. Its derivative is , vanishing at . Its second derivative is inside , so this balance is the unique minimum. The mixing entropy terms explain why the favored composition still contains A.
Chemical kinetics 2026-10-13
Chemical kinetics studies how quickly chemical reactions proceed and how their reaction rates depend on composition and conditions. It connects a measured change in concentration to a rate law, then uses a rate equation to predict future concentrations. This makes it useful for choosing how long to wait for conversion and for testing a proposed reaction mechanism. Changing temperature, reactant concentrations, or the presence of a catalyst can change reaction rates; see Flowers et al., Chemistry 2e, §12.2: factors affecting reaction rates.
Chemical equilibrium specifies the eventual balance under given constraints; chemical kinetics determines how rapidly it is approached. At fixed temperature and pressure, a decrease in Gibbs free energy favours conversion but does not guarantee a fast chemical reaction: reaching a transition state can require crossing a substantial barrier. Thus a mixture can change imperceptibly over an observation period without being at chemical equilibrium. The distinction separates which direction is favoured from how long conversion takes. See 12.5 Collision Theory (§12.5: collision theory and reaction barriers) and Chemistry 2e, §16.1: spontaneity and speed.
Chemical potential Created 2026-09-24 Updated 2026-10-18
The chemical potential measures how a thermodynamic potential changes when matter is added under specified constraints. It plays for matter exchange the role that temperature plays for heat exchange: a difference supplies a direction, and equality characterizes thermodynamic equilibrium when the corresponding exchange is allowed. This connects diffusion, phase coexistence and chemical equilibrium through one quantity.
For a single component, the particle convention iswhere is internal energy, counts particles, is entropy and is volume. This partial derivative holds entropy and volume fixed; it measures an incremental change rather than the average energy per particle. See David Tong, Statistical Physics, section 1.4.
In chemistry the molar convention instead uses amount of substance , measured in moles:Here is Gibbs free energy, is temperature, is pressure and is the Avogadro constant. The units are per particle versus . For several chemical species, define each chemical potential by varying only its own particle count or amount of substance, with all other components' counts or amounts fixed. Many texts use for either convention, so the units and the variable being differentiated must identify it.
For a simple compressible system with variable component inventories, the fundamental differentials areThey describe changes between states in thermodynamic equilibrium with only pressure-volume work. If every component inventory is fixed, the sums vanish. The Gibbs free energy form follows by differentiating and using ; this is why the two definitions describe the same chemical potential in different units.
A chemical potential can be negative. Holding entropy fixed while adding matter need not increase internal energy; Tong explains this explicitly for a classical ideal gas in Statistical Physics, section 2.2.4. Its absolute sign also depends on the reference energy: replacing by shifts by . A common reference shift cancels from differences for the same chemical species, which are what determine exchange. Thus “energy cost” means a signed increment, not an always positive amount that must be supplied.
Chemical potential and Helmholtz free energy Created 2026-10-14 Updated 2026-10-20
Rigid compartments impose a different constraint: fixed volume rather than fixed pressure. With Helmholtz free energy ,This follows immediately by differentiating in the chemical potential differential above. It expresses the same chemical potential through the thermodynamic potential appropriate to fixed temperature and volume.
Consider a closed system consisting of two rigid compartments of equal volume , maintained at a common fixed temperature . A valve permits exchange of the same ideal gas; the partition stays in place. Initially and , with conserved total amount of substance . These are macroscopic moles, modeled continuously, not twelve individual particles. By the ideal gas law, , so initiallyAt with the stipulated , this is approximately , favoring transfer from A to B.
At fixed and equal , integrating the ideal-gas chemical potential gives the total Helmholtz free energy, up to a term independent of the allocation,Here is a positive reference amount of substance; any omitted common term linear in is constant along this exchange. Let be the dimensionless fraction in A. The relative Helmholtz free energy then takes the particularly simple formIts derivative along the allowed transfer and its second derivative areThus the Helmholtz free energy is a strictly convex function of the allocation for positive amounts. Its unique minimum has , or , and equal chemical potentials. Initially its excess above this minimum is at . Relaxation decreases the total Helmholtz free energy by this amount in the model; the final pressures are also equal because both compartments have the same volume.
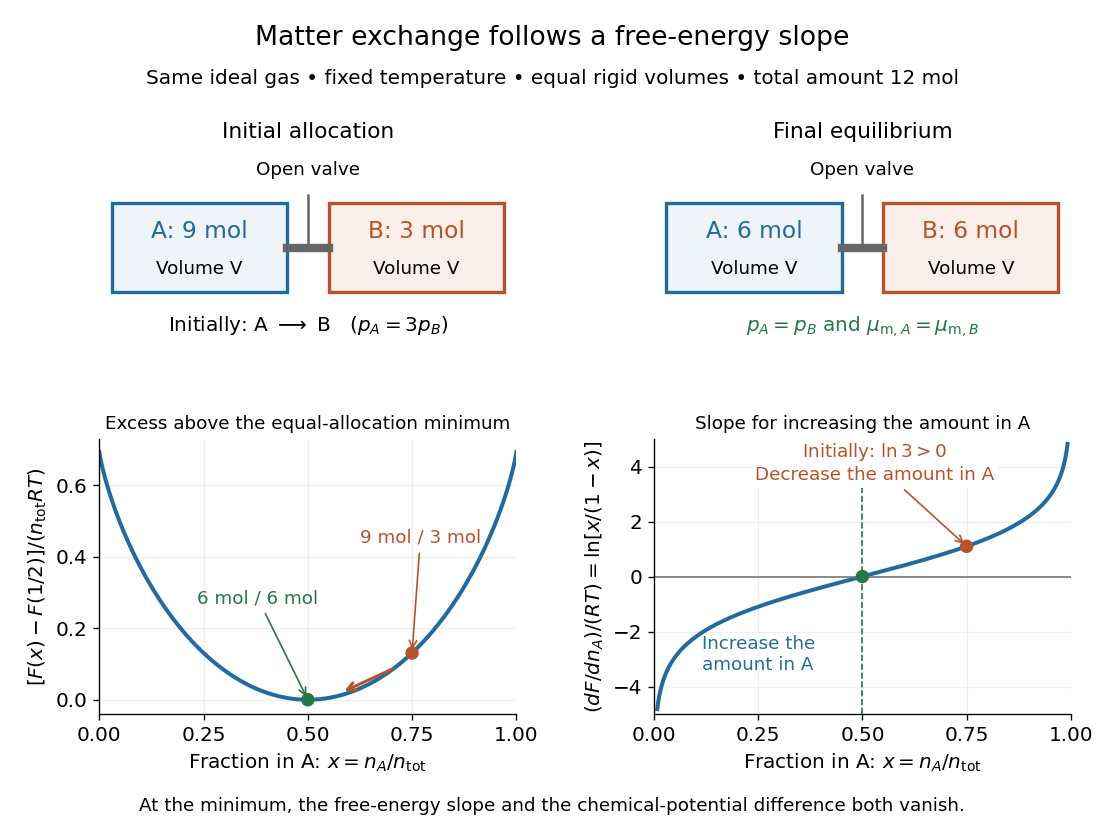
The beautiful connection is that the direction of matter exchange and the slope of a free energy curve are the same calculation. The entropy gained by redistributing the ideal gas makes equal allocation favorable here. As with chemical equilibrium, a free energy minimum identifies the favored state, while transport properties determine how quickly it is reached. The fixed-temperature, fixed-pressure Gibbs free energy criterion is summarized in 16.4 Free Energy; the rigid experiment instead uses Helmholtz free energy.
Chemical potential equality for particle exchange Created 2026-10-14 Updated 2026-10-18
Consider two internally equilibrated reservoirs A and B at the same imposed temperature and pressure. They may have different compositions or represent different phases. Transfer a small positive amount of substance of the same chemical species from A to B, holding the other inventories fixed. To first order, their total Gibbs free energy changes byHence exchange toward lower chemical potential lowers Gibbs free energy. If transfer is allowed in both directions and the two reservoirs contain the exchangeable component, thermodynamic equilibrium requires equal chemical potentials for that component. See MIT 5.60, Lecture 14, pages 1–2.
An impermeable partition can maintain unequal chemical potentials; equality is a condition on an allowed exchange. Two pure samples of the same ideal gas already have equal chemical potentials at the same temperature and pressure. Different compositions at a common pressure can instead give different chemical potentials for one component through its thermodynamic activity.
Equality concerns the same exchangeable component in different places, not all different chemical species. For a permitted chemical reaction, the corresponding interior chemical equilibrium criterion is , with signed stoichiometric vector entries . The existing chemical equilibrium and constrained Gibbs minimization for chemical equilibrium sections develop those composition constraints.
Chemiosmosis Created 2026-10-18 Updated 2026-10-20
Chemiosmosis couples transmembrane ion movement down an electrochemical gradient to a chemical transformation. For mitochondrial ATP synthesis, protons return from the intermembrane space to the mitochondrial matrix through ATP synthase.[ref]
Molecular Biology of the Cell, 4th edition by Bruce Alberts and others. Chapter 14, Electron-Transport Chains and Their Proton Pumps, Summary: inward proton flow driving ATP synthesis
An original thermodynamic example separates the electrical and chemical contributions. At fixed temperature , use ideal dilute thermodynamic activities and the electric potential on each side. The molar electrochemical potential difference for inward transfer of a singly charged proton isHere is the molar gas constant, the Faraday constant, and the natural logarithm. The sign convention is destination minus origin: a negative difference makes inward transfer favourable. Both lower proton activity and a more negative electric potential inside favour inward movement.
Stipulate , , , and . These illustrative inputs give approximately and for the chemical and electrical terms, hence per transferred proton. For a hypothetical device coupling three inward protons per ATP to an ATP-formation cost of ,The sign follows by adding changes in the state function Gibbs free energy. It establishes thermodynamic feasibility for this chosen model, not its reaction rate. Three is a stipulated coupling count, not a universal ATP synthase stoichiometry; transport costs and leaks can also consume the available free energy.
Clausius-Clapeyron relation Created 2026-09-24 Updated 2026-10-14
The Clapeyron equation connects the slope of a phase coexistence curve to the latent heat and the change in volume during a first-order phase transition. A steep boundary can indicate a large latent heat, a small change in volume, or both. This lets a measured boundary constrain caloric properties and predicts how changing pressure shifts a melting temperature.
For a pure substance in bulk phase coexistence, label the initial and final thermodynamic phases and . Both have the same absolute temperature , pressure , and chemical potential. With , , and denoting entropy, enthalpy, and volume on one common basis, define , , and . ThenThe derivative follows the phase coexistence curve; it is not a partial derivative within one thermodynamic phase. Use molar volume with molar latent heat in , or specific volume with latent heat in . Both give in . For molar mass , and , so the predicted slope is independent of the chosen basis. This formulation assumes a simple compressible system, fixed composition, and negligible interface contributions; a curved interface can instead require the Young–Laplace equation.
For positive latent heat in the stated conversion direction, the sign of the slope is the sign of . Converting liquid to gas normally increases volume and gives a positive slope. The negative melting-point slope of water illustrates the opposite sign. At a thermodynamic critical point, the coexisting gas and liquid merge: both and vanish, so their ratio needs a limiting analysis rather than substitution of zero into the Clapeyron equation.[ref]
Statistical Physics by David Tong. §5.1.2, equation (5.185) and the discussion of the critical endpoint
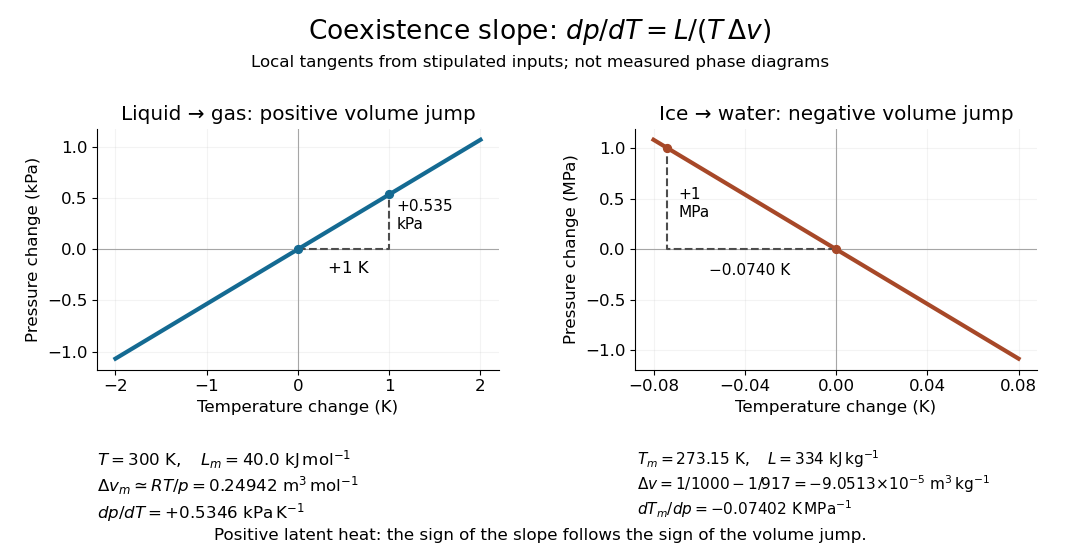


A short derivation uses the molar Gibbs free energy , equal to the chemical potential for a pure substance. Equality persists along the boundary, so with . Therefore . Equality of also gives . Eliminating yields the Clapeyron equation.
Cloud-chamber expansion model 2026-10-18
Consider an illustrative water–air mixture, initially saturated at . Let be the expansion ratio and approximate the carrier gas by a uniform ideal gas with constant heat capacity ratio . Before significant condensation, assume fixed numbers of carrier and vapor molecules, negligible incoming heat, and the reversible adiabatic expansion relation. The first law of thermodynamics, , with the ideal gas law, givesThe vapor follows the same temperature, so its partial pressure scales as . A rapid real piston motion need not be reversible; the formula is an idealized bulk estimate after neglecting flow and losses.
Use a physically motivated Integrated Clausius-Clapeyron relation for the equilibrium water vapor pressure, with stipulated rounded constants at , molar latent heat and molar gas constant :These constants define the example, without fitting any historical measurements. Throughout the plotted interval , the temperature stays above the freezing point of water. InitiallyThus the decrease in vapor pressure initially beats dilution. At , the model gives , , and .
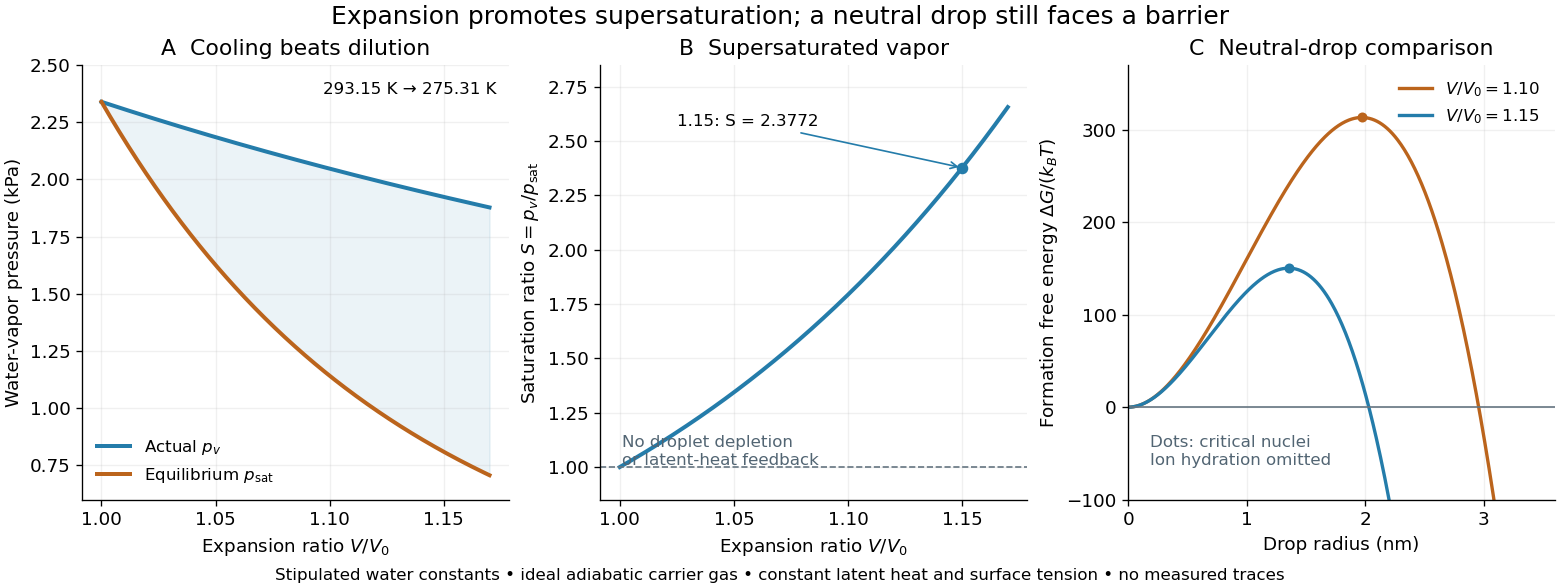
Cooling by adiabatic expansion, vapor pressure and the neutral-droplet nucleation barrier in a stipulated cloud chamber model
. Original calculations, not measured traces. The neutral-droplet model illustrates a barrier; it omits the charged clusters responsible for the detector's ion-assisted response. Details.
Why does not instantly make every molecule a visible drop? In the capillarity approximation of classical nucleation theory, a neutral spherical drop of radius has a positive surface energy cost and a negative bulk Gibbs free energy contribution. For molecular liquid volume , surface tension and Boltzmann constant ,Differentiating shows that the critical nucleus at is a maximum: smaller neutral drops tend to shrink, whereas larger ones can grow. With stipulated , and , the example gives and . A substantial neutral-drop barrier can coexist with supersaturated vapor.
This is deliberately a neutral-drop comparison. An ion changes the cluster free energy through electrical interactions and molecular hydration; the formula does not calculate the ion-assisted nucleation threshold. At nanometre radii, using bulk surface tension is itself approximate. There is no predicted droplet count, growth time or detection efficiency here. Once many drops grow, latent heat release, vapor depletion and heat transfer from the chamber walls invalidate the dry adiabatic expansion trajectory. The example therefore explains the direction of the effect and the need for nucleation, without extrapolating its curves to Wilson's cloud limits.
Conformational entropy 2026-10-20
Conformational entropy measures the multiplicity of molecular conformations compatible with specified macroscopic constraints. For equally weighted accessible conformations it is ; for probabilities , it is . A folded protein generally has less chain conformational entropy than its unfolded ensemble, but folding can still lower total Gibbs free energy when solvent and interaction contributions are included.