Thermodynamics studies energy transfers and the constraints on macroscopic changes. Its vocabulary connects internal energy, entropy, temperature, pressure and volume to heat and work. The first law of thermodynamics balances energy; the Second law of thermodynamics decides which changes can occur spontaneously and limits how much work a heat engine can deliver. These principles let us bound performance before choosing a material or a detailed machine design. Applications include power generation, propulsion and refrigerators. [ref]
https://web.mit.edu/16.unified/www/FALL/thermodynamics/notes/node10.html. MIT, What thermodynamics describes
Start by choosing a system boundary. A closed system can exchange energy but no matter; an isolated system exchanges neither. A state function, such as internal energy or entropy, depends on the equilibrium state. A process specifies how that state changes. Thus a body's temperature is a property it has, whereas heat records energy crossing its boundary during a process. A body contains internal energy, rather than a stored quantity of heat.
Two insulated bodies initially at different positive temperatures approach thermal equilibrium when brought into thermal contact. Their combined energy is conserved, but their combined entropy increases. They do not spontaneously separate again into a hotter body and a colder one. This familiar asymmetry makes thermodynamics interesting: conserving energy leaves many imaginable changes available, and entropy supplies a further constraint. The same distinction underlies both entropy maximization under thermal contact and the thermal efficiency limit in Carnot's theorem.
Conversion of organized work into internal modes with macroscopic entropy production. Total energy remains conserved. For a passive linear material, the loss coefficient must give nonnegative supplied work over a completed cycle.
Suppose a small volume change obeys , with its displayed coefficients held constant. Work on the body is . Two rectangular pressure-temperature paths between the same endpoints differ by , where path a changes temperature first. The straight interpolation samples the mean pressure during its temperature change, giving . Variable material coefficients need not preserve that average relation.
Thermal energy is a context-dependent name for the microscopic energy associated with a system's thermal state, rather than a separate conserved substance. The characteristic single-particle energy scale is , with Boltzmann constant and absolute temperature . Comparing this scale with an excitation gap or energy barrier helps decide which microscopic processes are thermally accessible.
Under the classical equipartition theorem, each independent quadratic term in the Hamiltonian contributes on average. Three translational terms therefore give per particle of a monatomic ideal gas. Quantum levels can invalidate that count: a degree of freedom with excitation spacing much larger than can remain nearly frozen. A measured heat capacity tests how the stored internal energy changes with temperature; it does not simply count a fixed amount of heat stored in the sample.
Maxwell's demon is a thought experiment in which a controller measures microscopic motion and selectively operates a barrier, apparently driving a system away from thermal equilibrium without the usual thermodynamic cost. For example, admitting fast gas particles preferentially to one side seems to create a temperature difference that a heat engine could exploit.
The controller must be included in the physical accounting. Its measurement record becomes correlated with the gas, and a repeatable cyclic device must deal with that record rather than treating information storage as an unlimited free resource. Landauer's principle connects logically irreversible memory erasure with a thermodynamic cost under its stated conditions. This does not mean that every measurement automatically dissipates exactly one bit's erasure cost. The point is to track the complete cycle, correlations and resources. A controller that reduces one subsystem's entropy can still respect the Second law of thermodynamics by increasing entropy elsewhere.
Landauer's principle relates logically irreversible information erasure to thermodynamic cost. Resetting an initially equiprobable, uncorrelated one-bit memory to one standard state while coupled to a bath at temperature requires average heat release of at least to the bath in the ideal cyclic setting.
The restriction matters: erasing one unknown bit is different from reversibly transforming a known state, and real devices can dissipate more. The entropy reduction of the memory must be compensated elsewhere. The bound therefore concerns information loss under specified physical conditions, rather than a universal cost for every logic operation.[ref]
Irreversibility and Heat Generation in the Computing Process by Rolf Landauer. 1961, logical irreversibility and heat generation
A later experiment represented a bit by the two wells of a potential trapping one colloidal particle. Driving the particle distribution into a chosen well allowed dissipated heat during erasure to be compared with the bound. Approaching the reversible limit requires slow controlled operations; measurement of the work and heat connects the information statement to a physical apparatus.[ref]
https://www.nature.com/articles/nature10872. Bérut and others, 2012, abstract and single-particle erasure experiment
A state of matter is a broad category of material behavior. A solid resists a sustained change of shape, a liquid flows while remaining difficult to compress, and a gas expands to fill its container. These distinctions are useful first descriptions, but they do not enumerate all thermodynamic phases. Plasma adds collective behavior of charged particles, while liquid crystals show that flow and structural order need not disappear together.
Distinguish three questions: what broad state of matter does the material resemble, which thermodynamic phase is present, and what are its temperature, pressure and other state variables? Two samples at different temperatures can be in the same liquid thermodynamic phase. Conversely, crystal polymorphism gives different solid thermodynamic phases of the same substance, distinguished by their crystal structures. A phase transition between them need not change the broad state-of-matter category.
For an imagined substance with crystalline forms A and B, melting A produces a solid–liquid phase transition, while converting A to B produces a solid–solid phase transition. A sample containing A and B has two thermodynamic phases even though all its material is solid. Cutting one uniform crystal into two pieces does not create a second thermodynamic phase: count distinct equilibrium forms, rather than pieces or grains. This distinction matters when applying the phase rule.
A phase diagram locates stable thermodynamic phases using temperature and pressure. Its liquid–gas phase coexistence curve ends at a liquid-gas critical point, around which a path connects liquid and gas without a first-order phase transition. Broad categories can therefore remain useful without a sharp boundary.[ref]
Statistical Physics by David Tong. §5.1.2, printed pp. 140–141, Figures 40–41: pressure–temperature coexistence and disappearance of the liquid–gas distinction
A solid resists sustained changes of shape on the timescale being considered. It can support a static shear stress, whereas an ordinary liquid continues to deform under sustained shear. Small loads often produce recoverable elastic deformation; larger loads can cause plastic deformation or fracture.
A solid need not be a crystal. A crystalline specimen has long-range structural order, while an amorphous solid such as ordinary glass lacks the same repeating lattice arrangement. The mechanical classification and the structural classification answer different questions. Applying a small lateral load to the top of a block while holding its base fixed tests shear resistance; a finite shear displacement at static balance is a solid-like response. At long times some solids also creep, so the observation interval and applied stress matter when comparing a material's response with an ideal elastic model.
A crystal is a solid with long-range spatial order in its constituents. In a periodic crystal, translation by a lattice vector reproduces the structure; a unit cell and its constituent arrangement describe the repeating pattern. The atomic structure is distinct from the visible external shape: an irregularly cut specimen can remain a single ordered crystal.
Diffraction probes that order because radiation scattered from many regularly arranged sites can reinforce at selected directions. The resulting pattern reveals structural spacings and orientation, rather than merely showing the specimen's outer faces. A single crystal maintains one coherent orientation across the specimen, while a polycrystalline material contains many crystal grains. Crystal orientation can make properties anisotropic, so a test along one direction need not predict response along another. Surface shape additionally depends on equilibrium surface free energy and growth kinetics.
A crystal lattice records the translation symmetry of a periodic crystal. With a primitive basis of three linearly independent lattice vectors , its translation vectors formThe crystal structure additionally specifies the arrangement of atoms associated with each lattice point. A lattice point is therefore a symmetry marker, not necessarily one atom.[ref]
Matrices, Mappings and Crystallographic Symmetry by Hans Wondratschek. Section 1.5, Definition D 1.5.2 and primitive bases
For example, a one-dimensional chain with alternating species at spacing has translation period , not : shifting by exchanges distinct species. It can be described by lattice with a two-atom basis at offsets zero and . In a diffraction measurement, the translation lattice selects possible reciprocal-space peak positions, while the atomic basis affects their intensities and systematic absences. This distinction lets diffraction reveal internal structure rather than just the visible shape of the specimen.
A magnetic unit cell repeats the ordered magnetic moments as well as the atomic positions. It can exceed the chemical primitive unit cell when translations interchange oppositely ordered magnetic sublattices. For a simple-cubic Néel pattern , translations preserving the sum's parity form an FCC lattice with conventional parameter twice the chemical spacing and a two-spin primitive basis.
A macroscopically planar crystal boundary characterized by a crystallographic orientation. Its Miller index specifies that orientation; visible facet area depends on surface thermodynamics and growth kinetics. A low surface energy and a slow growth rate are different properties.
A solid containing many differently oriented crystalline crystal grains. grain boundaries impose compatibility and act as transport paths or sinks for mobile defects.
A grain in a polycrystal is a region whose crystal structure has approximately one common orientation, separated from differently oriented regions by grain boundaries. The atoms remain ordered within each grain even when a macroscopic specimen contains a complicated mosaic of orientations. A grain is therefore different from an amorphous patch or a separate chemical phase.
The boundaries enforce geometric compatibility between neighboring crystals and can obstruct dislocation glide, provide rapid transport paths or act as sinks for defects. A mechanical measurement over many grains averages both the intragrain response and those boundary effects. Observing a prepared cross-section can reveal grain shape, but a single two-dimensional section need not directly give every grain's three-dimensional size. This distinction matters when comparing an elongated grain structure with one described by a single mean grain size.
A measure of crystal grain elongation. It changes directional boundary spacings and can influence anisotropic defect transport independently of average grain size.
A measure of the spatial scale of crystal grains. Its definition must be stated for elongated crystal grains; size, aspect ratio and orientation distribution have distinct physical effects.
Crystal packing describes how constituents are arranged and make contact within a crystal. It includes both the repeated geometry and, for molecular crystals, the orientations and intermolecular contacts of the molecules. Composition alone does not determine this arrangement: the same substance can have different crystal polymorphs with different packing.
A useful geometric model replaces constituents by equal hard spheres. Hexagonal close packing and face-centered cubic packing put each sphere next to twelve nearest neighbours but arrange successive close-packed layers differently. Their local contact geometry can therefore agree while their longer-range order differs. Real molecular shapes and directional intermolecular forces supply further constraints. Packing affects mass density, available slip geometry and directional response, making diffraction a way to connect structural measurements with macroscopic properties rather than merely assign a repeating pattern.
A cocrystal is a single crystalline phase containing two or more distinct molecular components in a shared ordered structure. Its components occupy defined roles in the repeating arrangement, rather than simply forming separate crystals mixed together. A stylized arrangement of molecules A and B illustrates the idea without assuming that every cocrystal has that ratio.
Intermolecular forces and molecular geometry influence which combined structure is favorable. A diffraction measurement tests the shared ordering, while a phase diagram distinguishes a stable combined phase from coexistence of separate A-rich and B-rich phases. This is useful because a shared lattice can change properties such as melting behavior and mass density even though the identities of the molecular components remain recognizable. The word describes a structural phase assignment and does not by itself predict which component combinations will form one.
Close packing of triangular layers with ABAB stacking. Its nearest-layer geometry agrees with face-centered cubic stacking, but the further-layer registry differs.
A cubic crystal structure with lattice sites at cube corners and face centres. Its close-packed layers are (111) planes, stacked ABCABC.
The fixed-volume equilibrium crystal shape obtained by minimizing orientation-dependent surface free energy. Facet support-plane distances are proportional to their interfacial free energies under the construction assumptions.
An equilibrium construction for a supported particle that modifies the Wulff construction using particle-support and support-medium interfacial energies. The support truncates the corresponding free-particle shape.
A single crystal is a crystalline specimen whose ordered structure has one continuous crystallographic orientation, rather than being divided into differently oriented crystal grains. It can still contain dislocations, impurities and other defects; the term does not mean an absolutely perfect lattice.
A diffraction pattern from one well-oriented specimen resolves its crystallographic directions, while rotating it changes which reflections are observed. Mechanical or thermal measurements along different directions can reveal intrinsic anisotropy that would be averaged over many orientations in a polycrystalline material. The absence of internal grain boundaries also changes transport and deformation mechanisms that depend on those boundaries. A specimen's outline is not the criterion: cutting a piece from a single crystal changes its external shape without necessarily changing its internal crystallographic identity.
A liquid flows under a sustained shear stress while usually resisting a change of volume much more strongly than a gas. It adopts the shape of a container but, when it coexists with a gas and gravity permits, can form a free surface rather than expanding to fill all available space. These are mechanical observations, not a requirement that every liquid be exactly incompressible.
At rest in a uniform gravitational field, a liquid develops the hydrostatic pressure variation needed to support its weight. During flow, viscosity relates rate of deformation to viscous stress for a Newtonian fluid. Releasing a small surface disturbance probes its dynamical response through gravity waves and surface tension. Thus its static pressure, resistance to rapid compression and resistance to shear rate are distinct measurable properties. Liquid and gas can also connect continuously around a liquid-gas critical point, so the broad labels do not imply a first-order boundary along every thermodynamic path.
A droplet is a bounded amount of liquid surrounded by another phase, such as a gas or an immiscible liquid. Its interface carries surface energy. For a free droplet at fixed volume with uniform positive surface tension, reducing surface area lowers this contribution to the free energy; when other effects are negligible, the equilibrium shape is a sphere. This is the physical setting behind the isoperimetric inequality.
For a spherical droplet of radius and constant surface tension , write and . Quasistatic expansion against the pressure difference givesThis is the spherical Young–Laplace equation: the pressure inside exceeds the outside pressure. An illustrative gives at , and twice that pressure difference at half the radius.
Real surface tension depends on material and temperature; the IAPWS release, for example, tabulates measured water values and supplies an interpolation formula.[ref]
Revised Release on Surface Tension of Ordinary Water Substance by International Association for the Properties of Water and Steam. Printed p. 3, correlation and validity range; Table 1, measured values.
Gravity, contact with a solid and motion can deform a droplet, so the spherical calculation states a regime rather than defining every possible droplet shape.In a homogeneous isotropic one-component fluid, is the conditional number density at separation from a tagged particle divided by the bulk number density . Thus the displayed shell contains the mean number of neighbours. For an uncorrelated ideal gas away from the tagged particle; repulsive cores suppress small separations, while liquid coordination shells can produce damped oscillations approaching one at large distance. This is a pair correlation, not a normalized one-particle radial probability density. The static structure factor measures its Fourier-space counterpart.
For a differentiable central pair potential in a classical homogeneous three-dimensional fluid, the mechanical virial theorem gives the displayed pressure. The radial distribution function accounts for the probability of each pair separation. In the dilute limit, , and integration by parts yields . Thus the second virial coefficient controls . A few virial terms are a low-density approximation, not generally a converged description of a dense liquid. A discontinuous hard core needs its contact form or a limiting argument.
For a classical monatomic homogeneous fluid with pairwise central pair potential , the mean interaction energy is one half the sum over neighbours of a tagged particle. Using the radial distribution function gives the displayed formula; the factor one half prevents counting each pair twice. The kinetic term follows from the equipartition theorem. Its use requires the relevant integral to exist and a thermodynamic state specified by number density and temperature. Quantum kinetic contributions, many-body forces and internal molecular modes require modifications.
A gas is a fluid that expands to occupy its available container and is readily compressed compared with an ordinary liquid. Its pressure reflects microscopic momentum transfer to boundaries. In a sufficiently dilute classical gas, molecular separation is large enough that the ideal gas law can approximate its equilibrium response.
A real gas can depart from that law because intermolecular interactions and finite molecular size become significant. Compressing a gas or changing its temperature therefore tests an equation of state, rather than a universally exact proportionality. A vapor is the gaseous form of a substance under conditions where a condensed phase can be reached by suitable compression or cooling. The same substance can pass continuously between liquid-like and gas-like states around its liquid-gas critical point, so a complete phase description requires a phase diagram as well as the broad word gas.
A historical physical name for a gas or vapour, emphasizing its mobility and appreciable compressibility. It does not imply an exactly ideal gas or prevent a vapour from undergoing condensation.
A gas law relates equilibrium variables such as pressure, volume, temperature and amount of substance. The ideal gas law provides a useful dilute-gas model. Holding temperature and particle number fixed predicts ; holding pressure and particle number fixed predicts with absolute temperature.
A piston and cylinder provide a direct comparison: vary one controlled variable slowly enough for the gas to equilibrate, then record the remaining pair. The constraint is part of the law being tested; rapid compression can heat the gas and need not follow an isothermal curve. At larger density, real gas effects require a more suitable equation of state. A gas law describes possible equilibrium states, while the first law of thermodynamics additionally determines the heat and work along a specified process.
A vapor is a substance in a gaseous state at a temperature where a condensed phase can also be reached under suitable pressure conditions. For a pure substance below its liquid-gas critical point, isothermal compression can reach phase coexistence and then convert some vapor to liquid. At coexistence the equilibrium vapor pressure depends on temperature.
A sealed cell containing both water and water vapor illustrates the distinction between amount and state: at fixed temperature, changing the liquid fraction can leave the equilibrium vapor pressure unchanged while both phases remain. A dilute vapor can still approximate an ideal gas, so vapor does not mean that ideal-gas calculations are automatically invalid. Conversely, the broad term gas does not imply ideal behavior. Specifying composition, temperature and pressure makes the usage physically meaningful.
Crystal polymorphism is the existence of different crystal structures for the same chemical substance. Each crystal polymorph can have its own mass density, melting behavior and range of stability on a phase diagram. A different external shape of the same crystal structure alone does not establish a different crystal polymorph.[ref]
polymorph. IUPAC Compendium of Chemical Terminology, fifth edition, entry “polymorph” (15225), definition and Note 2
At fixed temperature and pressure, compare the chemical potentials of the possible crystal polymorphs. The lower value determines the stable bulk thermodynamic phase; equality permits phase coexistence when neither form is beaten by another thermodynamic phase. A higher-chemical potential form can persist through metastability, so finding it in a sample does not by itself locate an equilibrium boundary.
A solid thermodynamic phase with a particular crystal structure, among distinct crystalline forms of the same chemical substance. Crystal polymorphism names the existence of these alternatives; a crystal polymorph names one of the forms.
A thermodynamic system is the part of the physical world selected for an energy and matter balance. Draw its boundary before classifying an interaction as heat, work or transported matter. The boundary may move: a sealed piston encloses a closed system even while its volume changes. An isolated system additionally excludes all external energy transfers.[ref]
Thermodynamics and Propulsion. Thermodynamics and Propulsion, §1.2.2: system boundaries and closed systems
Boundary choice changes the ledger. If an electrical heater warms water, choosing only the water counts incoming heat; including the heater counts electrical work entering through its leads. The combined internal energy records storage, without separate stored categories for the two transfer mechanisms.[ref]
Thermodynamics and Propulsion. Thermodynamics and Propulsion, §1.3.4, Figure 1.14: resistor and water with alternative system boundaries
An equilibrium thermodynamic state specifies sufficient independent macroscopic properties, together with material identity and constraints, to determine the other equilibrium properties. For a uniform fixed-composition ideal gas, its amount of substance, temperature and volume determine its pressure through the ideal gas law. These properties specify a state, without specifying a thermodynamic process that prepared it.[ref]
A thermodynamic process changes a thermodynamic system between states. A quasistatic process supplies equilibrium intermediate states and therefore a path on equilibrium coordinates. An irreversible thermodynamic process can have equilibrium endpoints while lacking a uniform intermediate temperature or pressure. Endpoint values still determine changes in a state function; computing heat and work requires the actual interaction history.
A thermodynamic system for which the reversible mechanical work is entirely pressure-volume work. At fixed composition its internal energy is specified by entropy and volume; additional work modes require additional state variables.
A thermodynamic system is in thermodynamic equilibrium when its macroscopic state has no spontaneous tendency to change through any exchange allowed by its constraints. For an isolated system, stable thermodynamic equilibrium maximizes entropy over the accessible states at fixed total energy and conserved material inventories. A wall can forbid an exchange, so always specify what can cross or move. Thermal, mechanical and chemical conditions answer different questions about that same constrained state. [ref]
For example, two compartments separated by a rigid, impermeable wall that transmits heat can reach thermal equilibrium while retaining different pressures and compositions. Allowing an unloaded piston to move adds a mechanical equilibrium condition. Allowing one chemical species to cross adds chemical potential equality for particle exchange; allowing a chemical reaction adds its chemical equilibrium condition. A fixed barrier can therefore support a constrained thermodynamic equilibrium that ceases to be an equilibrium when the barrier is removed.
Uniformity is not the definition of thermodynamic equilibrium. An isothermal atmosphere in hydrostatic equilibrium can have a common temperature while its pressure and mass density vary with height: the pressure gradient balances gravity. In an external field, the exchange condition must include the relevant potential energy; for charged matter this leads to the electrochemical potential. Relativistic gravitational corrections to thermal equilibrium are described separately by the Tolman temperature law.
A steady state only says that the macroscopic description is constant in time. A bar maintained between hot and cold thermal reservoirs has steady heat flux and produces entropy, so it is not in thermodynamic equilibrium. Even matching temperatures at one perfect-contact interface can coexist with heat transfer through the surrounding material; matching those two local values does not establish equilibrium of the whole apparatus. See thermal conduction for that spatial transport problem.
The useful simplification is that equilibrium conditions predict an endpoint without computing every collision or flow during relaxation. The first law of thermodynamics supplies the conserved quantities, and the Second law of thermodynamics supplies the direction of change. For a system held at fixed temperature and volume, the corresponding criterion is minimization of Helmholtz free energy; at fixed temperature and pressure, it is minimization of Gibbs free energy, with the material and exchange constraints still imposed.
Coexistence without a thermodynamic driving force for further exchange. Phases able to exchange heat and constituents have equal temperature and chemical potentials of the exchanging species. Mechanical balance is also needed; interface curvature can require a pressure jump rather than equal bulk pressures.
At fixed temperature and pressure, liquid-liquid equilibrium is stable coexistence of distinct liquid thermodynamic phases that minimizes total Gibbs free energy for the specified component amounts. Their compositions can differ even though each exchanging chemical species has the same chemical potential in both phases. For neutral, nonreacting components in bulk phases at common temperature and pressure , neglecting external-field and interface-curvature contributions,To see why, transfer an amount from phase to phase while holding the total amount of each component fixed. The change in Gibbs free energy is . If the coefficient is nonzero, one transfer direction lowers . Equality removes that driving term; stable equilibrium additionally requires a global minimum over the allowed phase arrangements. Here each chemical potential is the partial derivative of Gibbs free energy with respect to that component amount at fixed temperature, pressure and the other component amounts.[ref]
Charged species instead require equality of the corresponding electrochemical potential. Equal chemical potentials for neutral species do not require equal mole fractions. With a common pure-component reference, and , where is thermodynamic activity, an activity coefficient, and the molar gas constant. The activity equality for coexisting liquids is . Different nonideal environments can therefore support different compositions at the same chemical potential.
For a binary mixture, let be the mole fraction of component B and let be the homogeneous molar Gibbs free energy at fixed . Differentiating givesEquality of both component chemical potentials between two compositions becomes the common-tangent construction for phase coexistence:In this molar representation, the tangent slope is and its intercept is . Stable coexistence uses a supporting tangent lying below the homogeneous curve over all admissible compositions. This constructs the relevant convex envelope; two stationary points or equal slopes alone do not establish stability.
An original regular solution calculation makes this distinction visible. Because linear composition bias leaves coexistence compositions unchanged, subtract the linear pure-component reference and writeThe dimensionless parameter penalizes unlike contacts when it is positive. Take . Symmetry makes the coexistence tangent horizontal, and the two outer solutions of give the symmetric regular-solution binodal:These are model compositions, rather than measured hexane–water values. At overall mole fraction , the liquid-liquid tie line and lever rule give the fraction of total moles in phase :Changing within the coexistence interval changes phase amounts while leaving the two endpoint compositions fixed. These mole-based phase fractions need not equal volume fractions.
The spinodal answers a different question: local stability of a homogeneous mixture. Here , so at the two local-stability limits are . Between them, composition fluctuations lower the bulk free energy. Between a binodal endpoint and the nearer spinodal, the homogeneous state has positive local curvature but a lower-energy phase-separated state exists: this is metastability. Thus the observed coexistence compositions are not obtained merely by setting .
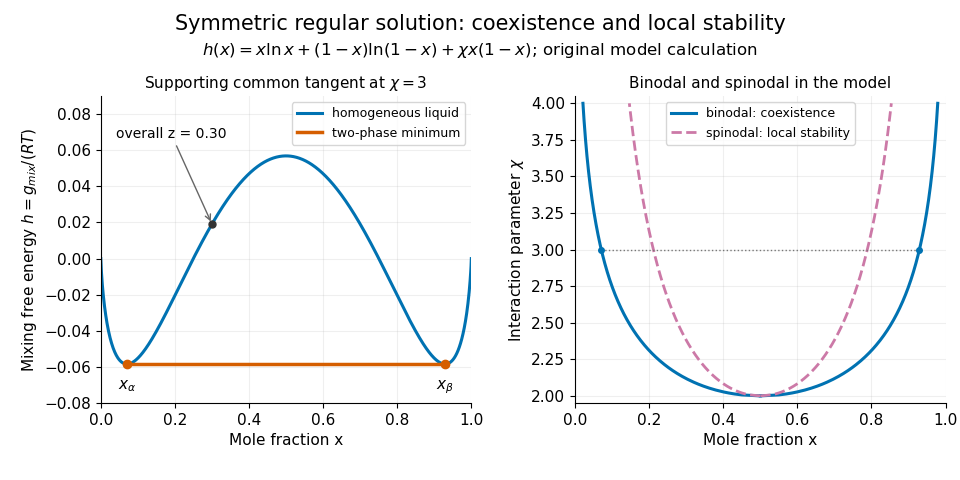
Original regular-solution model: the supporting tangent gives liquid coexistence, while the spinodal marks loss of local homogeneous stability.
Details. A useful experiment measures compositions as well as the presence of layers. One hexane–water record reports overnight shaking, microsyringe sampling of the aqueous solution, and gas chromatography to measure dissolved hydrocarbon. The small dissolved amount is evidence of finite mutual solubility, consistent with liquid immiscibility over other bulk compositions.[ref]
Hexane with water in the IUPAC-NIST Solubility Database. Record37_269, Experimental Data and Method/Apparatus/Procedure; no ambiguous table values are used here
At controlled temperature, repeated sampling of both phases and checks from different initial compositions help test whether the endpoint compositions have stabilized. Critically evaluated reference systems provide a way to validate such instruments and techniques.[ref]Reference Materials for Phase Equilibrium Studies. 1. Liquid-Liquid Equilibria by Ala Bazyleva and others. Abstract and Introduction: seven binary reference systems, accepted data and uncertainty estimates
For exchanging neutral components at a common temperature and pressure, activity equality for coexisting liquids expresses liquid-liquid equilibrium as , provided both thermodynamic activities use the same standard state. The basic equilibrium condition is equality of chemical potentials; equality of numerical activities follows only after their references have been matched.
Write the definition of thermodynamic activity aswhere is the molar gas constant and is temperature. Subtraction at equilibrium givesThe ratio is one when the two standard chemical potentials are the same. IUPAC defines activity relative to the specified standard chemical potential and distinguishes pure-phase and solute reference conventions.[ref]
Quantities, Units and Symbols in Physical Chemistry by IUPAC. §2.11, pp.57,59; §2.11.1(iv), pp.61–62.
Mixing conventions can therefore produce unequal activities for physically coexisting phases. For instance, if one reference exceeds the other by , its corresponding activity is half as large at the same chemical potential. Standard-state activity conversion changes the numerical coordinate, not the physical driving force.Equal thermodynamic activities also need not mean equal mole fractions. With a common pure-liquid reference, , where is an activity coefficient. An original symmetric regular solution example makes this quantitative. Set the dimensionless interaction parameter to . The model has coexisting component-A mole fractions and , withThus and , whileComponent B satisfies the analogous equality by symmetry. The different compositions are compensated by different nonideal environments. These are chosen model values, not experimental compositions. As in liquid-liquid equilibrium, satisfying equalities must be supplemented by stability against competing phase arrangements.
Standard-state activity conversion changes the numerical thermodynamic activity assigned to a fixed physical state when its reference chemical potential changes. The observable equilibrium and physical chemical potential stay the same. The activity and its standard chemical potential must therefore be converted together.
At fixed temperature , writeIf , subtraction givesHere is the molar gas constant. This follows from the IUPAC definition of relative activity and its specified standard state.[ref]
Quantities, Units and Symbols in Physical Chemistry by IUPAC. §2.11, pp.57,59, and §2.11.1(iv), pp.61–62: activity and reference conventions.
Changing just the activity in the first equation would instead change the calculated physical chemical potential.For an invented conversion , the compensating reference shift is . An old activity becomes ; the two expressions for are identical. An activity of one consequently identifies a reference condition under the chosen convention, not an invariant amount of material.
An activity conversion of an equilibrium constant follows the same logic. For the reaction A B, take at equilibrium and convert by , . Then . If , the converted value is , whilebecause originally . For any physical composition, the corresponding reaction quotient also changes by , so remains unchanged. These are reference changes, not altered reaction chemistry.
The same requirement governs activity equality for coexisting liquids: compare activities only after identifying their standard states, composition scales and reference conventions. Equal numerical values expressed under incompatible conventions do not, by themselves, establish liquid-liquid equilibrium.
Activity conversion of an equilibrium constant changes the numerical equilibrium constant when a different standard state convention rescales the thermodynamic activities used to define it. The equilibrium composition is unchanged. The essential requirement is to convert the reaction quotient, the equilibrium constant and the standard Gibbs energy of reaction together.[ref]
Quantities, Units and Symbols in Physical Chemistry by IUPAC. §2.11, printed pp. 57–58: relative activity, reaction quotient, standard reaction Gibbs energy and standard equilibrium constant; notes 12, 13 and 18
At fixed temperature , write a chemical reaction as , with positive stoichiometric coefficients for products and negative ones for reactants. Let a standard-state activity conversion give , where each is positive and independent of composition at the chosen conditions. From the product defining the reaction quotient,At chemical equilibrium, and , soA scale change for a reactant enters with a negative exponent; multiplying every activity by the same number need not leave unchanged when the sum of stoichiometric coefficients is nonzero.
The reference chemical potentials must compensate. Since , the converted standard values are , where is the molar gas constant. ConsequentlyThe physical Gibbs energy of reaction and the ratio are invariant. A change in the numerical equilibrium constant alone therefore does not show that the reaction has become more favorable.
For an original example, consider , with , and choose , . Then . A composition with has ; both conventions give and . The original standard Gibbs energy of reaction is , whereas the converted value is . Their opposite signs identify different reference conventions, while the actual reaction driving Gibbs energy at the chosen composition agrees exactly. This calculation assumes a reference rescaling, rather than a composition-dependent alteration of the physical activity model.
A liquid-liquid tie line joins the compositions of two liquid phases that coexist at a specified temperature and pressure. Its endpoints describe the separate phases; an interior point describes an overall composition formed from suitable amounts of them. The liquid-liquid tie line therefore connects equilibrium chemistry with material balance rather than describing a spatial boundary between layers.
For a binary mixture, use component-B mole fractions , and overall fraction . If is the mole-based phase fraction in phase , conservation of component B gives the lever rule:The endpoints must be different and . For invented coexistence endpoints , and overall , the result is . In a -mole sample, the phases contain and moles; their component-B amounts are and moles, totaling . These mole amounts do not specify the two layer volumes.
A ternary liquid-liquid tie line obeys the same vector balance. For chosen endpoint vectors and , a phase fraction gives . Every component must give the same . A normalized composition vector need not lie on this particular line; normalization alone does not establish compatibility with the proposed endpoints.
The lever rule determines amounts after equilibrium endpoints have been established; it does not select those endpoints. They can come from measured phase compositions or a stable common-tangent construction for phase coexistence. Reliable experiments control temperature and sample purity, measure the separated compositions and distinguish genuine liquid-liquid equilibrium from other phase behavior. An IUPAC reference-system assessment specifically warns that turbidity can confuse liquid–liquid and solid–liquid transitions.[ref]
Reference Materials for Phase Equilibrium Studies. 1. Liquid-Liquid Equilibria by Ala Bazyleva and others. Introduction, pp.1–2: composition measurement, emulsions, solid–liquid discrimination and reference-system validation.
A ternary liquid-liquid tie line joins the compositions of two coexisting liquid phases in a three-component mixture at a specified temperature and pressure. IUPAC defines a tie line by its coexisting equilibrium endpoints.[ref]
Tie-line in IUPAC Gold Book. Entry15367: line connecting compositions of independent coexisting phases.
Each composition is a vector of mole fractions satisfying . Only two coordinates are independent; a triangular composition diagram displays this constraint.If is the mole-based phase fraction in phase , conserving every component givesThis is the lever rule in vector form. Wherever , each component must give the same valueA component identical at both endpoints cannot determine , but its overall fraction must equal that common endpoint value.
For an original material-balance example, stipulate endpoint compositions and . With , the overall composition is . All three component calculations give . By contrast, still sums to one, but gives apparent phase fractions , and . Under the stipulated exact endpoints, it does not lie on this tie line. A ternary tie-line consistency residual records the failure of the vector balance; experimental use must account for composition uncertainties.
These chosen endpoints illustrate conservation, not a claim that an actual mixture has these equilibrium compositions. Establish endpoints from equilibrium measurements or a stable Gibbs free energy model. Moving along one liquid-liquid tie line changes phase amounts while preserving its endpoint compositions. Moving elsewhere in the two-phase region can select a different pair of endpoints; all tie lines need not be parallel. Finally, mole-based and mass-based diagrams require consistent fractions: a mole-based is not automatically a volume or mass fraction.
A ternary tie-line consistency residual quantifies how far an overall three-component composition is from the mass balance predicted by specified endpoints of a ternary liquid-liquid tie line. It is a geometric consistency check on a proposed two-phase decomposition. A thermodynamic tie line connects coexisting equilibrium compositions; the calculation below checks the lever rule for those endpoints, rather than independently proving that they are equilibrium phases.[ref]
Tie-line in IUPAC Gold Book. Entry 15367: tie line connects compositions of independent coexisting equilibrium phases
Use mole fraction vectors for the endpoint liquids and for the overall composition, all normalized to sum to one. Put . For a proposed mole-based phase fraction , defineExact mass balance requires every component of to vanish. Closure gives automatically, so this residual has only two independent components. A zero sum alone does not imply that the component balances hold.
When the phase fraction is unknown and the endpoints are distinct, an unweighted least squares tie-line phase-fraction fit minimizes the squared Euclidean norm of . Differentiating givesThis is an orthogonal projection onto the line through the endpoints. A physically admissible phase fraction also lies in ; if the unconstrained fit lies outside that interval, the closest admissible composition is an endpoint of the line segment. Coincident endpoints do not determine a phase fraction through this formula.
For the hypothetical endpoints and , the composition gives and zero residual. Changing the overall composition to preserves closure but givesNo adjustment of a single phase fraction can eliminate that discrepancy under the stipulated exact endpoints. For experimental use, its significance depends on measurement uncertainty in both the overall composition and endpoint compositions. Unweighted distance is a chosen metric, not an automatic statistical rejection threshold; correlated composition errors require their covariance matrix on the independent composition coordinates. Even a tiny residual establishes only compatibility with this mass balance, not equality of chemical potentials or stability of the proposed liquid-liquid equilibrium.
A tie-line phase-fraction fit estimates how much of each liquid phase is needed to reproduce an overall composition from specified liquid-liquid tie line endpoints. If are endpoint mole fraction vectors and is the mole-based phase fraction of phase , the mass balance predictsThe endpoints must be supplied by equilibrium measurements or a suitable thermodynamic model; fitting does not establish that an arbitrary pair of compositions is an equilibrium tie line.[ref]
Tie-line in IUPAC Gold Book. Entry15367: endpoints are compositions of coexisting equilibrium phases
For a ternary liquid-liquid tie line, use two independent composition coordinates, such as the first two mole fractions, since the third is fixed by their sum. Let be these two measured coordinates, the corresponding phase- coordinates, the difference between phase- and phase- coordinates, and a positive-definite covariance matrix for . With exact endpoints, generalized least squares minimizesDifferentiation yields the unconstrained estimateThe second expression imposes the physical bounds. It follows because is a convex quadratic when the endpoints differ in the measured coordinates. The ternary tie-line consistency residual then records the remaining failure of the mass balance.
For an original hypothetical example, take endpoint compositions and , with overall measurement . Suppose the first two measured mole fractions have independent standard measurement uncertainties and , so . ThenThe predicted composition is , giving residual . The separate first- and second-component balances would suggest and ; the fit combines them according to their stated measurement uncertainties, rather than averaging those two numbers without regard to precision.
With Gaussian composition errors, fixed endpoints and an interior unconstrained fit, the tie-line phase-fraction uncertainty follows directly from the linear estimator:for this example. Endpoint uncertainty, a fit on a boundary, or a nearly vanishing endpoint difference needs additional treatment. In particular, all three normalized mole fractions must not be treated as independent observations with an invertible three-coordinate covariance matrix: their exact closure creates dependence. A small fitted residual is evidence of consistency with the specified mass balance and error model, not a substitute for testing chemical potential equality or phase stability.
Tie-line phase-fraction uncertainty describes the uncertainty of the phase amount inferred from a tie-line phase-fraction fit. It depends on the uncertainties of the overall composition, the endpoint compositions and the model used to fit them. A conditional estimate treating endpoints as exact omits their contribution. All composition vectors below use two independent coordinates of a ternary liquid-liquid tie line; the third mole fraction is fixed by closure.
For fixed endpoints , let and let the overall composition measurement have covariance matrix . With and an interior unconstrained fit,Here denotes standard uncertainty. The variance formula is exact for this linear estimator with fixed endpoints and known covariance; Gaussian errors are needed for a Gaussian sampling distribution, not for the variance identity itself. This is an instance of covariance-based uncertainty propagation.[ref]
Guide to the expression of uncertainty in measurement by JCGM. §§5.1–5.2, sensitivity coefficients, combined variance and correlated input quantities
Endpoint errors enter through tie-line phase-fraction sensitivity. Near an exactly balanced composition , with the weight matrix held fixed, differentiating the fit givesIf overall and endpoint measurements are mutually independent, their combined first-order contribution isShared instruments or common reference data can correlate these measurements; then cross-covariances, including tie-line endpoint covariance, must also be propagated rather than discarded.[ref]
Guide to the expression of uncertainty in measurement by JCGM. §5.2, equation(13), and §§5.2.4–5.2.5, significant covariance from common influences
For an original example, use endpoints and with overall composition , so . Let the first two overall mole fractions have independent standard uncertainties and . Then and , giving with exact endpoints. If each endpoint instead has independent coordinate uncertainties , with all three measurements independent, the endpoint contribution isThe combined value becomes .
When the endpoints approach one another, their difference becomes less informative about phase fraction; at coincident endpoints this inference is undefined. Near the physical bounds, clipping an unconstrained fit to the interval from zero to one changes its sampling distribution, potentially creating a point mass at an endpoint. A symmetric Gaussian error summary can then be misleading. Propagating a joint input model through the actual bounded fit with a Monte Carlo method is one possible analysis; it must retain composition closure and the specified correlations. Neither a small uncertainty nor a small ternary tie-line consistency residual independently verifies chemical potential equality or phase stability.
Tie-line phase-fraction sensitivity measures how an inferred phase fraction changes when the overall composition or the measured liquid-liquid tie line endpoints change. A sensitivity coefficient is a local partial derivative of the inference formula, rather than an uncertainty by itself. It becomes an uncertainty contribution only after combination with the input covariance matrix.[ref]
Guide to the expression of uncertainty in measurement by JCGM. §§5.1.3–5.1.4, sensitivity coefficients and their experimental determination
For one composition coordinate of a binary mixture, write the lever rule aswhere are endpoint mole fractions, is the overall mole fraction and is the mole-based amount of phase . Differentiating the mass balance givesThus a measurement error in the overall composition and an equal measurement error in the phase- endpoint have different effects. The conditioning of tie-line phase-fraction inference deteriorates as the endpoint separation becomes small: the same composition measurement error then produces a larger change in the inferred phase fraction.
For an original example, take , , , giving . Increasing only by gives a first-order change . Direct substitution gives , a change of . For the closer endpoints , and balanced composition , the local sensitivity to the same endpoint error is ten times larger. This is a property of the inference geometry; it does not establish whether either hypothetical pair is in liquid-liquid equilibrium.
The multidimensional tie-line phase-fraction fit has an additional detail away from an exactly balanced composition. On independent mole fraction coordinates, hold a symmetric positive-definite matrix fixed and definewith distinct endpoints. Define row vectors and . Differentiating the generalized least squares normal equation givesThe residual correction in tie-line sensitivity vanishes when . Otherwise, moving an endpoint also rotates the fitted line relative to the nonzero ternary tie-line consistency residual. Omitting that contribution is an additional approximation. If itself changes with the measured inputs, its derivative must also enter the calculation.
These formulas describe the unconstrained fit locally. Imposing introduces boundary effects, and coincident endpoints make this inference undefined. A practical sensitivity check recomputes the fit after small positive and negative perturbations of one measured coordinate, keeping composition closure and the stated weight matrix. Agreement as the perturbation shrinks tests the derivative calculation; it does not validate the underlying equilibrium model.
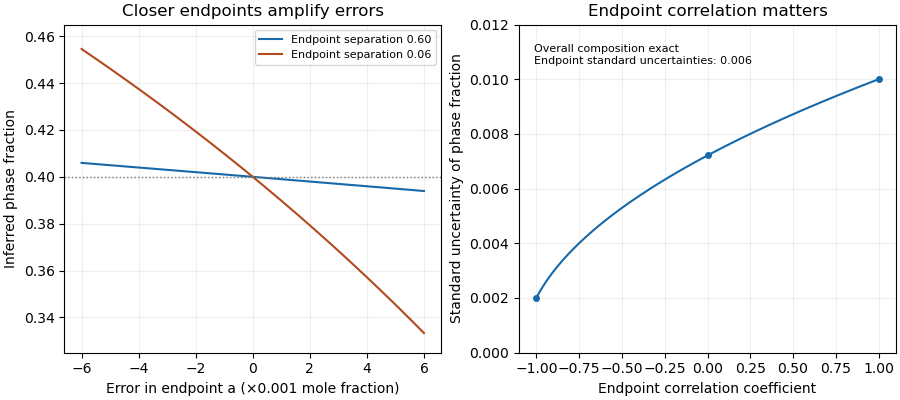
Closer tie-line endpoints amplify composition errors; endpoint correlation changes the inferred phase-fraction uncertainty
. Original binary examples with phase fraction0.40. Left: exact endpoint perturbations at separations0.60 and0.06. Right: first-order standard uncertainty with uncertain endpoints and exact overall composition; common calibration of all three measurements is a different model. Details.The residual correction in tie-line sensitivity accounts for rotation of a fitted liquid-liquid tie line when the measured overall composition does not lie exactly on it. In independent mole fraction coordinates, set , hold a symmetric positive-definite matrix fixed, and use the interior unconstrained tie-line phase-fraction fit. With , residual , and row vectors , , its differential isThe full derivation is in tie-line phase-fraction sensitivity. The normal equation makes orthogonal to in the chosen metric; it does not make the residual zero.
For an original example, take , , and . The third mole fraction is fixed by closure. These choices giveIncreasing only the first coordinate of by predicts a phase fraction change . Omitting the correction misses the second contribution even though the fit satisfies its normal equation. A centered finite-difference recalculation as the perturbation shrinks tests this derivative directly.
This is a local sensitivity, not an uncertainty until combined with the joint input covariance matrix.[ref]
Guide to the expression of uncertainty in measurement by JCGM. §§5.1.3–5.1.4, sensitivity coefficients; §5.2.2, correlated-input propagation
The formula assumes distinct endpoints and an interior fit with fixed . A changing metric requires sensitivity of a tie-line fit to its weight matrix; a constrained boundary requires a separate local analysis. A nonzero residual can reflect composition noise or model mismatch and does not independently establish liquid-liquid equilibrium.Sensitivity of a tie-line fit to its weight matrix describes how changing the metric changes an inferred phase fraction, even when the measured compositions are held fixed. For distinct endpoints , define , and the interior unconstrained tie-line phase-fraction fitDifferentiating the normal equation , with fixed, givesThis term supplements the composition and endpoint derivatives in tie-line phase-fraction sensitivity; it is not included in their fixed-metric formulas.
For an original example, use independent mole fraction coordinates , and . With , the fitted fraction is , and . Increasing the second diagonal weight from to predictsDirect refitting gives , a change of . The small difference is the neglected higher-order term.
Two invariances clarify the result. An exactly balanced composition has and the same fraction under every positive-definite metric. Also, replacing by for positive leaves the fraction unchanged: the scalar rescaling invariance of a tie-line weight matrix follows from cancellation in its ratio. The differential agrees because .
If for a composition covariance matrix, its perturbation is to first order. Uncertain fitted weights can therefore add sensitivity, with correlations to the composition measurements retained in the measurement model. This is an application of the guide's general sensitivity-coefficient framework, not a tie-line formula quoted from it.[ref]
Guide to the expression of uncertainty in measurement by JCGM. §5.1.3, derivatives of the measurement model; §5.2.2, correlated inputs
Boundary-constrained fits and coincident endpoints need separate treatment.Scalar rescaling invariance of a tie-line weight matrix means that multiplying every fit weight by the same positive number leaves the fitted phase fraction unchanged. Let endpoint compositions and overall composition be expressed in independent mole fraction coordinates. With , a fixed symmetric positive-definite matrix defines the tie-line phase-fraction fit by minimizingFor , . Multiplication by a positive constant preserves every comparison between objective values, so the sets of minimizers agree. The proof also applies when the same constraint is imposed in both fits.
For distinct endpoints, the unconstrained formula makes the cancellation visible:The differential in sensitivity of a tie-line fit to its weight matrix agrees: for a perturbation , its numerator is by the fit's normal equation. Thus the scalar direction is insensitive even when the residual is nonzero.
In an original example, take , , and . The remaining composition coordinate is fixed by closure. These values giveReplacing by gives the same fraction, although the minimum objective increases from0.00072to0.00504. Changing only the second diagonal weight from4to7 instead changes the fitted fraction to0.640625. Relative weights determine the geometry; their common multiplier does not.
Point-estimate invariance does not by itself establish invariance of physical measurement uncertainty. Merely rewriting the same objective with leaves propagation through the same measured-input covariance matrix unchanged. If instead represents absolute precision and asserts a genuinely different covariance , that is a different noise model. With exact endpoints, the linear-model variance then decreases by a factor . The measurement model and covariance interpretation must be stated.[ref]
Guide to the expression of uncertainty in measurement by JCGM. Sections 5.1.3 and 5.2.2, model sensitivities and covariance propagation; tie-line result derived here
The conditioning of tie-line phase-fraction inference describes how strongly composition perturbations change an inferred phase fraction. For a binary mixture with distinct endpoints, the lever rule gives . Its absolute sensitivity to , with endpoints fixed, is . This is an absolute amplification coefficient; a relative condition number additionally depends on the chosen input and output scales.
For an interior , the differential in tie-line phase-fraction sensitivity yieldsto first order. If each composition perturbation is bounded by , this becomes . For original stipulated , endpoint separations and give local worst-case bounds and . Tenfold closer endpoints make the same composition precision tenfold less informative about phase amount. Correlations can change the attainable combinations of errors, as common affine calibration cancellation in the lever rule illustrates.
For fixed multicomponent endpoints and overall-composition covariance matrix , the tie-line phase-fraction uncertainty isThe denominator is endpoint separation after a whitening transformation: separation must be compared with measurement precision, not only plotted distance. This follows from covariance-based uncertainty propagation and the linear fit.[ref]
Guide to the expression of uncertainty in measurement by JCGM. §§5.1–5.2, sensitivity coefficients and covariance propagation
Near coincident endpoints the inverse problem becomes poorly informative, and at coincidence it is undefined. Near the physical bounds, a large propagated uncertainty also warns that the interior linear approximation may fail. More numerical digits in the fitted answer do not resolve this measurement limitation.
Tie-line endpoint covariance records how the measurement errors of the two measured liquid-liquid tie line endpoints vary together. Using independent mole fraction coordinates, represent the two endpoint estimates by column vectors . Their cross-covariance matrix isAn individual off-diagonal block need not be symmetric; the full joint covariance matrix must be a symmetric positive semidefinite matrix. The uncertainties of the two endpoint vectors alone do not determine their cross-covariance matrix.
Near an exactly balanced, interior tie-line phase-fraction fit, let be the row-vector sensitivity coefficient defined in tie-line phase-fraction sensitivity. If the overall composition measurement is independent of both endpoints, first-order uncertainty propagation givesThe cross term has a positive coefficient because the two endpoint perturbations enter the phase fraction with the same sign. Correlated endpoints can therefore increase the result. The general rule is to propagate the full joint covariance matrix, rather than assume that correlation always increases or always reduces uncertainty.[ref]
Guide to the expression of uncertainty in measurement by JCGM. §5.2.2, equations13–16, correlated inputs
For a stipulated binary mixture example, use , , , so . Treat as exact and give each endpoint standard uncertainty with correlation coefficient . The lever rule yieldsThis gives for , about for , and for . These are first-order standard uncertainties under the stipulated joint error model, not bounds on all possible errors.
A shared calibration error affecting the overall composition as well changes the calculation. Suppose the same additive error affects identically. ThenThe shared offset cancels exactly. More generally, a common affine map with also cancels in this binary ratio. This common affine calibration cancellation in the lever rule concerns only a truly common contribution; separate sample errors remain. If the shared offset has standard uncertainty , incorrectly treating its three appearances as independent would assign a spurious contribution instead of zero.
In general, stack the inputs as with joint covariance matrix . Let be the corresponding Jacobian matrix row from tie-line phase-fraction sensitivity, including its residual correction when needed. Then automatically includes correlations with the overall composition. The measurement guide specifically identifies common instruments, measurement standards and reference data as sources of such correlations.[ref]
An instructive measurement design analyzes the two equilibrium phases and the overall mixture using one instrument calibration, then repeats the entire procedure with independent instrument calibrations. The within-run contrasts and between-run changes probe different error contributions. Repeated readings under one unchanged instrument calibration cannot by themselves establish the uncertainty of its shared offset. The measurement model must identify which influences are common before the joint covariance matrix can represent them.
Common affine calibration cancellation in the lever rule is the exact invariance of the binary mixture phase fraction under the same affine map applied to the overall composition and both endpoints. For ,Both a shared offset and a shared scale cancel. The map must be common to all three measurements; separately fitted calibration curves or sample-dependent effects do not satisfy that assumption.
An original example has , , and . Applying gives measured values , and , still yielding . If only the overall-composition reading then receives an additional error, its inferred fraction changes by . Common calibration cancellation leaves independent sample errors intact.
The covariance matrix formulation explains why treating the shared errors as independent gives a false uncertainty. In input order , the Jacobian matrix row isIt annihilates both the common-offset direction and the common-scale direction , since . A joint covariance matrix representing only these shared affine influences therefore gives . The measurement guide explicitly identifies common instruments and reference data as sources of correlation that must be retained.[ref]
Guide to the expression of uncertainty in measurement by JCGM. §5.2.2, equation13; §5.2.4, common influences
Analyzing the overall sample and separated phases under one unchanged instrument calibration can realize this common contribution; repeating only one sample under another calibration breaks it. The cancellation is a property of this binary ratio. It does not remove nonlinear response, composition closure errors, endpoint misidentification or uncertainty in whether the samples establish liquid-liquid equilibrium.
Mechanical equilibrium requires force balance for the mechanical displacements that are allowed. Two uniform fluids separated by a freely moving, unloaded, frictionless piston with equal face areas require equal pressures. A supported piston can sustain a pressure difference, and a fluid in gravity instead obeys hydrostatic equilibrium. For an upward height coordinate , that balance is , where is mass density and is the downward gravitational acceleration. Mechanical equilibrium alone does not establish thermal equilibrium or chemical equilibrium.
Two fluids at a stationary, freely movable interface require equal pressures when surface tension and other normal stresses are negligible. This local pressure equilibrium does not require equal temperatures or mass densities. Curvature with nonzero surface tension or an applied load changes the normal-stress balance. Comparing points at different heights also requires the hydrostatic equilibrium condition.
Thermodynamic stability means that allowed small perturbations do not lower the appropriate thermodynamic potential from an equilibrium state. The constraints select that potential: an isolated system maximizes entropy, a system at fixed temperature and volume minimizes Helmholtz free energy, and one at fixed temperature and pressure minimizes Gibbs free energy.
For an unconstrained scalar order parameter , a stationary state has , and positive curvature gives local stability. A distant lower minimum can still make it metastable rather than globally stable. In an ordinary additive fluid, positive isothermal compressibility expresses resistance to density perturbations: increasing pressure should reduce volume. Loss of the corresponding curvature can mark a spinodal. These statements depend on the ensemble and permitted perturbations; mechanical equilibrium alone does not establish thermal, chemical or global stability.
For a mechanical coordinate at fixed temperature, total volume and particle numbers, the relevant thermodynamic potential is the Helmholtz free energy . Its generalized force is . At mechanical equilibrium, , and a linearly restoring force has , equivalently . A Taylor expansion then proves a strict local free-energy minimum. If is a strictly convex function throughout the accessible domain, the stationary state is the unique global minimum.
A closed system exchanges no matter with its surroundings, but can exchange energy through heat or work. A sealed cylinder with a moving piston is a typical idealized example. “Closed” fixes the material inventory, rather than the volume or the internal energy. An insulated cylinder can receive compression work; a rigid sealed vessel can receive heat. Neither is necessarily an isolated system.
An isolated system exchanges neither matter nor energy with its surroundings. Its total energy is fixed, while the Second law of thermodynamics requires its total entropy to be nondecreasing. Its internal energy alone is fixed when bulk kinetic energy and external potential energy also remain unchanged. For example, an initially spinning stirrer inside an insulated rigid vessel can lose kinetic energy while the contents gain internal energy, with no external input.
An isolated system need not yet be in thermodynamic equilibrium. Two enclosed bodies at different temperatures can exchange heat internally until they reach thermal equilibrium; their combined energy stays constant. Their internal transfer is absent from the outer boundary ledger. A vacuum-insulated container is only an approximation to isolation: residual thermal conduction and thermal radiation can still carry energy across its boundary.
A quasistatic process is an ideal limit in which a changing thermodynamic system passes through a sequence of states arbitrarily close to thermodynamic equilibrium. Its intermediate pressure, temperature and volume are therefore well defined and obey its equation of state. Driving must be slow compared with the system's own equilibration, rather than merely taking a long time on a clock. A reversible thermodynamic process additionally allows both system and surroundings to be restored without net change; friction can produce entropy even in a quasistatic process.[ref]
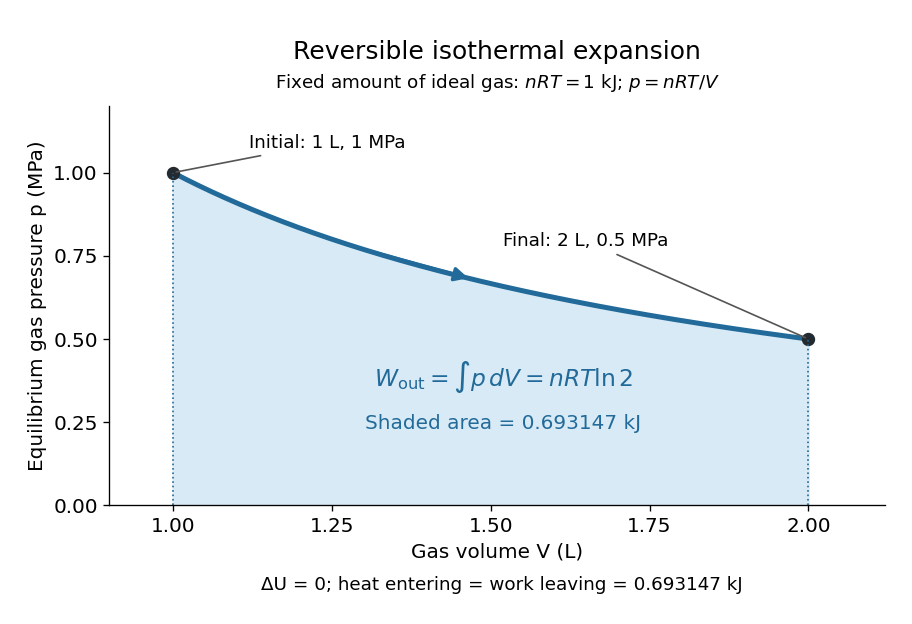
For a computed example, take a fixed amount of substance of ideal gas in a sealed cylinder with a frictionless piston, in contact with a heat bath at absolute temperature . Reduce the external load in infinitesimal steps so that the piston expands from to . In the reversible limit, the gas pressure exceeds the external pressure by an infinitesimal amount during expansion, and the heat transfer occurs across an infinitesimal temperature difference. Exact mechanical equilibrium would give no driving force; the limiting imbalance supplies motion while preserving equilibrium intermediate states.[ref]
Along this reversible isothermal expansion, the ideal gas law givesHere is the molar gas constant; specifying fixes an energy scale, without choosing and separately. The initial and final equilibrium states are and . Count work delivered to the surroundings as positive. The reversible work is the shaded area beneath the pressure-volume curve:The integral of supplies the natural logarithm. Since internal energy of an ideal gas depends only on temperature, ; the first law of thermodynamics, , gives to the displayed precision. The heat bath replaces the energy delivered as work.[ref]
Now start from the same initial state, but abruptly lower the external pressure to the constant value . There is a finite initial pressure imbalance, so this is an irreversible expansion rather than a quasistatic process. Assume the piston begins and ends at rest, has no net change in stored energy, and has no external friction; relaxation occurs within the gas and its heat bath. After motion has settled and the heat bath has restored temperature , mechanical equilibrium and the ideal gas law give . The work delivered to the constant load isThis work follows directly from load force times piston displacement, using for piston area . Intermediate gas states need not have a single equilibrium pressure or temperature. In particular, neither at the bath temperature nor describes the actual gas during this irreversible motion. The rectangle below plots the known external load; its upper edge is not an equilibrium gas path.
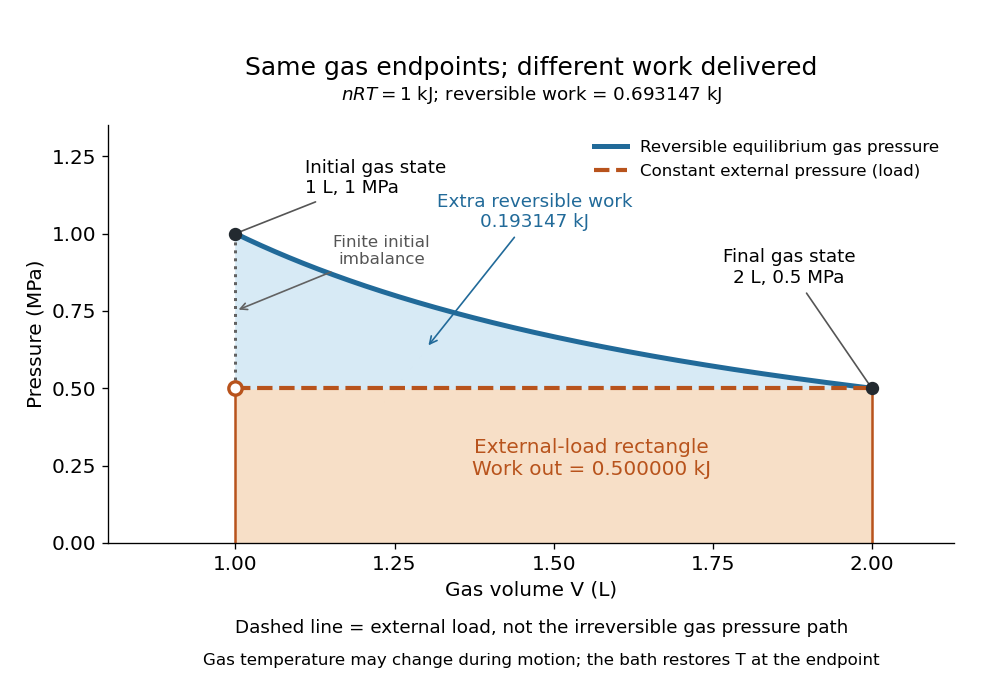
The same equilibrium endpoints give different work for reversible and constant-load expansion
. The blue curve is the reversible ideal gas path. The dashed orange line is constant external pressure, whose rectangle gives of load work. The additional blue shading is available in the reversible case. Black points mark the common gas endpoints; the open orange point marks the initial external load, and the dotted gap indicates the initial imbalance. No irreversible internal gas path is drawn. Details.The entropy accounting makes the difference precise. Because entropy is a state function, both expansions have the same gas entropy change, computed along the reversible isothermal expansion:The thermal reservoir stays at and loses heat , so . Assume the ideal load stores work without an entropy change. The combined entropy production is thereforeNo numerical entropy unit is implied by alone: or must also be specified. The lost work opportunity is for this comparison. This calculation uses equilibrium endpoints and the reservoir's temperature, without assigning equilibrium intermediate states to the irreversible gas. [ref]
https://openstax.org/books/university-physics-volume-2/pages/4-6-entropy. OpenStax, §4.6: entropy as a state function and reversible reference paths
For another boundary choice, let the same ideal gas expand into an initially evacuated compartment of volume inside a rigid, insulated vessel of total volume . Removing the partition causes free expansion: no load is moved and no heat crosses the outer boundary, so and . After equilibration the temperature is again , the pressure is , and the gas entropy has increased by . The outer vessel has fixed volume; the gas's accessible volume doubles. Restoring the initial gas state requires compression and removal of heat, changing the surroundings. Equal endpoints do not make free expansion reversible. [ref]
https://openstax.org/books/university-physics-volume-2/pages/4-1-reversible-and-irreversible-processes. OpenStax, §4.1: insulated free expansion and restoration
A slowly moving piston with dry friction supplies a different caution: its gas can remain arbitrarily close to thermodynamic equilibrium, yet rubbing still converts work into heat and produces entropy. If the frictional force has a nonzero slow-motion limit, its dissipated energy over a fixed travel does not vanish with speed. Reversing the motion reverses friction, so infinitesimally changing the external load cannot retrace the same operation and restore the surroundings. The heat capacity calculations additionally exclude friction and other dissipative work.[ref]
The usefulness of a quasistatic process is that it replaces detailed motion by a readable sequence of equilibrium states. An equation of state and an integral then turn a pressure-volume drawing into an energy budget. The beautiful distinction is that internal energy and entropy depend on the endpoints, while heat and work record how the change was performed. Reversible benchmarks expose how much work an irreversible device sacrifices, while entropy production explains why simply making it slower need not recover that opportunity.
A state function assigns a value to each equilibrium thermodynamic state of a specified thermodynamic system. Its change depends only on the initial and final states. Internal energy , entropy and enthalpy are state functions; here is pressure and is volume. Transferred heat and work belong to a thermodynamic process, rather than to a state. A system stores internal energy; it does not store separate inventories of “heat received” and “work received”.[ref]
Thermodynamics and Propulsion. Thermodynamics and Propulsion, §2.1, points 3–5 and equation 2.2: internal energy, transfer notation and signs
Use the explicit convention for heat entering and for work done by the thermodynamic system on its surroundings. For a closed system with unchanged endpoint bulk kinetic energy and external potential energy, the first law of thermodynamics readsThe notation denotes an exact differential of a state function; and denote incremental transfers. Along a reversible thermodynamic process of a simple compressible system,where is absolute temperature. These are rules for particular thermodynamic processes, not definitions of stored heat or stored work. In particular, a quasistatic process with friction can remain near thermodynamic equilibrium while producing entropy.[ref]
Thermodynamics and Propulsion. Thermodynamics and Propulsion, §5.3: reversible transfer formulas distinguished from relations between state properties
For any state function and two equilibrium states , an integral of its exact differential givesThe proof is telescoping: add the changes along a subdivided path, and all intermediate values cancel. The smooth limit gives the integral. Conversely, if an integrable differential form has zero integral around every accessible closed path in a connected region, define by integrating from a reference state. Two routes differ by a closed path, so the definition is unambiguous. This argument needs every accessible loop, rather than a local test that might miss a hole in the domain; see path independence.
The same cancellation does not hold separately for heat and work. A thermodynamic cycle has but can have nonzero net transfers, with . Nor does the fact that entropy is a state function make its general change: the formula uses a reversible thermodynamic process and its varying temperature.
This distinction makes endpoint calculations powerful. We can compute an internal energy, entropy or enthalpy difference along a convenient reversible reference path even when the actual thermodynamic process is irreversible. Its actual heat and work must still be obtained from its own boundary interactions. The appealing cancellation is visible in two reversible ideal-gas paths between the same states: a different mechanical output requires a matching difference in thermal input, while stored internal energy arrives at the same value.
Take a closed system consisting of one mole of a classical monatomic gas, idealized as an ideal gas with constant heat capacity at constant volume . Stipulate amount of substance and molar gas constant for this calculation. Set initial temperature and volume , and define and . The specified equilibrium endpoints areThus has volume , temperature and pressure . The fixed material inventory and ideal gas law make these complete endpoint specifications for this model. All numbers below are calculated values, rather than measurements of a particular gas.
Use a frictionless piston, ignore bulk mechanical energy changes, and allow only pressure-volume work. Each step is the limit of a reversible thermodynamic process: adjust loads by infinitesimal pressure differences, and supply heat through infinitesimal temperature differences. Heating with changing temperature requires a succession of matching thermal reservoirs; contact with a single hotter heat bath would give an irreversible finite difference. On a fixed-volume leg the piston is locked. These assumptions justify using the actual equilibrium path's pressure inAn endpoint chord is not the path; substituting a guessed straight line would compute a different thermodynamic process.[ref]
Thermodynamics and Propulsion. Thermodynamics and Propulsion, §1.3.3, Figures 1.10–1.11: external pressure versus equilibrium gas pressure in boundary work
The upper route heats at fixed volume from to , then undergoes isothermal expansion at to . The lower route undergoes isothermal expansion at from to , then heats at fixed volume to . The tuples use the same ordering as the endpoints. Vertical legs do zero expansion work, and the two curved legs have different pressures at every shared intermediate volume.
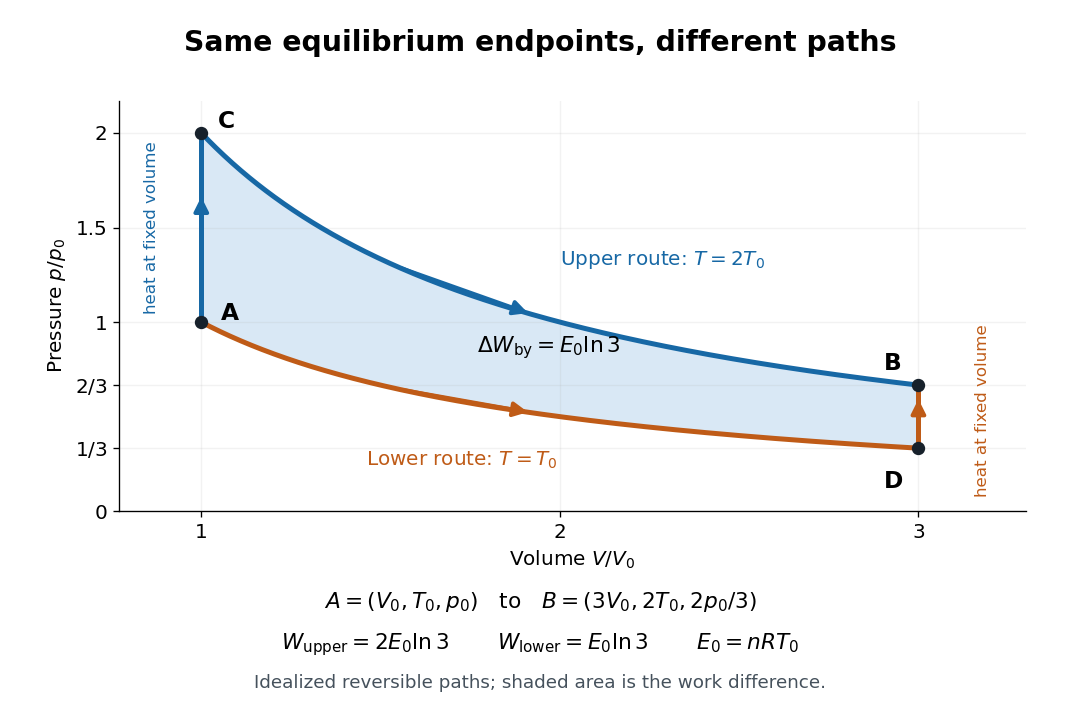
Two reversible ideal gas paths connect the same equilibrium endpoints
. Original idealized pressure-volume diagram. The upper route is and the lower route is ; the shaded gap is the difference in work, not an experimental uncertainty. Both are reversible thermodynamic processes under the stated piston and thermal reservoir assumptions. Details.The internal energy of an ideal gas gives the same endpoint difference for both routes:On either fixed-volume heating leg, and . On each isothermal expansion, , so the first law of thermodynamics gives . The ideal gas law and the integral of then giveHere is the natural logarithm. Rounded to , the upper route has and the lower route has . Subtracting separately rounded entries can differ by ; the unrounded balances are exact in the stipulated model.
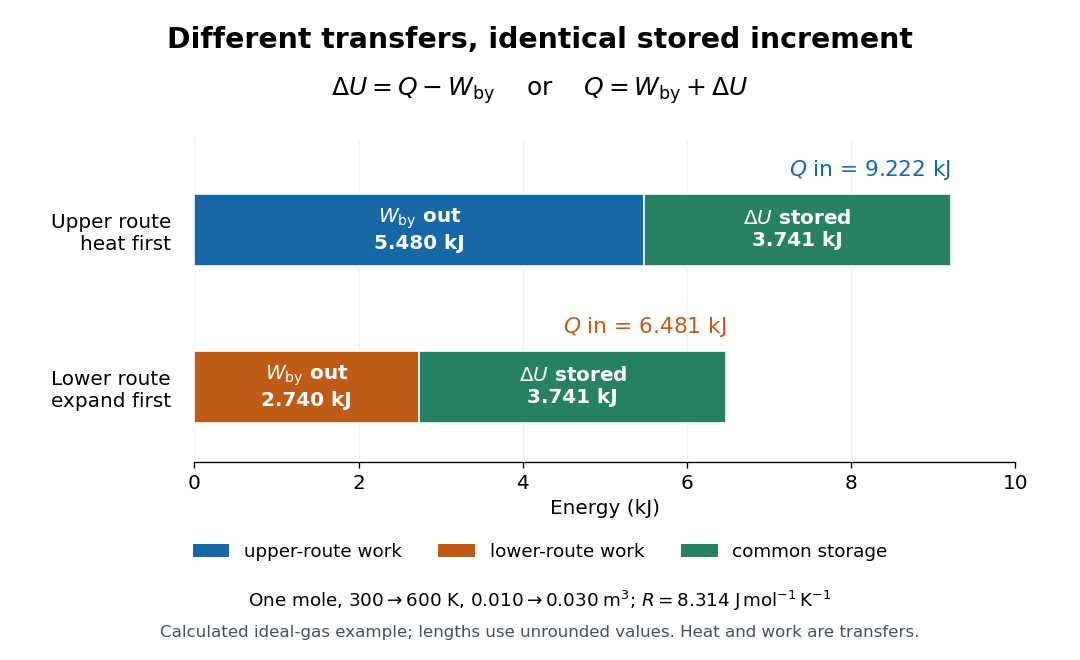
The shaded area between the curves explains the entire difference:Following the upper route forward and the lower route backward makes a thermodynamic cycle with this positive net work and matching net heat. Its internal energy change is zero. The two open routes themselves are reversible, so their different work outputs should not be mistaken for a loss caused by entropy production: they receive different heat transfers at different temperatures.
Two other state functions provide independent endpoint checks. With enthalpy in this model,To obtain the entropy change along either route, integrate on its heating leg, giving , and on its isothermal expansion, giving . Reordering the legs changes their heat totals but leaves the sum of their entropy changes unchanged. In particular, neither route's total divided by its final temperature equals , and neither total equals : both routes include an isothermal expansion, rather than a single fixed-pressure heating step.
The UCSB demonstration uses an aluminum cylinder turned by a crank, with a wrapped rope supporting a hanging mass. An embedded electrical thermometer connects through sliding contacts to an electrical resistance meter; a supplied electrical resistance–temperature table or fitted equation converts the readout. A counter records turns. Record the initial temperature, crank steadily, then stop and convert the minimum recorded electrical resistance to the final temperature.[ref]
52.33 -- Mechanical equivalent of heat. UCSB Physics Lecture Demonstrations; UCSB demonstration 52.33, opening apparatus paragraphs and “Operating the demonstration”: cylinder, rope, turn counter and resistance calibration
The proposed control is to start below room temperature and finish comparably above it, approximately balancing incoming and outgoing environmental heat. Neglected accessory heat capacities and rough temperature estimates limit accuracy. The mechanical input and thermal response test an energy balance; warming alone does not establish quantitative equivalence.[ref]
52.33 -- Mechanical equivalent of heat. UCSB Physics Lecture Demonstrations; UCSB demonstration 52.33, final operating paragraphs: symmetric temperature preparation, environmental heat exchange and omitted apparatus capacities
This is a modern companion to Joule's paddle-wheel experiment, with a different apparatus. Choose the cylinder as the closed system. When endpoint bulk mechanical energy changes are negligible, a model for its thermal storage iswhere is its participating heat capacity, approximately constant over the run, and is signed environmental heat. The cylinder is not an isolated system, since mechanical work enters. The temperature rise estimates a change in internal energy, rather than an amount of externally supplied heat. This boundary distinction connects a measured thermal response to the same stored quantity used in two reversible ideal-gas paths between the same states.
A heat engine uses a repeating thermodynamic cycle to turn part of an incoming heat transfer into net outgoing work. In the two-thermal reservoir model, the hot thermal reservoir supplies heat at fixed temperature , and the cold thermal reservoir receives heat of magnitude at fixed temperature . A Stirling engine makes the repeated motion visible; the abstract energy budget applies independently of its particular mechanism.[ref]
3.2 Generalized Representation of Thermodynamic Cycles. §3.2, Figure 3.3: two reservoirs and net work output
Choose the entire cyclic device as a closed system, with no net change in bulk kinetic energy or potential energy. Count heat entering as positive and net work delivered as positive, as in the first law of thermodynamics. Thus enters with a plus sign and , although a positive magnitude, enters with a minus sign. Assume these two heat transfers and the net work are the only transfers across the boundary. Over one complete thermodynamic cycle, all stored energy, including internal energy , returns to its starting value:The device does not accumulate heat from cycle to cycle. More precisely, heat is a transfer, while internal energy is what can be stored; even a component that temporarily stores internal energy must return to its initial state for this boundary to be cyclic.[ref]
For engine operation, and . Its thermal efficiency isUse net work, after subtracting any work supplied during compression or required by auxiliaries included in the chosen device. For periodic operation, average power satisfies and , with the rates averaged over complete thermodynamic cycles.[ref]
For a stipulated model at and , let and per thermodynamic cycle. Then and . At two thermodynamic cycles per second, these become incoming heat transfer, outgoing heat transfer, and net power. These are constructed values, not measurements.
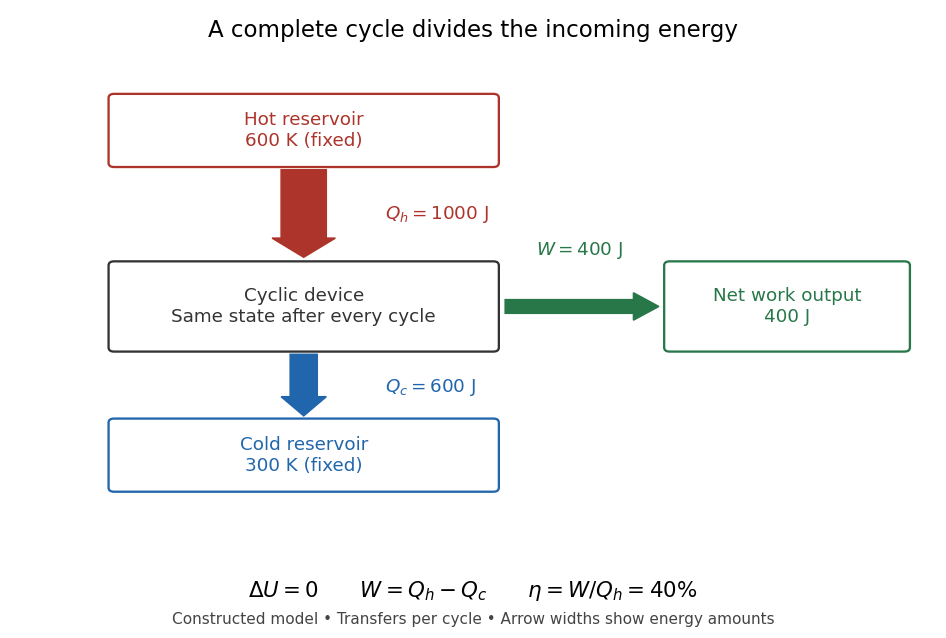
Different signs describe different tasks. With the same directions of heat transfer, gives : a device transmitting heat without net work output. If , net work is supplied to that device. A refrigerator or heat pump instead extracts heat from the cold side and delivers heat to the hot side, using positive input work and the balance . Their useful output is heat transfer, measured by a coefficient of performance. Finally, a one-time isothermal expansion can have and while leaving the volume changed: it is not a complete thermodynamic cycle. A warming device also fails the cyclic assumption, and a fuel-consuming device requires an accounting of matter crossing its boundary.
For two fixed thermal reservoirs at positive thermodynamic temperatures, the Second law of thermodynamics strengthens the energy balance toEquality requires the overall operation to be a reversible thermodynamic process, including its heat transfer to the thermal reservoirs. The Carnot cycle explains how the reversible benchmark is constructed, and Carnot's theorem explains its universality and the entropy production penalty. Use kelvin values, never ratios of degree Celsius readings. The example above lies below the bound: its total entropy production is per thermodynamic cycle.
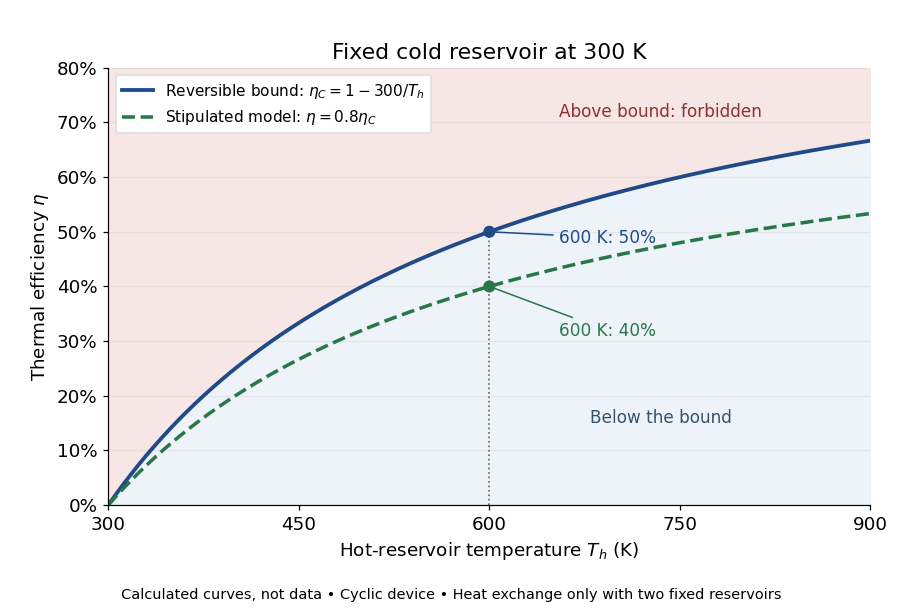
This is useful before building a machine: warmer cooling surroundings reduce the possible thermal efficiency, even if the mechanism is unchanged. A small observed thermal efficiency does not by itself test the Second law of thermodynamics; heat leaks, friction, incomplete recovery of work, and the measurement boundary can all reduce the reported ratio. A meaningful comparison requires the actual thermal reservoir temperatures, matched heat and net work measurements, and the hypotheses of Carnot's theorem. Finite bodies whose temperatures change are treated separately there.
The practical question already appears in the 1824 Réflexions sur la puissance motrice du feu et sur les machines propres à développer cette puissance by Sadi Carnot: pp. 6–8 ask whether machines driven by heat have a fundamental limit and whether changing the working substance can improve their output. Pages 9–11 describe heating and cooling in real machines, but use the historical caloric theory, rather than the modern first law of thermodynamics balance. The historical discussion under Carnot's theorem explains that distinction.
A vapour-power cycle using a liquid pump, high-pressure heat addition, turbine expansion and low-pressure condensation. Ideal components are reversible apart from heat transfer through finite temperature differences. With states numbered pump inlet, pump outlet, turbine inlet and turbine outlet, the net thermal efficiency is . Liquid pumping avoids the impractical compression of a wet two-phase mixture in a steam realization of the Carnot cycle. Named after William John Macquorn Rankine.
Consider an ideal-gas cycle with constant heat-capacity ratio : expand isobarically from to , cool isochorically, then compress reversibly and adiabatically to the initial state. With , closure gives and . The heat input is and the rejected heat magnitude is . Thus . Reservoir temperatures determine feasibility and entropy generation but need not coincide with the gas temperatures at heat-exchange endpoints.
The heat input to a cycle sums positive transfers of heat into its working substance. It is not the net heat over the entire cycle, which equals net work. A single geometric branch can contain both heat absorption and rejection, so grouping the branch into one net transfer can overestimate a heat engine's thermal efficiency.
An ideal cyclic heat engine producing no entropy overall. Between two equilibrium reservoirs its efficiency reaches the Carnot bound; finite reservoir temperatures can be treated as varying through successive reversible cycles.
A Stirling engine repeatedly heats and cools a confined gas, whose expansion and compression drive a moving mechanical load. This makes it a particularly accessible example of a heat engine: both the repeating motion and the two regions exchanging heat can be observed.
In the published Demonstration experiments with a Stirling engine by Christopher G Deacon and others, an electrical heater warms one end of a cylinder and flowing water cools the other at . A moving displacer shuttles gas through copper wool that temporarily stores and returns internal energy. A laser spot traces a closed pressure-volume path on a screen: mechanical motion controls one coordinate, pressure the other. Calibrating both scales converts the enclosed area to work per thermodynamic cycle; multiplying by rotation frequency gives gas-cycle power. At heater input, the paper reports gas-cycle power, while a separate Prony brake measures shaft output. The cold-side temperature is controlled and two higher heater settings are compared. The gas has a nonuniform temperature, and heater energy also escapes to the room: neither output/input ratio measures for an ideal two-thermal reservoir engine. These observations demonstrate work production and losses, not a direct measurement of the bound in Carnot's theorem. The inspected apparatus, readouts and results are on pp. 180–182, Figures 1–3 and Table 1.
For an ideal working gas and , two isotherms at produce net work . Without a regenerator, isochoric heating requires additional external heat , giving the displayed thermal efficiency, strictly below Carnot efficiency. A perfect reversible regenerator recovers this heat and attains the bound instead. Reversing the unregenerated cycle gives cooling coefficient of performance ; positive cooling requires a sufficiently large expansion ratio.
A Prony brake measures shaft power using a resisting band: its force difference gives torque , and rotation gives , where is angular velocity. The Stirling engine experiment above uses this independent load measurement to distinguish shaft output from gas-cycle work.
Heat-engine calorimetry measures heat transfers so that observed work can be compared with the correct energy input. Measuring motion alone cannot supply the denominator of thermal efficiency.
The institutional Laboratory 16: Stirling Engine describes a modern teaching arrangement: an ethanol lamp drives a Stirling engine coupled to an electrical output device, which supplies a resistor immersed in water in a calorimeter. The prescribed readouts include fuel mass consumed, the pressure-volume loop, electrical readings, and water temperature. The sequence separates fuel input, gas-cycle work, electrical output, and Joule heating of the water. Warming of the water is the final useful output; it is not the engine's rejected . The handout distinguishes warm-up from steady operation and asks students to compare the final warming with direct heating by the lamp. It gives a procedure, not a table of measured efficiencies. Its pp. 1–3 therefore establish an apparatus and measurement design without supplying a new experimental result.
For a proposed calibration, use a known input of electrical work to determine the effective heat capacity of the water, vessel and immersed parts; stir to make the measured temperature representative. A matched run with no input estimates exchange with the room. For constant and no phase transition, the resulting model iswhere is the signed net heat lost to the room during the measurement. Here is supplied electrical work when the resistor lies inside the calibration boundary; for a separate external heat input it is the received heat. These are proposed controls, not controls reported in that handout. Slow cooling, changing material properties and imperfectly matched room exchange limit this inference; the approach to thermal equilibrium and heat capacity matter as much as the thermometer reading.
To measure the actual engine-input , one must instrument its hot-side transfer separately, or close a calibrated energy balance around a specified heater boundary. A heater's electrical work includes changes in its stored internal energy and heat transfer that bypasses the engine. At periodic operation a cold-side flowing-water measurement can instead estimate , if mass flow rate and specific heat capacity at constant pressure are known, with negligible storage and other heat exchange in that measurement boundary. It measures only the rejection captured by that stream. Include all remaining transfers before using the residual to infer . This measurement discipline makes a comparison with Carnot's theorem meaningful.
Calorimetry measures heat transfers. A calorimeter, a calibrated heat capacity and an accounting of unwanted heat transfer connect an observed temperature change to an energy balance, as in heat-engine calorimetry and heat-pump calorimetry.
The central idea is to measure a response that is easier to observe than the transfer itself: a known heat capacity turns a temperature change into an energy estimate. Mixing calorimetry can determine an unknown specific heat capacity, while a chemical reaction can be studied by measuring the response of its surroundings. The useful observable is a change in temperature, together with a stated thermodynamic system boundary; a hot object does not contain a separately conserved store of heat.
For a closed system with unchanged bulk kinetic energy and potential energy, let be net inward heat and net inward work. The first law of thermodynamics givesIf the boundary includes a hot sample, receiving water and the participating vessel, their mutual heat transfer is internal. With no external input, the total storage change is minus the signed outward heat leakage, not the positive amount transferred from sample to water. A calorimeter calibration with an immersed resistor instead supplies electrical work across this boundary; Joule heating converts it into internal energy. Choosing only the receiving water as the boundary changes which transfers count as external.
The mechanical constraint decides which storage quantity the readout represents. For a closed system at fixed volume, with no other work, . At constant external pressure, with only expansion work , , where is enthalpy. Thus the corresponding warming balances use heat capacity at constant volume or heat capacity at constant pressure. Electrical or shaft work must be included separately when present. These endpoint balances do not require the actual exchange to be a reversible thermodynamic process.
A covered cup open to atmospheric pressure approximately realizes the second constraint when escaping material is negligible. A sealed rigid reaction vessel realizes the first; its surrounding water, vessel and immersed parts form the calibrated receiver. The reaction's outward heat equals the receiver's gain only after losses and ignition input have been accounted for. The distinction concerns the reaction boundary, even if its receiver is open to atmospheric pressure.[ref]
A historical quantitative result appears in James Prescott Joule's On the Mechanical Equivalent of Heat (1850): Table IX gives foot-pounds of mechanical work per warming of one pound of water by one degree Fahrenheit, with weighing reduced to vacuum. His final rounded statement specifies water between and degrees Fahrenheit. He acknowledged residual vibration and sound as possible upward bias. This is an original reported conversion result, not the definition of the modern joule; the apparatus and resting controls are described in Joule's paddle-wheel experiment.[ref]
On the Mechanical Equivalent of Heat by James Prescott Joule. p. 82, Table IX and concluding paragraphs: water-series equivalent and residual losses
Water calorimetry measures an energy transfer using the thermal response of water. In radiation metrology, it determines absorbed dose to water from a small measured temperature rise. The ideal conversion follows from and :where is water's specific heat capacity. The sample mass cancels because dose is energy per unit mass.[ref]
An Absorbed Dose Water Calorimeter: Theory, Design, and Performance by Steve R. Domen. Abstract and opening description: calibrated thermistors and temperature-rise dose measurement.
For an original scale estimate, take as an illustrative near-room-temperature value. An absorbed dose of would then give only , about . Detecting such a small change requires careful thermometry and separation from background thermal drift.
The physical conversion also has chemical assumptions. Heat defect describes the fraction of absorbed energy that does not appear as heat because of radiation-induced chemical change. Thermal conduction, convection and sensor effects are additional measurement questions, rather than alternate names for that chemical correction. This makes water calorimetry an example of an apparently direct energy measurement whose interpretation still needs a well-defined model.
The heat defect in water calorimetry measures the fraction of absorbed energy that does not appear as heat because of radiation-induced chemical changes. Writing absorbed energy as and the energy appearing as heat as , the convention used by Norman V. Klassen and Carl K. Ross isA positive value means less heat than the absorbed-energy equivalent; a negative value means the chemical response adds heat.[ref]
Water Calorimetry: The Heat Defect by Norman V. Klassen and Carl K. Ross. Printed p.63, Eqs1–2 and accompanying definition.
If other thermal corrections are treated separately, the absorbed dose isAn illustrative means that represents 98% of the dose, so the chemical correction multiplies that thermal estimate by . The example fixes a sign convention; it is not a measured universal value for water.
The motivating evidence included variations with dissolved gases and comparison of radiolysis calculations with chemical measurements of hydrogen peroxide. Those tests address chemical energy accounting. An external heat leak or a drifting thermometer can also alter a temperature reading, but belongs to a different correction mechanism.[ref]
Water Calorimetry: The Heat Defect by Norman V. Klassen and Carl K. Ross. Abstract and Introduction: radiolysis model and independently measured chemical product.
The sealed-water calorimeter of Steve R. Domen is an apparatus for measuring absorbed dose through the radiation-induced temperature rise of a controlled water volume. Domen's 1994 design enclosed the inner water in a thin-walled cylindrical glass vessel about 110 mm long and 33 mm in diameter, immersed in a larger water bath. The vessel also acted as a barrier against convection.[ref]
A Sealed Water Calorimeter for Measuring Absorbed Dose by Steve R. Domen. Section2.1, Design: vessel dimensions, surrounding water and convective-barrier role.
Two calibrated thermistors served as temperature sensors in opposite arms of a Wheatstone bridge, increasing the electrical response to their common temperature rise. The apparatus therefore connected a small thermal change to a measurable resistance change, rather than directly counting radiation events.[ref]
A Sealed Water Calorimeter for Measuring Absorbed Dose by Steve R. Domen. Introduction, Eq1 and Section2: two-thermistor bridge and temperature conversion.
A central experiment compared water fillings with different dissolved-gas conditions. The temperature response could change because radiation also drove chemical reactions, motivating investigation of the heat defect. This comparison illustrates a useful apparatus principle: controlling the geometry and reading the thermometer carefully does not, by itself, establish that every joule deposited in the water becomes measurable heat.[ref]
A Sealed Water Calorimeter for Measuring Absorbed Dose by Steve R. Domen. Abstract and results: repeated measurements with different fillings and heat-defect investigation.
Indirect calorimetry estimates an energy-release rate from measured chemical exchanges and a model relating those exchanges to chemical energy. Gas-analysis implementations measure oxygen disappearance and often carbon dioxide production; stoichiometry and energy equivalents then supply the inference. The physical measurement chain therefore contains both observations and assumptions, rather than a direct measurement of the entire heat transfer.
A controlled combustion system makes this logic concrete. A gas stream passes through a reaction region, exhaust flow and composition are measured, and the difference between incoming and outgoing oxygen amounts gives an uptake rate. In oxygen consumption calorimetry, that rate is combined with an appropriate energy-per-oxygen factor to estimate heat release rate. A falling oxygen fraction alone is insufficient: dilution and changes of total gas flow must also be accounted for.[ref]
National Fire Research Laboratory by NIST. Facility description, exhaust system and oxygen-consumption measurement of fire heat release rate
For a deliberately stipulated model with and measured oxygen uptake ,This factor is an assumption for the calculation, not a universal physical constant. Different reaction mixtures, incomplete oxidation or changing product composition can require a different model. Gas volumes must likewise be tied to stated temperature and pressure before they become comparable amounts.
An independent reference tests the chain. NIST describes natural-gas reference burners whose fuel-consumption-based heat input cross-checks its oxygen-consumption calorimetry system. The agreement then tests gas sampling, flow measurement and the adopted energy accounting together, rather than merely checking one sensor.[ref]
Natural Gas Confirmation Burner by NIST. Research Overview, reference burners and mutual confirmation of fuel- and oxygen-consumption calorimetry
Oxygen consumption calorimetry estimates combustion heat release rate from the rate at which a fire consumes oxygen. Its practical usefulness comes from an approximate regularity: many common combustible materials release roughly per kilogram of consumed oxygen. The coefficient is an empirical energy equivalent for an applicable class of fuels, not a universal physical constant.[ref]
Standard Practice for Full-Scale Oxygen Consumption Calorimetry Fire Tests (ASTM E2067-20) by ASTM International. Public Significance and Use, §4.1: approximate energy release per unit mass of consumed oxygen
Let be the consumed oxygen mass flow rate, obtained from a consistent incoming/outgoing gas balance. A simplified model isFor an illustrative measured consumption rate of , this model givesThe observable must include gas throughput: an oxygen fraction by itself cannot determine a consumed mass per unit time.
In an exhaust-collection apparatus, a hood captures combustion products together with entrained air. Measurements of exhaust flow and gas composition allow the consumed oxygen rate to be inferred despite dilution. Gas sampling, mixing and the flow measurement must represent the same stream; gas amounts also need consistent wet/dry conventions and reference temperature and pressure. The NIST National Fire Research Laboratory uses exhaust systems to measure large-fire heat release rate by this principle.[ref]
National Fire Research Laboratory by NIST. Facility description, exhaust collection and oxygen-consumption calorimetry
[ref]Standard Practice for Full-Scale Oxygen Consumption Calorimetry Fire Tests (ASTM E2067-20) by ASTM International. Public Significance and Use, §4.3: oxygen concentration and exhaust-flow measurements
A useful experimental check is an independently metered reference burner. Its fuel-consumption-based energy input can be compared with the heat release rate inferred from oxygen depletion; NIST describes natural-gas burners for this mutual confirmation. This tests the measurement chain rather than merely exposing a gas analyzer to a known mixture.[ref]
Natural Gas Confirmation Burner by NIST. Research Overview, comparison of fuel-consumption and oxygen-consumption calorimetry
Incomplete capture, changing fuel composition, incomplete combustion and analyzer transport delays can all affect the inferred curve. They concern different parts of the inference: lost exhaust biases the gas balance, an inappropriate energy equivalent changes the conversion, and delayed sampling shifts the response in time. A reported heat release rate therefore needs both its conversion assumptions and its measurement conditions.
Heat release rate is the rate at which a process releases energy as heat, commonly used to characterize combustion. It has the dimensions of power and is measured in watts, with . Using for cumulative released energy,The heat release rate is the height of the curve; total released energy is its area.
For a synthetic fire whose heat release rate rises linearly from zero to in and falls linearly to zero over the next , the area is a triangle:A second fire with a smaller peak but longer duration can release more total energy. A peak value alone therefore cannot describe the full event.
A fuel-based estimate multiplies fuel mass flow rate by the effective released energy per unit mass, under stated assumptions about composition and combustion. For a deliberately stipulated effective value and fuel rate , the estimate is . In oxygen consumption calorimetry, measured gas flow and oxygen depletion instead provide an estimate of heat release rate. NIST uses independently metered reference burners to compare these two inference routes.[ref]
Natural Gas Confirmation Burner by NIST. Research Overview, fuel-consumption heat input and confirmation of oxygen-consumption calorimetry
A fire's heat release rate differs from the heat transfer received by one nearby surface or from the temperature of its exhaust. Released energy can leave by several paths, while a receiving object can store internal energy as well as lose heat. Even a gas-based calorimeter needs a defined measurement boundary, capture conditions and time response. The NIST National Fire Research Laboratory provides an example of using exhaust collection to quantify large-fire heat release rate.[ref]
National Fire Research Laboratory by NIST. Facility description, large-fire calorimetry and exhaust systems
Direct calorimetry measures a process's thermal output or absorption through the response of a calorimeter. The observations can be a receiver's temperature change or a measured heat transfer rate across a controlled boundary. Converting those observations into energy or power requires a calibrated thermal response and an accounting of storage and losses.
A flow calorimeter provides a concrete physical arrangement. A heated component transfers energy to a coolant stream; inlet and outlet thermometers measure its temperature rise, while a flow measurement supplies the mass flow rate. With negligible changes in bulk kinetic energy and height, the stream's enthalpy gain isHere is specific heat capacity at the relevant pressure and temperature. Manufacturer documentation for a water calorimeter likewise describes monitoring water flow and temperature to determine absorbed incident power.[ref]
Water calorimeter power detectors by Gentec-EO. Publisher description, water-flow and temperature monitoring for calorimetric power measurement
For an illustrative water flow of , a rise and approximately ,Over 60 seconds at those conditions, the stream carries away about . These assumed values describe a calculation, not a product specification.
A broader energy balance can readIf the vessel is still warming, some input is stored rather than carried away. Heat leakage can also leave the chosen boundary through another path. Electrical calorimetry can check the response by supplying a measured heater input and comparing it with the readout. Establishing a steady state, matched thermometer calibration and a clear thermodynamic system boundary makes the inferred source power credible.
A flow calorimeter uses a flowing material to carry energy from a measurement region. In a common arrangement, a heated receiver exchanges energy with a coolant stream. Measurements of mass flow rate and inlet/outlet temperatures determine the stream's enthalpy gain, while the vessel and environmental paths are included in the overall balance.[ref]
Water calorimeter power detectors by Gentec-EO. Publisher explanation, measuring water flow and temperature to determine absorbed power
For a steady stream with negligible changes in bulk kinetic energy and height,The approximation uses a suitable approximately constant specific heat capacity. An absorber, inlet and outlet thermometers, and a calibrated flow measurement turn this equation into an apparatus rather than just a temperature reading.
For a synthetic example, let and . A rise gives . If a 30-watt electrical reference produces this response under matched conditions, the observed response fraction is . Under an explicitly tested proportional-response assumption, a later 36-watt fluid reading would correspond to a 45-watt source.
A single calibration point does not establish that assumption at every power. The full balance includes receiver storage and other losses:Warm-up, leakage, thermometer offsets and transport delays can all affect a time-varying measurement. Calorimeter calibration and electrical calorimetry can test the response across a range, with the same thermodynamic system boundary used for calibration and the unknown source.
Use measured electrical energy input to calibrate or infer a thermal response. A heater's terminal electric potential difference and electric current determine the supplied energy; receiver heat capacity and losses must also be accounted for.
Infer an unknown heat capacity by thermal contact with a calibrated receiver and a measured common final temperature. The energy lost by hotter components equals the energy gained by colder components, with the calorimeter and surrounding heat transfer included. For appreciably varying heat capacities, use temperature integrals rather than constant terms.
differential scanning calorimetry measures differences in heat flow between sample and reference during a controlled temperature scan. Transition peaks can reveal cooperative lipid-chain melting, with scan rate, composition and reversibility affecting interpretation.
Mixing calorimetry determines heat capacities by bringing bodies at different temperatures into thermal contact. The striking feature is that the final temperature is a weighted arithmetic mean with heat capacities as weights, rather than weights given by mass. The vessel participates in the exchange even though it is not the sample being studied.
For a hot nonreacting sample of mass , initial temperature and unknown specific heat capacity at constant pressure , let receiving water have mass , initial temperature and known specific heat capacity at constant pressure . Let the vessel and immersed parts start at and have combined heat capacity , excluding that water. Assume a common final temperature , approximately constant pressure and heat capacities, no phase transition, no chemical reaction and negligible external nonexpansion work. For signed outward heat loss , the combined enthalpy balance isHenceThe loss is positive if the room receives net heat, negative if the room supplies it. If the vessel starts at a different temperature , replace its warming term by ; do not assume the vessel shared the water's initial temperature. For variable material properties, replace each by the corresponding integral and each vessel term by . The inferred constant then represents an average over the sample's cooling interval.[ref]
For a synthetic no-loss example, mix of hot water at with of cold water at in a vessel initially at . Stipulate and . The enthalpy balance givesThe hot water releases : the cold water receives and the vessel . Omitting the vessel predicts , despite perfect insulation. These are computed numbers from stipulated properties, not laboratory observations.
The vessel changes the mixing temperature by sharing the heat transfer
. Original computed no-loss water example. Left: transfers sum to the hot portion's released heat. Right: increasing the initially cold vessel's heat capacity lowers the common final temperature; the marked point uses . Each vessel setting is a separate modeled experiment. Details.For an unknown-sample example using the same cold water and vessel, stipulate , and . With no loss, . Inverting the full balance returns ; ignoring vessel storage returns about . With a positive outward loss, ignoring that loss also biases the inferred downward under these hot-sample conditions. A small water temperature rise makes sensor offsets and loss estimates relatively influential; propagation of uncertainty must include the measured temperatures, masses and calibrated vessel term.
A calorimeter infers transferred heat from a measured change, such as a temperature change and a calibrated heat capacity. The heat-engine calorimetry example requires accounting for the vessel and heat transfer to the surroundings as well as the water.
A practical receiver has an inner vessel, insulation or an outer shield, a lid, a thermometer and a stirrer. Its participating heat capacity includes the immersed parts that follow the water's temperature. Insulation reduces heat transfer to the room; stirring makes a single reading more representative of the contents. These serve different purposes: a well-insulated vessel can still have substantial heat capacity, and a uniform temperature does not imply zero external heat transfer.
The University of Toronto's Heat Capacity (University of Toronto laboratory handout) describes a copper inner vessel, outer shield, thermometer and heated metal block. Weigh the block and receiving water, heat the block in a bath, then transfer it quickly into enough water to cover it. The observed temperature rises, reaches a maximum, then falls. The inferred specific heat capacity comes from the block's cooling and the warming of both water and vessel. In its cooling exercise the handout deliberately exposes the inner vessel to room air, recommends readings every , and uses the later common cooling to estimate the earlier loss. Starting below room temperature reduces exchange without guaranteeing cancellation. The handout notes limited time and thermometry for small corrections; it supplies no dataset.[ref]
Heat Capacity (University of Toronto laboratory handout). pp. 1–3, The Method of Mixtures and Cooling Correction, Figure 1
For a proposed modern version, independently perform thermometer calibration and calorimeter calibration, record bath and room temperatures, and keep immersion depth, lid position and stirring rate fixed. Ensure the block actually reaches bath temperature; minimize and account for adhering hot bath water. Compare repeated transfers with different delays and a reference sample of known specific heat capacity. A resting run tests environmental heat transfer, while a stirred run with no added sample checks mechanical input. These are proposed controls, not claimed observations from the handout. Sensor delay, changing heat capacity, incomplete thermal equilibrium and material escaping the boundary limit the resulting estimate.
A calorimeter that infers transferred heat from ice melting at a specified equilibrium temperature. The latent heat sets the proportionality after correcting for other thermal changes and extraneous melting.
Calorimeter calibration establishes the response of the actual measurement assembly to a known energy input. It determines a participating heat capacity, not just the empty vessel's mass. For an electrical calibration with an immersed resistor, measure voltage across that resistor and electric current through it. If that electrical work is deposited inside the chosen boundary, its input isFor a covered receiver at constant pressure, describes enthalpy storage; for a rigid fixed-volume assembly, use the matching heat capacity at constant volume. Negligible endpoint mechanical storage and no escaping material are assumed. Thus overestimates if outward loss is ignored. Account for lead losses outside the boundary and maintain the same water charge and immersed hardware in calibration and sample runs.
An alternative calibration mixes known hot and cold water portions. With the vessel initially at the cold portion's temperature , the mixing calorimetry balance givesThe previous synthetic water example returns . The sign matters: attributing room losses to the vessel makes this estimate too large. With both and the loss unknown, one final temperature supplies only one equation; separate cooling information or controlled independent runs are needed.
A calorimeter heat-loss correction can make the electrical balance quantitative. For a second synthetic example, stipulate of water with and immersed hardware with , so . Supply for , starting at a room temperature of , with loss coefficient . The model below gives , storage and outward loss . The uncorrected ratio gives about ; subtracting the loss recovers .
Electrical input splits into thermal storage and room losses during calibration
. Original synthetic computed traces for the stated constant-heat capacity, constant-loss model. The heater switches off at ; stored energy then declines while cumulative outward heat increases. The no-loss comparison reaches . No points represent experimental measurements. Details.A calorimeter heat-loss correction estimates the signed heat exchanged with the room during the measurement. Under Newton's law of cooling, a constant room temperature and a single representative assembly temperature give the modelHere is deposited input power, and is the apparatus loss coefficient in . After input ceases, the temperature excess shows exponential decay with time constant :A cooling trace determines the ratio , not the two factors separately. Combining the cooling trace with a known electrical energy input supplies the missing information. For constant input , starting at , the heating solution is ; these solutions generate the calibration figure.
For mixing calorimetry, infer cooling behavior only after the sample, water and vessel approach a common temperature. Applying a single-body cooling model while the sample remains much hotter than the water can misestimate the earlier loss. The Toronto handout's area correction assumes the same loss mechanism represents both the mixing interval and subsequent cooling.[ref]
Heat Capacity (University of Toronto laboratory handout). p. 3, Figure 1 and the area-ratio cooling correction
A backward extrapolation to the transfer time is therefore a model-dependent estimate, not the ultimate equilibrium with the room. Extend the model or report a limitation if sensor delay, substantial temperature gradients, changing room temperature, or escaping material invalidate that assumption. In particular, the relevant after adding a sample includes its heat capacity; an empty-receiver cooling rate need not apply to the loaded receiver.The historical caloric theory treated heat as a conserved substance. The modern first law of thermodynamics conserves energy, allowing net heat input to become work in a heat engine.
Rumford's cannon-boring experiment tested whether frictional heating could be explained as release of a finite store of caloric. In his 1798 paper, Count Rumford describes rotating gun-metal against a borer while enclosing the heated cylinder in a water-filled wooden box. In Experiment No. 3, the cylinder turned at about revolutions per minute, and the surrounding water boiled after hours minutes.[ref]
An inquiry concerning the source of the heat which is excited by friction by Benjamin Thompson. Experiment No. 3: water-tight box, rotation rate and temperature record
A separate control compared hot metal chips with thin strips cut from the same metal. Mixing equal masses into equal quantities of cold water gave nearly equal final temperatures, challenging an explanation based on a large change in specific heat capacity during cutting.[ref]
An inquiry concerning the source of the heat which is excited by friction by Benjamin Thompson. Opening metal-chip/strip calorimetry comparison and repeated trials
Continued mechanical input, rather than depletion of an evident material heat store, was the striking result. A modern quantitative energy balance must also include warming of the apparatus, environmental losses and measured mechanical work; boiling by itself does not determine an exact mechanical equivalent of heat.The hypothetical heat substance of caloric theory. The modern distinction between heat transfer and conserved energy replaces that substance picture.
A refrigerator removes heat from the region we want to keep cold and releases heat to warmer surroundings. Here we consider a work-driven cyclic device: its useful output is the cold-side extraction , and its input is net work . A food compartment is a familiar example; the same cooling objective applies when removing heat from a room. The coefficient of performance measures extraction per unit work, rather than the fraction of input energy converted into output. [ref]
https://openstax.org/books/university-physics-volume-2/pages/4-3-refrigerators-and-heat-pumps. Ling, Moebs and Sanny, University Physics Volume 2, §4.3: refrigerators and useful output
Choose the complete machine as the thermodynamic system. Let be heat entering from the colder side, be heat delivered to the hotter side, and be net work supplied, all over the same complete cycle. Assume these are its only energy transfers, no matter crosses the outer boundary, and there is no net change in stored energy. Returning to the initial state gives , so the first law of thermodynamics readsThe heat leaving the warmer side includes both the extracted heat and the supplied work. For instance, stipulated transfers and require ; the cooling coefficient of performance is . This is an illustrative budget, not measured performance. At steady operation the same balance holds for average transfer rates, , where is input power.
This budget explains a useful counterexample: operating a refrigerator with its door open inside an otherwise isolated room cannot cool the room as a whole. If both heat transfers occur within the room and the electrical input comes from outside, the room gains each cycle. Locally cold air does not imply net cooling. The Second law of thermodynamics permits cold-to-hot transfer because work is supplied; it forbids a cyclic device whose sole effect is that transfer. See the Clausius statement of the second law.
A Carnot refrigerator runs a reversible Carnot cycle in the direction that removes heat from a cold reservoir at and delivers to a hot reservoir at . It requires supplied work . Zero total entropy production gives .
Its coefficient of performance, defined as useful cooling divided by supplied work, is thereforeAll temperatures here are absolute. The ratio can exceed one because supplied work moves heat rather than being the only energy delivered to the cold-side process. For reservoirs at and , the reversible value is five. A real refrigerator produces entropy and requires more work for the same cooling between those reservoirs. The ideal result is a limit, not a claim of finite cooling power without thermal gradients.
Vapor-compression refrigeration circulates a working fluid through cold and hot coils. In the cold coil, liquid becomes gas while absorbing heat from the refrigerated region. Compression supplies work and raises the pressure and temperature; the hot coil releases heat as the gas returns to liquid. A restriction reduces the pressure before the next cold-side passage. Changing the pressure changes the temperature of the phase transition, letting one circulating fluid receive and release heat at different temperatures. This physical implementation serves both refrigerators and heat pumps; its ordinary dissipative operation differs from the reversible Carnot cycle used to bound performance. [ref]
https://openstax.org/books/university-physics-volume-2/pages/4-3-refrigerators-and-heat-pumps. Ling, Moebs and Sanny, University Physics Volume 2, §4.3, Figure 4.7: circulating coolant
[ref]A heat pump uses work to deliver heat to the region we want to keep warm. Its useful output is the hot-side delivery . The same operating machine can be described as a refrigerator when the cold-side extraction is the objective: which output is useful changes the coefficient of performance, while the energy balance stays . Heating a water bath by taking heat from a colder water bath makes this distinction visible in the heat-pump calorimetry experiment below. [ref]
https://labs.phys.utk.edu/mbreinig/phys136core/modules/m3/refrigerators.html. University of Tennessee, Refrigerators, air conditioners, and heat pumps: heating objective
In the illustrative budget from refrigerator, supplying of work delivers of useful heat, giving a heating coefficient of performance of . The additional came from the colder region. Compare an ideal electrical heater whose entire input work becomes heat in the target region: it delivers for that input. The appeal of a heat pump is this ability to move existing energy, while the Second law of thermodynamics limits the work saving for a given pair of temperatures.
One energy budget, two useful outputs, and Carnot's theorem bounds on the coefficient of performance
. Original quantitative diagram, with stipulated , and . Arrow widths follow the transfer magnitudes. The curves are reversible bounds for fixed cold temperature and varying hot temperature ; they are not measured performance. At , the bounds are for cooling and for heating, above the illustrative machine's and . Details.For a sinusoidal ground-temperature model whose coldest surface day is , source temperature on that day is , where . The first and largest thermal maximum occurs at , not at infinite depth. This maximizes the winter coefficient of performance for a reversible pump with a fixed indoor temperature. With a heating load even about that day, the same optimum minimizes annually integrated reversible heating work. Excavation cost, finite heat extraction and real heat-pump behaviour can change the optimum.
A reversible heat pump maintaining against an external has heating coefficient of performance . If its maximum work input is and the building loses heat at rate , maintaining equilibrium requires . This yields the displayed minimum external temperature, provided it lies in the physical positive-temperature range. If the rating instead specifies maximum delivered heat, the bound is ; a heat-output rating cannot be substituted for a work-input rating in the Carnot formula.
Articles were limited to the first 100 out of 720 total.
Articles by others on the same topic

Thermodynamics is a branch of physics that deals with the relationships between heat, work, temperature, and energy. It encompasses the study of how energy is transferred and transformed, and how these processes influence matter, particularly in terms of its macroscopic properties and behavior. Thermodynamics is governed by four fundamental laws: 1. **Zeroth Law of Thermodynamics**: This law establishes the concept of temperature and thermal equilibrium.



